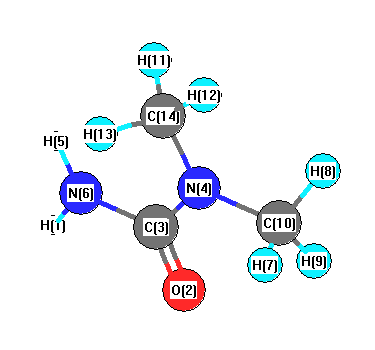
Vibrational Frequencies calculated at BLYP/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3514 |
3520 |
14.35 |
|
|
|
| 2 |
A |
3408 |
3413 |
6.48 |
|
|
|
| 3 |
A |
3081 |
3086 |
1.68 |
|
|
|
| 4 |
A |
3045 |
3050 |
9.66 |
|
|
|
| 5 |
A |
2974 |
2978 |
30.60 |
|
|
|
| 6 |
A |
2957 |
2962 |
59.30 |
|
|
|
| 7 |
A |
2914 |
2918 |
84.60 |
|
|
|
| 8 |
A |
2882 |
2886 |
82.73 |
|
|
|
| 9 |
A |
1702 |
1705 |
316.90 |
|
|
|
| 10 |
A |
1557 |
1559 |
89.35 |
|
|
|
| 11 |
A |
1477 |
1479 |
50.08 |
|
|
|
| 12 |
A |
1437 |
1440 |
12.19 |
|
|
|
| 13 |
A |
1429 |
1431 |
16.41 |
|
|
|
| 14 |
A |
1427 |
1430 |
8.38 |
|
|
|
| 15 |
A |
1410 |
1412 |
43.01 |
|
|
|
| 16 |
A |
1367 |
1369 |
8.15 |
|
|
|
| 17 |
A |
1361 |
1363 |
147.19 |
|
|
|
| 18 |
A |
1245 |
1247 |
34.20 |
|
|
|
| 19 |
A |
1222 |
1224 |
34.06 |
|
|
|
| 20 |
A |
1122 |
1124 |
2.33 |
|
|
|
| 21 |
A |
1083 |
1085 |
3.94 |
|
|
|
| 22 |
A |
1064 |
1066 |
88.81 |
|
|
|
| 23 |
A |
1041 |
1043 |
5.04 |
|
|
|
| 24 |
A |
985 |
986 |
34.09 |
|
|
|
| 25 |
A |
750 |
752 |
71.33 |
|
|
|
| 26 |
A |
737 |
738 |
6.28 |
|
|
|
| 27 |
A |
602 |
603 |
123.52 |
|
|
|
| 28 |
A |
570 |
571 |
20.60 |
|
|
|
| 29 |
A |
496 |
497 |
8.31 |
|
|
|
| 30 |
A |
410 |
410 |
17.54 |
|
|
|
| 31 |
A |
374 |
375 |
16.10 |
|
|
|
| 32 |
A |
307 |
308 |
9.19 |
|
|
|
| 33 |
A |
189 |
190 |
2.97 |
|
|
|
| 34 |
A |
152 |
152 |
0.87 |
|
|
|
| 35 |
A |
133 |
133 |
4.24 |
|
|
|
| 36 |
A |
88 |
88 |
1.88 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 25254.6 cm
-1
Scaled (by 1.0016) Zero Point Vibrational Energy (zpe) 25295.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at BLYP/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
H |
0.086 |
|
|
|
| 2 |
O |
-0.265 |
|
|
|
| 3 |
C |
0.134 |
|
|
|
| 4 |
N |
-0.203 |
|
|
|
| 5 |
H |
0.079 |
|
|
|
| 6 |
N |
-0.092 |
|
|
|
| 7 |
H |
0.012 |
|
|
|
| 8 |
H |
0.007 |
|
|
|
| 9 |
H |
0.035 |
|
|
|
| 10 |
C |
0.086 |
|
|
|
| 11 |
H |
0.024 |
|
|
|
| 12 |
H |
0.009 |
|
|
|
| 13 |
H |
0.015 |
|
|
|
| 14 |
C |
0.074 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.500 |
2.931 |
1.093 |
3.469 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-33.926 |
4.300 |
2.092 |
| y |
4.300 |
-37.790 |
1.287 |
| z |
2.092 |
1.287 |
-37.194 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.566 |
4.300 |
2.092 |
| y |
4.300 |
-2.230 |
1.287 |
| z |
2.092 |
1.287 |
-1.336 |
|
| Polar |
|---|
| 3z2-r2 | -2.672 |
|---|
| x2-y2 | 3.864 |
|---|
| xy | 4.300 |
|---|
| xz | 2.092 |
|---|
| yz | 1.287 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.723 |
0.014 |
0.004 |
| y |
0.014 |
8.708 |
0.258 |
| z |
0.004 |
0.258 |
5.420 |
<r2> (average value of r
2) Å
2
| <r2> |
0.000 |
| (<r2>)1/2 |
0.000 |