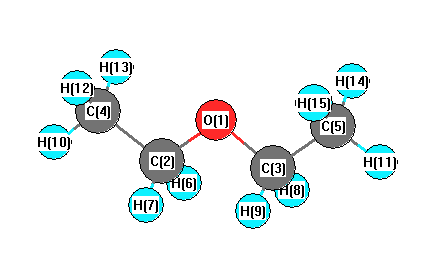
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3071 |
3053 |
26.72 |
|
|
|
| 2 |
A1 |
2982 |
2965 |
10.08 |
|
|
|
| 3 |
A1 |
2874 |
2857 |
116.25 |
|
|
|
| 4 |
A1 |
1462 |
1454 |
1.16 |
|
|
|
| 5 |
A1 |
1427 |
1419 |
3.08 |
|
|
|
| 6 |
A1 |
1387 |
1379 |
6.22 |
|
|
|
| 7 |
A1 |
1330 |
1322 |
0.12 |
|
|
|
| 8 |
A1 |
1138 |
1131 |
17.52 |
|
|
|
| 9 |
A1 |
1041 |
1035 |
14.32 |
|
|
|
| 10 |
A1 |
832 |
827 |
6.17 |
|
|
|
| 11 |
A1 |
431 |
428 |
0.03 |
|
|
|
| 12 |
A1 |
187 |
185 |
0.46 |
|
|
|
| 13 |
A2 |
3068 |
3051 |
0.00 |
|
|
|
| 14 |
A2 |
2895 |
2878 |
0.00 |
|
|
|
| 15 |
A2 |
1403 |
1395 |
0.00 |
|
|
|
| 16 |
A2 |
1234 |
1227 |
0.00 |
|
|
|
| 17 |
A2 |
1115 |
1109 |
0.00 |
|
|
|
| 18 |
A2 |
791 |
787 |
0.00 |
|
|
|
| 19 |
A2 |
245 |
244 |
0.00 |
|
|
|
| 20 |
A2 |
106 |
105 |
0.00 |
|
|
|
| 21 |
B1 |
3069 |
3051 |
43.91 |
|
|
|
| 22 |
B1 |
2889 |
2872 |
148.34 |
|
|
|
| 23 |
B1 |
1403 |
1395 |
12.15 |
|
|
|
| 24 |
B1 |
1236 |
1229 |
3.19 |
|
|
|
| 25 |
B1 |
1148 |
1142 |
12.61 |
|
|
|
| 26 |
B1 |
800 |
796 |
1.55 |
|
|
|
| 27 |
B1 |
249 |
247 |
1.38 |
|
|
|
| 28 |
B1 |
103 |
102 |
2.92 |
|
|
|
| 29 |
B2 |
3071 |
3053 |
13.71 |
|
|
|
| 30 |
B2 |
2982 |
2964 |
26.84 |
|
|
|
| 31 |
B2 |
2860 |
2844 |
13.37 |
|
|
|
| 32 |
B2 |
1440 |
1431 |
2.51 |
|
|
|
| 33 |
B2 |
1424 |
1415 |
1.72 |
|
|
|
| 34 |
B2 |
1345 |
1337 |
39.86 |
|
|
|
| 35 |
B2 |
1309 |
1302 |
39.63 |
|
|
|
| 36 |
B2 |
1133 |
1126 |
219.53 |
|
|
|
| 37 |
B2 |
1064 |
1058 |
0.80 |
|
|
|
| 38 |
B2 |
926 |
921 |
6.44 |
|
|
|
| 39 |
B2 |
418 |
416 |
3.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 28942.9 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 28775.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.282 |
|
|
|
| 2 |
C |
0.069 |
|
|
|
| 3 |
C |
0.069 |
|
|
|
| 4 |
C |
-0.060 |
|
|
|
| 5 |
C |
-0.060 |
|
|
|
| 6 |
H |
0.009 |
|
|
|
| 7 |
H |
0.009 |
|
|
|
| 8 |
H |
0.009 |
|
|
|
| 9 |
H |
0.009 |
|
|
|
| 10 |
H |
0.030 |
|
|
|
| 11 |
H |
0.030 |
|
|
|
| 12 |
H |
0.042 |
|
|
|
| 13 |
H |
0.042 |
|
|
|
| 14 |
H |
0.042 |
|
|
|
| 15 |
H |
0.042 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-0.869 |
0.869 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-33.206 |
0.000 |
0.000 |
| y |
0.000 |
-30.806 |
0.000 |
| z |
0.000 |
0.000 |
-33.632 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.987 |
0.000 |
0.000 |
| y |
0.000 |
2.613 |
0.000 |
| z |
0.000 |
0.000 |
-1.626 |
|
| Polar |
|---|
| 3z2-r2 | -3.253 |
|---|
| x2-y2 | -2.400 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.977 |
0.000 |
0.000 |
| y |
0.000 |
9.664 |
0.000 |
| z |
0.000 |
0.000 |
7.357 |
<r2> (average value of r
2) Å
2
| <r2> |
178.110 |
| (<r2>)1/2 |
13.346 |