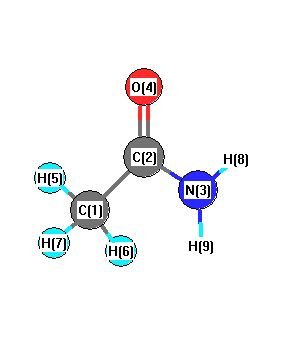
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3613 |
3592 |
22.05 |
|
|
|
| 2 |
A |
3471 |
3451 |
14.21 |
|
|
|
| 3 |
A |
3109 |
3091 |
4.41 |
|
|
|
| 4 |
A |
3056 |
3038 |
10.52 |
|
|
|
| 5 |
A |
2979 |
2961 |
7.23 |
|
|
|
| 6 |
A |
1756 |
1746 |
264.31 |
|
|
|
| 7 |
A |
1536 |
1527 |
95.10 |
|
|
|
| 8 |
A |
1408 |
1400 |
8.95 |
|
|
|
| 9 |
A |
1393 |
1384 |
10.06 |
|
|
|
| 10 |
A |
1337 |
1329 |
112.52 |
|
|
|
| 11 |
A |
1282 |
1274 |
29.52 |
|
|
|
| 12 |
A |
1082 |
1076 |
0.35 |
|
|
|
| 13 |
A |
995 |
989 |
3.66 |
|
|
|
| 14 |
A |
932 |
926 |
4.52 |
|
|
|
| 15 |
A |
826 |
821 |
3.83 |
|
|
|
| 16 |
A |
631 |
627 |
8.90 |
|
|
|
| 17 |
A |
536 |
533 |
15.36 |
|
|
|
| 18 |
A |
497 |
494 |
7.49 |
|
|
|
| 19 |
A |
404 |
401 |
3.70 |
|
|
|
| 20 |
A |
253 |
251 |
178.38 |
|
|
|
| 21 |
A |
49 |
48 |
0.43 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 15570.8 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 15480.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.055 |
|
|
|
| 2 |
C |
0.017 |
|
|
|
| 3 |
N |
-0.077 |
|
|
|
| 4 |
O |
-0.232 |
|
|
|
| 5 |
H |
0.056 |
|
|
|
| 6 |
H |
0.042 |
|
|
|
| 7 |
H |
0.061 |
|
|
|
| 8 |
H |
0.099 |
|
|
|
| 9 |
H |
0.090 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.111 |
-3.518 |
0.514 |
3.557 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-20.822 |
-2.305 |
0.572 |
| y |
-2.305 |
-25.066 |
-0.750 |
| z |
0.572 |
-0.750 |
-24.563 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.993 |
-2.305 |
0.572 |
| y |
-2.305 |
-2.374 |
-0.750 |
| z |
0.572 |
-0.750 |
-1.619 |
|
| Polar |
|---|
| 3z2-r2 | -3.238 |
|---|
| x2-y2 | 4.244 |
|---|
| xy | -2.305 |
|---|
| xz | 0.572 |
|---|
| yz | -0.750 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.638 |
-0.146 |
0.037 |
| y |
-0.146 |
5.688 |
-0.024 |
| z |
0.037 |
-0.024 |
3.228 |
<r2> (average value of r
2) Å
2
| <r2> |
75.888 |
| (<r2>)1/2 |
8.711 |