Jump to
S1C2
Energy calculated at QCISD/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -190.134287 |
| Energy at 298.15K | |
| HF Energy | -189.550222 |
| Nuclear repulsion energy | 87.392945 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3123 |
2997 |
25.39 |
|
|
|
| 2 |
A1 |
2307 |
2213 |
804.95 |
|
|
|
| 3 |
A1 |
1781 |
1709 |
36.56 |
|
|
|
| 4 |
A1 |
1484 |
1424 |
5.23 |
|
|
|
| 5 |
A1 |
920 |
882 |
3.85 |
|
|
|
| 6 |
B1 |
993 |
953 |
20.54 |
|
|
|
| 7 |
B1 |
640 |
614 |
21.81 |
|
|
|
| 8 |
B1 |
218 |
209 |
0.37 |
|
|
|
| 9 |
B2 |
3208 |
3078 |
9.72 |
|
|
|
| 10 |
B2 |
1067 |
1023 |
2.20 |
|
|
|
| 11 |
B2 |
445 |
427 |
12.58 |
|
|
|
| 12 |
B2 |
151i |
145i |
17.65 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 8017.2 cm
-1
Scaled (by 0.9594) Zero Point Vibrational Energy (zpe) 7691.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD/cc-pVDZ
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
-1.897 |
| C2 |
0.000 |
0.000 |
-0.564 |
| C3 |
0.000 |
0.000 |
0.732 |
| O4 |
0.000 |
0.000 |
1.917 |
| H5 |
0.000 |
0.935 |
-2.477 |
| H6 |
0.000 |
-0.935 |
-2.477 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
H5 |
H6 |
| C1 | | 1.3338 | 2.6289 | 3.8140 | 1.1005 | 1.1005 |
C2 | 1.3338 | | 1.2952 | 2.4802 | 2.1301 | 2.1301 | C3 | 2.6289 | 1.2952 | | 1.1850 | 3.3425 | 3.3425 | O4 | 3.8140 | 2.4802 | 1.1850 | | 4.4924 | 4.4924 | H5 | 1.1005 | 2.1301 | 3.3425 | 4.4924 | | 1.8705 | H6 | 1.1005 | 2.1301 | 3.3425 | 4.4924 | 1.8705 | |
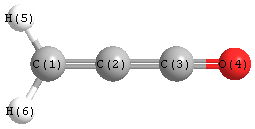 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
180.000 |
|
C2 |
C1 |
H5 |
121.807 |
| C2 |
C1 |
H6 |
121.807 |
|
C2 |
C3 |
O4 |
180.000 |
| H5 |
C1 |
H6 |
116.386 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at QCISD/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -190.136430 |
| Energy at 298.15K | |
| HF Energy | -189.549795 |
| Nuclear repulsion energy | 87.744231 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3222 |
3091 |
8.19 |
|
|
|
| 2 |
A' |
3133 |
3005 |
23.60 |
|
|
|
| 3 |
A' |
2171 |
2082 |
829.07 |
|
|
|
| 4 |
A' |
1740 |
1669 |
8.08 |
|
|
|
| 5 |
A' |
1482 |
1422 |
2.23 |
|
|
|
| 6 |
A' |
1071 |
1027 |
30.41 |
|
|
|
| 7 |
A' |
943 |
905 |
1.62 |
|
|
|
| 8 |
A' |
504 |
484 |
12.28 |
|
|
|
| 9 |
A' |
183 |
175 |
24.07 |
|
|
|
| 10 |
A" |
999 |
958 |
20.86 |
|
|
|
| 11 |
A" |
714 |
685 |
12.83 |
|
|
|
| 12 |
A" |
273 |
262 |
3.33 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 8215.7 cm
-1
Scaled (by 0.9594) Zero Point Vibrational Energy (zpe) 7882.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.165 |
-1.815 |
0.000 |
| C2 |
-0.379 |
-0.584 |
0.000 |
| C3 |
0.000 |
0.694 |
0.000 |
| O4 |
0.064 |
1.866 |
0.000 |
| H5 |
1.251 |
-1.994 |
0.000 |
| H6 |
-0.475 |
-2.708 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
H5 |
H6 |
| C1 | | 1.3455 | 2.5140 | 3.6827 | 1.1007 | 1.0988 |
C2 | 1.3455 | | 1.3327 | 2.4900 | 2.1549 | 2.1262 | C3 | 2.5140 | 1.3327 | | 1.1744 | 2.9646 | 3.4348 | O4 | 3.6827 | 2.4900 | 1.1744 | | 4.0389 | 4.6060 | H5 | 1.1007 | 2.1549 | 2.9646 | 4.0389 | | 1.8679 | H6 | 1.0988 | 2.1262 | 3.4348 | 4.6060 | 1.8679 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
139.667 |
|
C2 |
C1 |
H5 |
123.191 |
| C2 |
C1 |
H6 |
120.546 |
|
C2 |
C3 |
O4 |
166.594 |
| H5 |
C1 |
H6 |
116.263 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
