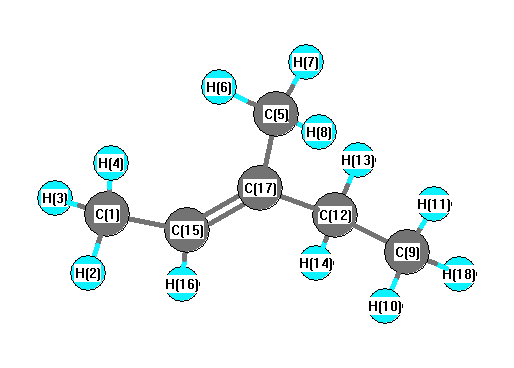
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3072 |
3054 |
26.22 |
|
|
|
| 2 |
A |
3067 |
3049 |
34.60 |
|
|
|
| 3 |
A |
3053 |
3035 |
29.67 |
|
|
|
| 4 |
A |
3051 |
3033 |
32.15 |
|
|
|
| 5 |
A |
3048 |
3030 |
2.62 |
|
|
|
| 6 |
A |
3011 |
2994 |
18.93 |
|
|
|
| 7 |
A |
3006 |
2989 |
17.82 |
|
|
|
| 8 |
A |
3000 |
2982 |
14.59 |
|
|
|
| 9 |
A |
2970 |
2952 |
30.50 |
|
|
|
| 10 |
A |
2957 |
2940 |
20.08 |
|
|
|
| 11 |
A |
2953 |
2936 |
49.70 |
|
|
|
| 12 |
A |
2936 |
2919 |
35.92 |
|
|
|
| 13 |
A |
1695 |
1685 |
1.80 |
|
|
|
| 14 |
A |
1430 |
1422 |
3.88 |
|
|
|
| 15 |
A |
1421 |
1413 |
7.59 |
|
|
|
| 16 |
A |
1414 |
1406 |
3.55 |
|
|
|
| 17 |
A |
1408 |
1400 |
5.42 |
|
|
|
| 18 |
A |
1404 |
1396 |
10.78 |
|
|
|
| 19 |
A |
1401 |
1392 |
2.44 |
|
|
|
| 20 |
A |
1392 |
1384 |
0.75 |
|
|
|
| 21 |
A |
1358 |
1350 |
3.00 |
|
|
|
| 22 |
A |
1345 |
1337 |
2.76 |
|
|
|
| 23 |
A |
1337 |
1329 |
0.70 |
|
|
|
| 24 |
A |
1304 |
1296 |
2.91 |
|
|
|
| 25 |
A |
1296 |
1288 |
2.52 |
|
|
|
| 26 |
A |
1231 |
1224 |
0.34 |
|
|
|
| 27 |
A |
1196 |
1189 |
5.24 |
|
|
|
| 28 |
A |
1107 |
1100 |
9.23 |
|
|
|
| 29 |
A |
1056 |
1050 |
0.89 |
|
|
|
| 30 |
A |
1050 |
1043 |
6.76 |
|
|
|
| 31 |
A |
1005 |
999 |
0.28 |
|
|
|
| 32 |
A |
999 |
994 |
1.75 |
|
|
|
| 33 |
A |
979 |
973 |
5.51 |
|
|
|
| 34 |
A |
960 |
954 |
9.18 |
|
|
|
| 35 |
A |
904 |
899 |
0.97 |
|
|
|
| 36 |
A |
818 |
813 |
10.45 |
|
|
|
| 37 |
A |
765 |
760 |
2.23 |
|
|
|
| 38 |
A |
744 |
739 |
1.52 |
|
|
|
| 39 |
A |
535 |
532 |
4.64 |
|
|
|
| 40 |
A |
480 |
477 |
2.98 |
|
|
|
| 41 |
A |
386 |
384 |
0.53 |
|
|
|
| 42 |
A |
310 |
308 |
0.60 |
|
|
|
| 43 |
A |
296 |
294 |
0.25 |
|
|
|
| 44 |
A |
223 |
222 |
0.11 |
|
|
|
| 45 |
A |
187 |
186 |
0.92 |
|
|
|
| 46 |
A |
141 |
140 |
0.03 |
|
|
|
| 47 |
A |
113 |
113 |
1.22 |
|
|
|
| 48 |
A |
53 |
53 |
0.02 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 34930.8 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 34728.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.014 |
|
|
|
| 2 |
H |
0.045 |
|
|
|
| 3 |
H |
0.044 |
|
|
|
| 4 |
H |
0.026 |
|
|
|
| 5 |
C |
-0.001 |
|
|
|
| 6 |
H |
0.027 |
|
|
|
| 7 |
H |
0.039 |
|
|
|
| 8 |
H |
0.045 |
|
|
|
| 9 |
C |
-0.059 |
|
|
|
| 10 |
H |
0.033 |
|
|
|
| 11 |
H |
0.032 |
|
|
|
| 12 |
C |
-0.009 |
|
|
|
| 13 |
H |
0.023 |
|
|
|
| 14 |
H |
0.009 |
|
|
|
| 15 |
C |
-0.088 |
|
|
|
| 16 |
H |
-0.027 |
|
|
|
| 17 |
C |
-0.153 |
|
|
|
| 18 |
H |
0.028 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.009 |
0.193 |
0.035 |
0.197 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-38.821 |
0.278 |
-0.640 |
| y |
0.278 |
-39.046 |
0.059 |
| z |
-0.640 |
0.059 |
-41.051 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.227 |
0.278 |
-0.640 |
| y |
0.278 |
0.890 |
0.059 |
| z |
-0.640 |
0.059 |
-2.118 |
|
| Polar |
|---|
| 3z2-r2 | -4.235 |
|---|
| x2-y2 | 0.225 |
|---|
| xy | 0.278 |
|---|
| xz | -0.640 |
|---|
| yz | 0.059 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
13.576 |
0.997 |
-0.484 |
| y |
0.997 |
10.512 |
0.030 |
| z |
-0.484 |
0.030 |
8.015 |
<r2> (average value of r
2) Å
2
| <r2> |
225.606 |
| (<r2>)1/2 |
15.020 |