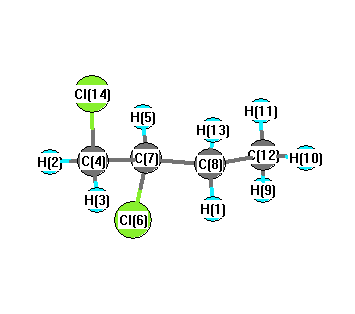
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3098 |
3080 |
2.78 |
|
|
|
| 2 |
A |
3069 |
3051 |
13.47 |
|
|
|
| 3 |
A |
3057 |
3039 |
24.77 |
|
|
|
| 4 |
A |
3026 |
3009 |
15.69 |
|
|
|
| 5 |
A |
3020 |
3003 |
1.02 |
|
|
|
| 6 |
A |
3009 |
2991 |
1.06 |
|
|
|
| 7 |
A |
2976 |
2959 |
23.87 |
|
|
|
| 8 |
A |
2968 |
2951 |
10.92 |
|
|
|
| 9 |
A |
1430 |
1422 |
6.89 |
|
|
|
| 10 |
A |
1418 |
1410 |
7.45 |
|
|
|
| 11 |
A |
1404 |
1396 |
3.95 |
|
|
|
| 12 |
A |
1396 |
1388 |
3.64 |
|
|
|
| 13 |
A |
1350 |
1342 |
1.90 |
|
|
|
| 14 |
A |
1342 |
1334 |
0.85 |
|
|
|
| 15 |
A |
1275 |
1268 |
4.37 |
|
|
|
| 16 |
A |
1262 |
1254 |
2.56 |
|
|
|
| 17 |
A |
1226 |
1218 |
6.51 |
|
|
|
| 18 |
A |
1184 |
1177 |
4.26 |
|
|
|
| 19 |
A |
1142 |
1135 |
16.29 |
|
|
|
| 20 |
A |
1101 |
1095 |
2.05 |
|
|
|
| 21 |
A |
1050 |
1044 |
1.57 |
|
|
|
| 22 |
A |
1032 |
1026 |
0.96 |
|
|
|
| 23 |
A |
1019 |
1013 |
2.74 |
|
|
|
| 24 |
A |
914 |
909 |
10.43 |
|
|
|
| 25 |
A |
810 |
805 |
5.16 |
|
|
|
| 26 |
A |
787 |
783 |
11.77 |
|
|
|
| 27 |
A |
712 |
707 |
49.89 |
|
|
|
| 28 |
A |
639 |
635 |
41.40 |
|
|
|
| 29 |
A |
450 |
448 |
1.67 |
|
|
|
| 30 |
A |
374 |
372 |
1.89 |
|
|
|
| 31 |
A |
286 |
284 |
0.09 |
|
|
|
| 32 |
A |
238 |
237 |
0.41 |
|
|
|
| 33 |
A |
210 |
209 |
3.16 |
|
|
|
| 34 |
A |
184 |
183 |
3.69 |
|
|
|
| 35 |
A |
111 |
111 |
3.08 |
|
|
|
| 36 |
A |
96 |
95 |
1.25 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 24333.1 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 24192.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
H |
0.034 |
|
|
|
| 2 |
H |
0.089 |
|
|
|
| 3 |
H |
0.086 |
|
|
|
| 4 |
C |
-0.015 |
|
|
|
| 5 |
H |
0.072 |
|
|
|
| 6 |
Cl |
-0.105 |
|
|
|
| 7 |
C |
-0.154 |
|
|
|
| 8 |
C |
-0.008 |
|
|
|
| 9 |
H |
0.047 |
|
|
|
| 10 |
H |
0.036 |
|
|
|
| 11 |
H |
0.036 |
|
|
|
| 12 |
C |
-0.047 |
|
|
|
| 13 |
H |
0.046 |
|
|
|
| 14 |
Cl |
-0.116 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.272 |
0.585 |
0.213 |
0.680 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-56.704 |
2.982 |
-0.185 |
| y |
2.982 |
-50.680 |
-0.186 |
| z |
-0.185 |
-0.186 |
-50.465 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-6.132 |
2.982 |
-0.185 |
| y |
2.982 |
2.905 |
-0.186 |
| z |
-0.185 |
-0.186 |
3.227 |
|
| Polar |
|---|
| 3z2-r2 | 6.454 |
|---|
| x2-y2 | -6.024 |
|---|
| xy | 2.982 |
|---|
| xz | -0.185 |
|---|
| yz | -0.186 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
12.079 |
-0.910 |
0.162 |
| y |
-0.910 |
10.214 |
-0.167 |
| z |
0.162 |
-0.167 |
7.598 |
<r2> (average value of r
2) Å
2
| <r2> |
326.475 |
| (<r2>)1/2 |
18.069 |