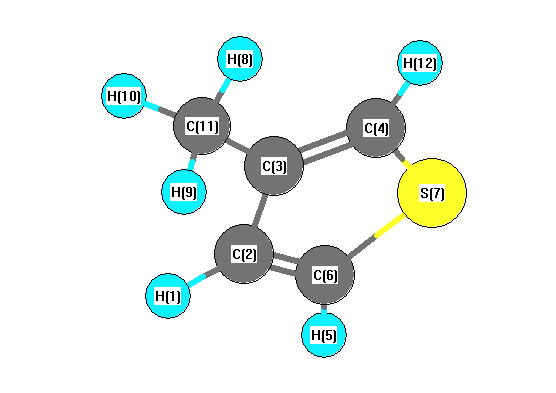
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3187 |
3168 |
0.57 |
|
|
|
| 2 |
A' |
3178 |
3159 |
0.71 |
|
|
|
| 3 |
A' |
3137 |
3119 |
4.41 |
|
|
|
| 4 |
A' |
3068 |
3050 |
11.92 |
|
|
|
| 5 |
A' |
2966 |
2949 |
33.22 |
|
|
|
| 6 |
A' |
1555 |
1546 |
3.54 |
|
|
|
| 7 |
A' |
1446 |
1437 |
8.81 |
|
|
|
| 8 |
A' |
1411 |
1402 |
2.21 |
|
|
|
| 9 |
A' |
1375 |
1367 |
5.84 |
|
|
|
| 10 |
A' |
1340 |
1333 |
0.64 |
|
|
|
| 11 |
A' |
1198 |
1191 |
2.71 |
|
|
|
| 12 |
A' |
1132 |
1125 |
3.18 |
|
|
|
| 13 |
A' |
1051 |
1045 |
3.86 |
|
|
|
| 14 |
A' |
968 |
962 |
10.09 |
|
|
|
| 15 |
A' |
920 |
915 |
2.19 |
|
|
|
| 16 |
A' |
846 |
841 |
13.65 |
|
|
|
| 17 |
A' |
815 |
811 |
4.46 |
|
|
|
| 18 |
A' |
649 |
645 |
0.37 |
|
|
|
| 19 |
A' |
533 |
529 |
0.29 |
|
|
|
| 20 |
A' |
313 |
312 |
0.88 |
|
|
|
| 21 |
A" |
3030 |
3013 |
13.73 |
|
|
|
| 22 |
A" |
1403 |
1395 |
6.14 |
|
|
|
| 23 |
A" |
1005 |
999 |
3.70 |
|
|
|
| 24 |
A" |
845 |
840 |
0.12 |
|
|
|
| 25 |
A" |
736 |
731 |
59.34 |
|
|
|
| 26 |
A" |
646 |
642 |
1.39 |
|
|
|
| 27 |
A" |
588 |
584 |
10.32 |
|
|
|
| 28 |
A" |
448 |
445 |
0.19 |
|
|
|
| 29 |
A" |
228 |
227 |
3.07 |
|
|
|
| 30 |
A" |
110 |
110 |
0.26 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 20063.1 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 19946.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
H |
-0.027 |
|
|
|
| 2 |
C |
0.045 |
|
|
|
| 3 |
C |
-0.046 |
|
|
|
| 4 |
C |
-0.157 |
|
|
|
| 5 |
H |
0.021 |
|
|
|
| 6 |
C |
-0.125 |
|
|
|
| 7 |
S |
0.141 |
|
|
|
| 8 |
H |
0.031 |
|
|
|
| 9 |
H |
0.051 |
|
|
|
| 10 |
H |
0.051 |
|
|
|
| 11 |
C |
0.008 |
|
|
|
| 12 |
H |
0.009 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.255 |
0.780 |
0.000 |
0.820 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-38.910 |
1.265 |
0.000 |
| y |
1.265 |
-40.157 |
0.000 |
| z |
0.000 |
0.000 |
-45.999 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
4.168 |
1.265 |
0.000 |
| y |
1.265 |
2.297 |
0.000 |
| z |
0.000 |
0.000 |
-6.465 |
|
| Polar |
|---|
| 3z2-r2 | -12.930 |
|---|
| x2-y2 | 1.247 |
|---|
| xy | 1.265 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
11.033 |
-0.287 |
0.000 |
| y |
-0.287 |
13.320 |
0.000 |
| z |
0.000 |
0.000 |
5.078 |
<r2> (average value of r
2) Å
2
| <r2> |
189.877 |
| (<r2>)1/2 |
13.780 |