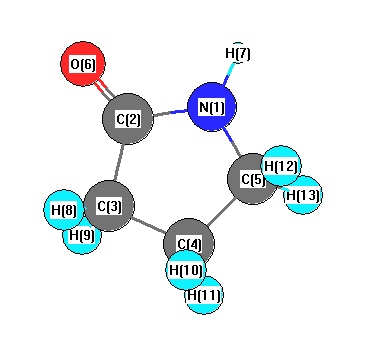
Vibrational Frequencies calculated at BLYP/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3492 |
3498 |
15.13 |
|
|
|
| 2 |
A |
3036 |
3041 |
21.02 |
|
|
|
| 3 |
A |
3025 |
3030 |
23.65 |
|
|
|
| 4 |
A |
2983 |
2988 |
57.51 |
|
|
|
| 5 |
A |
2970 |
2975 |
12.28 |
|
|
|
| 6 |
A |
2960 |
2965 |
17.55 |
|
|
|
| 7 |
A |
2891 |
2895 |
78.64 |
|
|
|
| 8 |
A |
1746 |
1749 |
357.14 |
|
|
|
| 9 |
A |
1465 |
1467 |
4.05 |
|
|
|
| 10 |
A |
1429 |
1431 |
5.60 |
|
|
|
| 11 |
A |
1400 |
1402 |
2.78 |
|
|
|
| 12 |
A |
1383 |
1385 |
26.70 |
|
|
|
| 13 |
A |
1312 |
1314 |
8.27 |
|
|
|
| 14 |
A |
1284 |
1286 |
12.19 |
|
|
|
| 15 |
A |
1244 |
1246 |
15.69 |
|
|
|
| 16 |
A |
1208 |
1210 |
53.89 |
|
|
|
| 17 |
A |
1189 |
1191 |
38.55 |
|
|
|
| 18 |
A |
1156 |
1158 |
1.28 |
|
|
|
| 19 |
A |
1137 |
1139 |
3.05 |
|
|
|
| 20 |
A |
1049 |
1050 |
0.54 |
|
|
|
| 21 |
A |
1034 |
1036 |
14.56 |
|
|
|
| 22 |
A |
968 |
969 |
10.36 |
|
|
|
| 23 |
A |
890 |
891 |
0.78 |
|
|
|
| 24 |
A |
875 |
877 |
1.56 |
|
|
|
| 25 |
A |
855 |
856 |
4.22 |
|
|
|
| 26 |
A |
780 |
781 |
5.48 |
|
|
|
| 27 |
A |
667 |
668 |
6.19 |
|
|
|
| 28 |
A |
610 |
611 |
13.95 |
|
|
|
| 29 |
A |
544 |
545 |
30.11 |
|
|
|
| 30 |
A |
475 |
476 |
55.06 |
|
|
|
| 31 |
A |
447 |
448 |
9.51 |
|
|
|
| 32 |
A |
194 |
195 |
3.51 |
|
|
|
| 33 |
A |
136 |
136 |
1.24 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 23417.4 cm
-1
Scaled (by 1.0016) Zero Point Vibrational Energy (zpe) 23454.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at BLYP/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.087 |
|
|
|
| 2 |
C |
0.049 |
|
|
|
| 3 |
C |
0.020 |
|
|
|
| 4 |
C |
0.012 |
|
|
|
| 5 |
C |
0.105 |
|
|
|
| 6 |
O |
-0.223 |
|
|
|
| 7 |
H |
0.068 |
|
|
|
| 8 |
H |
0.024 |
|
|
|
| 9 |
H |
0.015 |
|
|
|
| 10 |
H |
-0.005 |
|
|
|
| 11 |
H |
0.011 |
|
|
|
| 12 |
H |
0.006 |
|
|
|
| 13 |
H |
0.005 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-3.522 |
-0.735 |
0.318 |
3.612 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-43.142 |
-0.235 |
0.200 |
| y |
-0.235 |
-31.883 |
-0.185 |
| z |
0.200 |
-0.185 |
-35.635 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-9.383 |
-0.235 |
0.200 |
| y |
-0.235 |
7.506 |
-0.185 |
| z |
0.200 |
-0.185 |
1.877 |
|
| Polar |
|---|
| 3z2-r2 | 3.754 |
|---|
| x2-y2 | -11.260 |
|---|
| xy | -0.235 |
|---|
| xz | 0.200 |
|---|
| yz | -0.185 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.152 |
0.428 |
-0.115 |
| y |
0.428 |
8.229 |
0.023 |
| z |
-0.115 |
0.023 |
5.896 |
<r2> (average value of r
2) Å
2
| <r2> |
0.000 |
| (<r2>)1/2 |
0.000 |