Jump to
S1C2
Energy calculated at PBEPBEultrafine/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -277.993939 |
| Energy at 298.15K | |
| HF Energy | -277.993939 |
| Nuclear repulsion energy | 124.568594 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBEPBEultrafine/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
2960 |
2943 |
0.00 |
255.68 |
0.08 |
0.14 |
| 2 |
Ag |
1435 |
1427 |
0.00 |
23.55 |
0.70 |
0.82 |
| 3 |
Ag |
1387 |
1379 |
0.00 |
4.28 |
0.54 |
0.70 |
| 4 |
Ag |
1061 |
1055 |
0.00 |
6.70 |
0.47 |
0.64 |
| 5 |
Ag |
1039 |
1033 |
0.00 |
7.50 |
0.63 |
0.77 |
| 6 |
Ag |
443 |
441 |
0.00 |
3.28 |
0.55 |
0.71 |
| 7 |
Au |
3031 |
3014 |
68.93 |
0.00 |
0.00 |
0.00 |
| 8 |
Au |
1169 |
1162 |
2.00 |
0.00 |
0.00 |
0.00 |
| 9 |
Au |
789 |
784 |
0.03 |
0.00 |
0.00 |
0.00 |
| 10 |
Au |
111 |
110 |
10.14 |
0.00 |
0.00 |
0.00 |
| 11 |
Bg |
3005 |
2987 |
0.00 |
159.33 |
0.75 |
0.86 |
| 12 |
Bg |
1241 |
1234 |
0.00 |
14.92 |
0.75 |
0.86 |
| 13 |
Bg |
1125 |
1118 |
0.00 |
5.95 |
0.75 |
0.86 |
| 14 |
Bu |
2965 |
2948 |
71.16 |
0.00 |
0.00 |
0.00 |
| 15 |
Bu |
1446 |
1438 |
1.05 |
0.00 |
0.00 |
0.00 |
| 16 |
Bu |
1294 |
1287 |
12.82 |
0.00 |
0.00 |
0.00 |
| 17 |
Bu |
1056 |
1049 |
183.26 |
0.00 |
0.00 |
0.00 |
| 18 |
Bu |
258 |
257 |
15.28 |
0.00 |
0.00 |
0.00 |
Unscaled Zero Point Vibrational Energy (zpe) 12907.1 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 12832.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBEPBEultrafine/cc-pVDZ
Point Group is C2h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.425 |
0.632 |
0.000 |
| C2 |
-0.425 |
-0.632 |
0.000 |
| F3 |
-0.425 |
1.740 |
0.000 |
| F4 |
0.425 |
-1.740 |
0.000 |
| H5 |
1.071 |
0.665 |
0.904 |
| H6 |
1.071 |
0.665 |
-0.904 |
| H7 |
-1.071 |
-0.665 |
0.904 |
| H8 |
-1.071 |
-0.665 |
-0.904 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
F3 |
F4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5230 | 1.3962 | 2.3724 | 1.1120 | 1.1120 | 2.1768 | 2.1768 |
C2 | 1.5230 | | 2.3724 | 1.3962 | 2.1768 | 2.1768 | 1.1120 | 1.1120 | F3 | 1.3962 | 2.3724 | | 3.5827 | 2.0521 | 2.0521 | 2.6499 | 2.6499 | F4 | 2.3724 | 1.3962 | 3.5827 | | 2.6499 | 2.6499 | 2.0521 | 2.0521 | H5 | 1.1120 | 2.1768 | 2.0521 | 2.6499 | | 1.8079 | 2.5223 | 3.1033 | H6 | 1.1120 | 2.1768 | 2.0521 | 2.6499 | 1.8079 | | 3.1033 | 2.5223 | H7 | 2.1768 | 1.1120 | 2.6499 | 2.0521 | 2.5223 | 3.1033 | | 1.8079 | H8 | 2.1768 | 1.1120 | 2.6499 | 2.0521 | 3.1033 | 2.5223 | 1.8079 | |
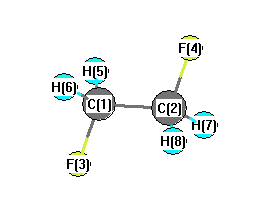 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
F4 |
108.639 |
|
C1 |
C2 |
H7 |
110.430 |
| C1 |
C2 |
H8 |
110.430 |
|
C2 |
C1 |
F3 |
108.639 |
| C2 |
C1 |
H5 |
110.430 |
|
C2 |
C1 |
H6 |
110.430 |
| F3 |
C1 |
H5 |
109.278 |
|
F3 |
C1 |
H6 |
109.278 |
| F4 |
C2 |
H7 |
109.278 |
|
F4 |
C2 |
H8 |
109.278 |
| H5 |
C1 |
H6 |
108.764 |
|
H7 |
C2 |
H8 |
108.764 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.143 |
|
|
|
| 2 |
C |
0.143 |
|
|
|
| 3 |
F |
-0.216 |
|
|
|
| 4 |
F |
-0.216 |
|
|
|
| 5 |
H |
0.036 |
|
|
|
| 6 |
H |
0.036 |
|
|
|
| 7 |
H |
0.036 |
|
|
|
| 8 |
H |
0.036 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-21.869 |
2.295 |
0.000 |
| y |
2.295 |
-28.016 |
0.000 |
| z |
0.000 |
0.000 |
-21.590 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.934 |
2.295 |
0.000 |
| y |
2.295 |
-6.286 |
0.000 |
| z |
0.000 |
0.000 |
3.352 |
|
| Polar |
|---|
| 3z2-r2 | 6.705 |
|---|
| x2-y2 | 6.147 |
|---|
| xy | 2.295 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.788 |
0.070 |
0.000 |
| y |
0.070 |
3.737 |
0.000 |
| z |
0.000 |
0.000 |
3.528 |
<r2> (average value of r
2) Å
2
| <r2> |
89.231 |
| (<r2>)1/2 |
9.446 |
Jump to
S1C1
Energy calculated at PBEPBEultrafine/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -277.995238 |
| Energy at 298.15K | |
| HF Energy | -277.995238 |
| Nuclear repulsion energy | 126.537106 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBEPBEultrafine/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
2999 |
2981 |
21.95 |
125.84 |
0.73 |
0.85 |
| 2 |
A |
2953 |
2936 |
41.89 |
219.99 |
0.02 |
0.04 |
| 3 |
A |
1411 |
1403 |
1.22 |
2.00 |
0.55 |
0.71 |
| 4 |
A |
1376 |
1368 |
17.39 |
5.24 |
0.60 |
0.75 |
| 5 |
A |
1249 |
1242 |
1.26 |
20.04 |
0.72 |
0.83 |
| 6 |
A |
1089 |
1083 |
78.23 |
3.66 |
0.75 |
0.86 |
| 7 |
A |
1082 |
1076 |
2.31 |
0.99 |
0.20 |
0.33 |
| 8 |
A |
849 |
844 |
23.24 |
4.96 |
0.33 |
0.50 |
| 9 |
A |
311 |
309 |
0.32 |
0.43 |
0.49 |
0.65 |
| 10 |
A |
146 |
145 |
2.77 |
0.07 |
0.75 |
0.86 |
| 11 |
B |
3013 |
2995 |
51.11 |
15.53 |
0.75 |
0.86 |
| 12 |
B |
2940 |
2923 |
15.43 |
45.98 |
0.75 |
0.86 |
| 13 |
B |
1405 |
1397 |
7.88 |
17.43 |
0.75 |
0.86 |
| 14 |
B |
1338 |
1331 |
11.10 |
1.20 |
0.75 |
0.86 |
| 15 |
B |
1204 |
1197 |
5.10 |
5.43 |
0.75 |
0.86 |
| 16 |
B |
1054 |
1048 |
50.32 |
3.49 |
0.75 |
0.86 |
| 17 |
B |
873 |
868 |
34.31 |
3.55 |
0.75 |
0.86 |
| 18 |
B |
471 |
468 |
12.88 |
0.70 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 12880.5 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 12805.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBEPBEultrafine/cc-pVDZ
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.268 |
0.706 |
0.508 |
| C2 |
-0.268 |
-0.706 |
0.508 |
| F3 |
-0.268 |
1.446 |
-0.547 |
| F4 |
0.268 |
-1.446 |
-0.547 |
| H5 |
-0.003 |
1.201 |
1.468 |
| H6 |
1.376 |
0.693 |
0.407 |
| H7 |
0.003 |
-1.201 |
1.468 |
| H8 |
-1.376 |
-0.693 |
0.407 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
F3 |
F4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5101 | 1.3948 | 2.3964 | 1.1143 | 1.1127 | 2.1521 | 2.1607 |
C2 | 1.5101 | | 2.3964 | 1.3948 | 2.1521 | 2.1607 | 1.1143 | 1.1127 | F3 | 1.3948 | 2.3964 | | 2.9410 | 2.0471 | 2.0437 | 3.3381 | 2.5914 | F4 | 2.3964 | 1.3948 | 2.9410 | | 3.3381 | 2.5914 | 2.0471 | 2.0437 | H5 | 1.1143 | 2.1521 | 2.0471 | 3.3381 | | 1.8124 | 2.4029 | 2.5691 | H6 | 1.1127 | 2.1607 | 2.0437 | 2.5914 | 1.8124 | | 2.5691 | 3.0811 | H7 | 2.1521 | 1.1143 | 3.3381 | 2.0471 | 2.4029 | 2.5691 | | 1.8124 | H8 | 2.1607 | 1.1127 | 2.5914 | 2.0437 | 2.5691 | 3.0811 | 1.8124 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
F4 |
111.105 |
|
C1 |
C2 |
H7 |
109.247 |
| C1 |
C2 |
H8 |
110.014 |
|
C2 |
C1 |
F3 |
111.105 |
| C2 |
C1 |
H5 |
109.247 |
|
C2 |
C1 |
H6 |
110.014 |
| F3 |
C1 |
H5 |
108.829 |
|
F3 |
C1 |
H6 |
108.661 |
| F4 |
C2 |
H7 |
108.829 |
|
F4 |
C2 |
H8 |
108.661 |
| H5 |
C1 |
H6 |
108.944 |
|
H7 |
C2 |
H8 |
108.944 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.152 |
|
|
|
| 2 |
C |
0.152 |
|
|
|
| 3 |
F |
-0.214 |
|
|
|
| 4 |
F |
-0.214 |
|
|
|
| 5 |
H |
0.024 |
|
|
|
| 6 |
H |
0.038 |
|
|
|
| 7 |
H |
0.024 |
|
|
|
| 8 |
H |
0.038 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
2.272 |
2.272 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-21.648 |
1.314 |
0.000 |
| y |
1.314 |
-25.646 |
0.000 |
| z |
0.000 |
0.000 |
-22.069 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.209 |
1.314 |
0.000 |
| y |
1.314 |
-3.788 |
0.000 |
| z |
0.000 |
0.000 |
1.579 |
|
| Polar |
|---|
| 3z2-r2 | 3.157 |
|---|
| x2-y2 | 3.998 |
|---|
| xy | 1.314 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.651 |
0.097 |
0.000 |
| y |
0.097 |
3.877 |
0.000 |
| z |
0.000 |
0.000 |
3.705 |
<r2> (average value of r
2) Å
2
| <r2> |
80.956 |
| (<r2>)1/2 |
8.998 |
