return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C8H6 (Calicene)

using model chemistry: QCISD/cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at QCISD/cc-pVDZ
 hartrees
Energy at 0K-307.388648
Energy at 298.15K-307.393335
HF Energy-306.312526
Nuclear repulsion energy296.890699
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A1 3320 3185 13.41      
2 A1 3260 3128 14.63      
3 A1 3235 3104 4.60      
4 A1 1831 1757 645.80      
5 A1 1585 1521 74.49      
6 A1 1534 1472 117.79      
7 A1 1420 1362 85.87      
8 A1 1219 1170 21.06      
9 A1 1074 1030 8.38      
10 A1 1018 977 18.49      
11 A1 949 910 8.55      
12 A1 878 843 51.89      
13 A1 485 465 1.27      
14 A2 892 856 0.00      
15 A2 887 851 0.00      
16 A2 714 685 0.00      
17 A2 568 545 0.00      
18 A2 217 208 0.00      
19 B1 896 859 0.00      
20 B1 751 720 77.04      
21 B1 692 664 40.07      
22 B1 595 571 5.74      
23 B1 415 398 8.65      
24 B1 127 122 5.58      
25 B2 3281 3148 5.63      
26 B2 3251 3119 24.97      
27 B2 3226 3095 1.35      
28 B2 1614 1549 0.26      
29 B2 1364 1309 0.32      
30 B2 1230 1180 1.29      
31 B2 1120 1075 13.23      
32 B2 1095 1050 5.84      
33 B2 940 902 6.20      
34 B2 827 794 1.70      
35 B2 491 471 1.24      
36 B2 145 139 1.44      

Unscaled Zero Point Vibrational Energy (zpe) 23572.9 cm-1
Scaled (by 0.9594) Zero Point Vibrational Energy (zpe) 22615.8 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/cc-pVDZ
ABC
0.22135 0.04658 0.03849

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/cc-pVDZ

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.000 -0.045
C2 0.000 0.000 1.313
C3 0.000 0.671 2.597
C4 0.000 -0.671 2.597
C5 0.000 1.181 -0.922
C6 0.000 -1.181 -0.922
C7 0.000 0.737 -2.221
C8 0.000 -0.737 -2.221
H9 0.000 1.601 3.168
H10 0.000 -1.601 3.168
H11 0.000 2.218 -0.576
H12 0.000 -2.218 -0.576
H13 0.000 1.364 -3.118
H14 0.000 -1.364 -3.118

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 C7 C8 H9 H10 H11 H12 H13 H14
C11.35862.72652.72651.47101.47102.29692.29693.58963.58962.28092.28093.36123.3612
C21.35861.44891.44892.52832.52833.61003.61002.44962.44962.91392.91394.63584.6358
C32.72651.44891.34233.55633.97724.81855.01961.09042.34223.53064.29185.75656.0662
C42.72651.44891.34233.97723.55635.01964.81852.34221.09044.29183.53066.06625.7565
C51.47102.52833.55633.97722.36221.37242.31634.11144.94621.09343.41692.20273.3608
C61.47102.52833.97723.55632.36222.31631.37244.94624.11143.41691.09343.36082.2027
C72.29693.61004.81855.01961.37242.31631.47415.45725.87362.21333.38211.09402.2842
C82.29693.61005.01964.81852.31631.37241.47415.87365.45723.38212.21332.28421.0940
H93.58962.44961.09042.34224.11144.94625.45725.87363.20103.79455.34796.28956.9490
H103.58962.44962.34221.09044.94624.11145.87365.45723.20105.34793.79456.94906.2895
H112.28092.91393.53064.29181.09343.41692.21333.38213.79455.34794.43662.68104.3919
H122.28092.91394.29183.53063.41691.09343.38212.21335.34793.79454.43664.39192.6810
H133.36124.63585.75656.06622.20273.36081.09402.28426.28956.94902.68104.39192.7276
H143.36124.63586.06625.75653.36082.20272.28421.09406.94906.28954.39192.68102.7276

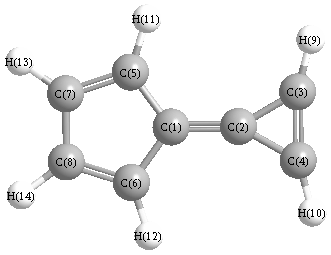
picture of Calicene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 152.405 C1 C2 C4 152.405
C1 C5 C7 107.714 C1 C5 H11 124.958
C1 C6 C8 107.714 C1 C6 H12 124.958
C2 C1 C5 126.593 C2 C1 C6 126.593
C2 C3 C4 62.405 C2 C3 H9 149.135
C2 C4 C3 62.405 C2 C4 H10 149.135
C3 C2 C4 55.190 C3 C4 H10 148.460
C4 C3 H9 148.460 C5 C1 C6 106.814
C5 C7 C8 108.879 C5 C7 H13 126.165
C6 C8 C7 108.879 C6 C8 H14 126.165
C7 C5 H11 127.328 C7 C8 H14 124.956
C8 C6 H12 127.328 C8 C7 H13 124.956
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability