return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C8H6 (Calicene)

using model chemistry: CCD/cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at CCD/cc-pVDZ
 hartrees
Energy at 0K-307.378326
Energy at 298.15K-307.383051
HF Energy-306.313189
Nuclear repulsion energy297.225444
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A1 3329 3185 14.99      
2 A1 3269 3127 14.06      
3 A1 3244 3103 4.10      
4 A1 1855 1775 694.88      
5 A1 1605 1535 85.90      
6 A1 1547 1480 123.14      
7 A1 1426 1364 90.60      
8 A1 1227 1174 25.06      
9 A1 1079 1032 7.86      
10 A1 1023 979 21.71      
11 A1 952 911 8.82      
12 A1 882 844 55.99      
13 A1 488 467 1.74      
14 A2 899 860 0.00      
15 A2 896 857 0.00      
16 A2 716 685 0.00      
17 A2 571 546 0.00      
18 A2 220 210 0.00      
19 B1 901 862 0.01      
20 B1 753 721 78.63      
21 B1 697 666 40.01      
22 B1 598 572 5.29      
23 B1 419 401 9.12      
24 B1 127 122 5.62      
25 B2 3290 3147 6.72      
26 B2 3260 3119 23.65      
27 B2 3234 3094 1.15      
28 B2 1630 1559 0.69      
29 B2 1373 1313 0.09      
30 B2 1236 1183 1.12      
31 B2 1132 1083 10.95      
32 B2 1099 1051 6.51      
33 B2 951 910 7.52      
34 B2 830 794 1.68      
35 B2 495 474 1.43      
36 B2 146 139 1.42      

Unscaled Zero Point Vibrational Energy (zpe) 23699.0 cm-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 22672.8 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at CCD/cc-pVDZ
ABC
0.22172 0.04671 0.03858

See section I.F.4 to change rotational constant units
Geometric Data calculated at CCD/cc-pVDZ

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.000 -0.044
C2 0.000 0.000 1.311
C3 0.000 0.670 2.594
C4 0.000 -0.670 2.594
C5 0.000 1.180 -0.922
C6 0.000 -1.180 -0.922
C7 0.000 0.736 -2.218
C8 0.000 -0.736 -2.218
H9 0.000 1.599 3.164
H10 0.000 -1.599 3.164
H11 0.000 2.217 -0.576
H12 0.000 -2.217 -0.576
H13 0.000 1.362 -3.114
H14 0.000 -1.362 -3.114

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 C7 C8 H9 H10 H11 H12 H13 H14
C11.35502.72182.72181.47071.47072.29492.29493.58483.58482.28022.28023.35893.3589
C21.35501.44741.44742.52512.52513.60453.60452.44772.44772.91142.91144.63014.6301
C32.72181.44741.33973.55223.97264.81215.01291.09002.33913.52744.28765.75016.0592
C42.72181.44741.33973.97263.55225.01294.81212.33911.09004.28763.52746.05925.7501
C51.47072.52513.55223.97262.36091.37022.31404.10714.94141.09303.41542.20053.3578
C61.47072.52513.97263.55222.36092.31401.37024.94144.10713.41541.09303.35782.2005
C72.29493.60454.81215.01291.37022.31401.47275.45075.86682.21113.37941.09362.2823
C82.29493.60455.01294.81212.31401.37021.47275.86685.45073.37942.21112.28231.0936
H93.58482.44771.09002.33914.10714.94145.45075.86683.19733.79115.34346.28326.9419
H103.58482.44772.33911.09004.94144.10715.86685.45073.19735.34343.79116.94196.2832
H112.28022.91143.52744.28761.09303.41542.21113.37943.79115.34344.43472.67864.3884
H122.28022.91144.28763.52743.41541.09303.37942.21115.34343.79114.43474.38842.6786
H133.35894.63015.75016.05922.20053.35781.09362.28236.28326.94192.67864.38842.7248
H143.35894.63016.05925.75013.35782.20052.28231.09366.94196.28324.38842.67862.7248

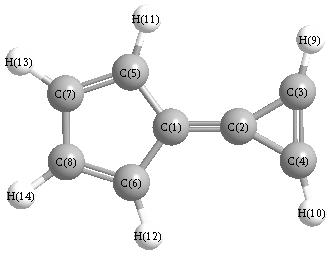
picture of Calicene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 152.432 C1 C2 C4 152.432
C1 C5 C7 107.706 C1 C5 H11 124.953
C1 C6 C8 107.706 C1 C6 H12 124.953
C2 C1 C5 126.617 C2 C1 C6 126.617
C2 C3 C4 62.432 C2 C3 H9 149.127
C2 C4 C3 62.432 C2 C4 H10 149.127
C3 C2 C4 55.135 C3 C4 H10 148.441
C4 C3 H9 148.441 C5 C1 C6 106.766
C5 C7 C8 108.911 C5 C7 H13 126.166
C6 C8 C7 108.911 C6 C8 H14 126.166
C7 C5 H11 127.341 C7 C8 H14 124.922
C8 C6 H12 127.341 C8 C7 H13 124.922
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability