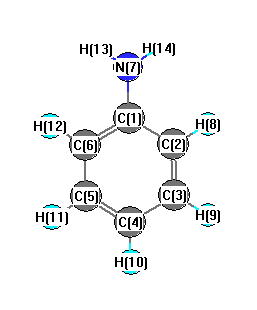
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3454 |
3434 |
9.41 |
|
|
|
| 2 |
A' |
3138 |
3120 |
13.09 |
|
|
|
| 3 |
A' |
3114 |
3096 |
2.73 |
|
|
|
| 4 |
A' |
3095 |
3077 |
19.08 |
|
|
|
| 5 |
A' |
1627 |
1618 |
117.46 |
|
|
|
| 6 |
A' |
1581 |
1571 |
29.11 |
|
|
|
| 7 |
A' |
1489 |
1480 |
52.73 |
|
|
|
| 8 |
A' |
1290 |
1283 |
36.65 |
|
|
|
| 9 |
A' |
1153 |
1147 |
6.92 |
|
|
|
| 10 |
A' |
1025 |
1019 |
3.12 |
|
|
|
| 11 |
A' |
976 |
970 |
0.37 |
|
|
|
| 12 |
A' |
948 |
943 |
0.07 |
|
|
|
| 13 |
A' |
847 |
842 |
6.01 |
|
|
|
| 14 |
A' |
815 |
810 |
1.50 |
|
|
|
| 15 |
A' |
737 |
733 |
47.88 |
|
|
|
| 16 |
A' |
683 |
679 |
17.99 |
|
|
|
| 17 |
A' |
581 |
578 |
259.64 |
|
|
|
| 18 |
A' |
518 |
515 |
3.66 |
|
|
|
| 19 |
A' |
490 |
487 |
61.00 |
|
|
|
| 20 |
A' |
214 |
213 |
4.87 |
|
|
|
| 21 |
A" |
3558 |
3537 |
9.14 |
|
|
|
| 22 |
A" |
3119 |
3101 |
35.17 |
|
|
|
| 23 |
A" |
3096 |
3078 |
4.90 |
|
|
|
| 24 |
A" |
1593 |
1584 |
3.71 |
|
|
|
| 25 |
A" |
1454 |
1445 |
0.75 |
|
|
|
| 26 |
A" |
1380 |
1372 |
9.32 |
|
|
|
| 27 |
A" |
1300 |
1292 |
0.02 |
|
|
|
| 28 |
A" |
1131 |
1125 |
0.93 |
|
|
|
| 29 |
A" |
1102 |
1095 |
3.25 |
|
|
|
| 30 |
A" |
1041 |
1035 |
4.26 |
|
|
|
| 31 |
A" |
926 |
921 |
0.00 |
|
|
|
| 32 |
A" |
793 |
789 |
0.08 |
|
|
|
| 33 |
A" |
610 |
607 |
0.33 |
|
|
|
| 34 |
A" |
402 |
400 |
0.23 |
|
|
|
| 35 |
A" |
371 |
369 |
0.21 |
|
|
|
| 36 |
A" |
300 |
298 |
17.23 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 24974.8 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 24829.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.024 |
|
|
|
| 2 |
C |
0.023 |
|
|
|
| 3 |
C |
0.028 |
|
|
|
| 4 |
C |
0.009 |
|
|
|
| 5 |
C |
0.028 |
|
|
|
| 6 |
C |
0.023 |
|
|
|
| 7 |
N |
-0.113 |
|
|
|
| 8 |
H |
-0.036 |
|
|
|
| 9 |
H |
-0.025 |
|
|
|
| 10 |
H |
-0.030 |
|
|
|
| 11 |
H |
-0.025 |
|
|
|
| 12 |
H |
-0.036 |
|
|
|
| 13 |
H |
0.089 |
|
|
|
| 14 |
H |
0.089 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.983 |
1.642 |
0.000 |
1.914 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-45.689 |
-3.068 |
0.000 |
| y |
-3.068 |
-34.020 |
0.000 |
| z |
0.000 |
0.000 |
-37.072 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-10.143 |
-3.068 |
0.000 |
| y |
-3.068 |
7.361 |
0.000 |
| z |
0.000 |
0.000 |
2.782 |
|
| Polar |
|---|
| 3z2-r2 | 5.564 |
|---|
| x2-y2 | -11.669 |
|---|
| xy | -3.068 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.269 |
-0.130 |
0.000 |
| y |
-0.130 |
14.739 |
0.000 |
| z |
0.000 |
0.000 |
12.186 |
<r2> (average value of r
2) Å
2
| <r2> |
194.210 |
| (<r2>)1/2 |
13.936 |