return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3CSNH2 (Ethanethioamide)

using model chemistry: CISD/cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at CISD/cc-pVDZ
 hartrees
Energy at 0K-531.163546
Energy at 298.15K 
HF Energy-530.641319
Nuclear repulsion energy154.992867
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CISD/cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3852 3551 56.94      
2 A 3708 3418 84.82      
3 A 3292 3034 1.76      
4 A 3212 2961 11.86      
5 A 3138 2892 16.81      
6 A 1695 1562 210.52      
7 A 1525 1406 9.11      
8 A 1523 1404 33.46      
9 A 1469 1354 129.48      
10 A 1442 1329 183.32      
11 A 1384 1276 25.68      
12 A 1067 984 0.50      
13 A 1057 975 38.32      
14 A 1039 957 5.66      
15 A 767 707 11.12      
16 A 641 591 8.93      
17 A 532 491 3.64      
18 A 441 407 1.01      
19 A 390 359 3.08      
20 A 245 225 184.99      
21 A 26 24 0.68      

Unscaled Zero Point Vibrational Energy (zpe) 16222.4 cm-1
Scaled (by 0.9218) Zero Point Vibrational Energy (zpe) 14953.8 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at CISD/cc-pVDZ
ABC
0.32630 0.16478 0.11180

See section I.F.4 to change rotational constant units
Geometric Data calculated at CISD/cc-pVDZ

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.276 0.061 -0.002
S2 -1.367 -0.117 0.000
C3 1.239 -1.104 -0.000
N4 0.877 1.263 -0.002
H5 0.694 -2.048 -0.037
H6 1.911 -1.037 -0.867
H7 1.855 -1.076 0.910
H8 1.879 1.353 0.007
H9 0.310 2.094 0.007

Atom - Atom Distances (Å)
  C1 S2 C3 N4 H5 H6 H7 H8 H9
C11.65281.51161.34382.15022.15152.14832.05902.0338
S21.65282.78712.63472.82473.51413.48253.56352.7758
C31.51162.78712.39421.09061.09891.09902.53923.3304
N41.34382.63472.39423.31592.66572.69351.00551.0064
H52.15022.82471.09063.31591.78701.78563.60194.1602
H62.15153.51411.09892.66571.78701.77802.54523.6239
H72.14833.48251.09902.69351.78561.77802.59163.6397
H82.05903.56352.53921.00553.60192.54522.59161.7344
H92.03382.77583.33041.00644.16023.62393.63971.7344

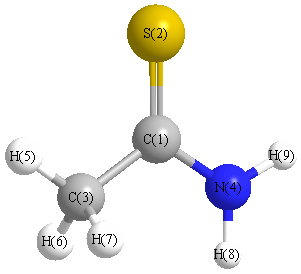
picture of Ethanethioamide state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C3 H5 110.393 C1 C3 H6 110.005
C1 C3 H7 109.749 C1 N4 H8 121.755
C1 N4 H9 119.143 S2 C1 C3 123.409
S2 C1 N4 122.761 C3 C1 N4 113.830
H5 C3 H6 109.396 H5 C3 H7 109.269
H6 C3 H7 107.986 H8 N4 H9 119.091
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability