return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3CSNH2 (Ethanethioamide)

using model chemistry: CCD/cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at CCD/cc-pVDZ
 hartrees
Energy at 0K-531.250615
Energy at 298.15K 
HF Energy-530.640421
Nuclear repulsion energy154.340272
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3771 3607 46.84      
2 A 3627 3470 81.37      
3 A 3217 3078 2.12      
4 A 3140 3004 12.64      
5 A 3066 2933 18.61      
6 A 1654 1582 209.44      
7 A 1489 1425 39.99      
8 A 1488 1423 8.72      
9 A 1431 1369 188.53      
10 A 1410 1349 123.95      
11 A 1352 1293 15.48      
12 A 1038 993 0.17      
13 A 1032 987 39.90      
14 A 1013 970 1.85      
15 A 755 723 7.35      
16 A 618 592 6.64      
17 A 514 492 6.09      
18 A 431 412 0.83      
19 A 382 365 2.67      
20 A 134 129 174.67      
21 A 47 45 3.71      

Unscaled Zero Point Vibrational Energy (zpe) 15804.3 cm-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 15120.0 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at CCD/cc-pVDZ
ABC
0.32354 0.16350 0.11092

See section I.F.4 to change rotational constant units
Geometric Data calculated at CCD/cc-pVDZ

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.273 0.060 -0.000
S2 -1.372 -0.116 0.000
C3 1.243 -1.109 0.000
N4 0.883 1.268 -0.004
H5 0.695 -2.060 -0.010
H6 1.899 -1.056 -0.889
H7 1.885 -1.067 0.900
H8 1.891 1.355 0.016
H9 0.316 2.107 0.009

Atom - Atom Distances (Å)
  C1 S2 C3 N4 H5 H6 H7 H8 H9
C11.65481.51911.35252.16162.16362.16312.07182.0467
S21.65482.79732.64602.83703.51833.51013.57942.7915
C31.51912.79732.40351.09781.10611.10622.54713.3464
N41.35252.64602.40353.33262.68642.69661.01191.0127
H52.16162.83701.09783.33261.79751.79703.61784.1835
H62.16363.51831.10612.68641.79751.78902.57513.6495
H72.16313.51011.10622.69661.79701.78902.57803.6504
H82.07183.57942.54711.01193.61782.57512.57801.7451
H92.04672.79153.34641.01274.18353.64953.65041.7451

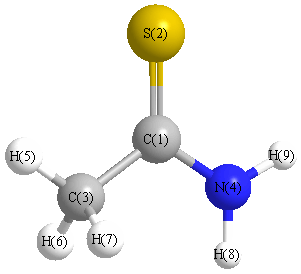
picture of Ethanethioamide state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C3 H5 110.344 C1 C3 H6 110.016
C1 C3 H7 109.968 C1 N4 H8 121.722
C1 N4 H9 119.150 S2 C1 C3 123.551
S2 C1 N4 122.931 C3 C1 N4 113.518
H5 C3 H6 109.296 H5 C3 H7 109.242
H6 C3 H7 107.932 H8 N4 H9 119.084
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability