return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C2H5NO3 (Nitric acid, ethyl ester)

using model chemistry: QCISD(T)/cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A'
Energy calculated at QCISD(T)/cc-pVDZ
 hartrees
Energy at 0K-358.628845
Energy at 298.15K-358.636730
HF Energy-357.532406
Nuclear repulsion energy249.707279
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD(T)/cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3151 3003        
2 A' 3071 2927        
3 A' 3059 2916        
4 A' 1741 1659        
5 A' 1514 1443        
6 A' 1489 1420        
7 A' 1423 1357        
8 A' 1387 1322        
9 A' 1320 1258        
10 A' 1145 1091        
11 A' 1069 1019        
12 A' 930 887        
13 A' 856 816        
14 A' 698 665        
15 A' 561 534        
16 A' 377 359        
17 A' 229 218        
18 A" 3160 3012        
19 A" 3126 2980        
20 A" 1474 1405        
21 A" 1282 1222        
22 A" 1178 1122        
23 A" 822 784        
24 A" 757 721        
25 A" 259 246        
26 A" 122 116        
27 A" 100 95        

Unscaled Zero Point Vibrational Energy (zpe) 18148.5 cm-1
Scaled (by 0.9531) Zero Point Vibrational Energy (zpe) 17297.3 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD(T)/cc-pVDZ
ABC
0.32830 0.07305 0.06116

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD(T)/cc-pVDZ

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
N1 1.110 -0.319 0.000
O2 0.000 0.578 0.000
O3 2.169 0.260 0.000
O4 0.867 -1.511 0.000
C5 -1.276 -0.110 0.000
C6 -2.323 0.996 0.000
H7 -1.351 -0.752 0.897
H8 -1.351 -0.752 -0.897
H9 -3.332 0.545 0.000
H10 -2.219 1.631 0.898
H11 -2.219 1.631 -0.898

Atom - Atom Distances (Å)
  N1 O2 O3 O4 C5 C6 H7 H8 H9 H10 H11
N11.42731.20731.21682.39503.67632.65502.65504.52533.96123.9612
O21.42732.19252.26221.44962.36032.09722.09723.33222.61492.6149
O31.20732.19252.19873.46504.55213.77123.77125.50874.68404.6840
O41.21682.26222.19872.56044.05772.51062.51064.67564.49514.4951
C52.39501.44963.46502.56041.52331.10531.10532.15802.17442.1744
C63.67632.36034.55214.05771.52332.19182.19181.10541.10461.1046
H72.65502.09723.77122.51061.10532.19181.79402.53162.53613.1068
H82.65502.09723.77122.51061.10532.19181.79402.53163.10682.5361
H94.52533.33225.50874.67562.15801.10542.53162.53161.79611.7961
H103.96122.61494.68404.49512.17441.10462.53613.10681.79611.7950
H113.96122.61494.68404.49512.17441.10463.10682.53611.79611.7950

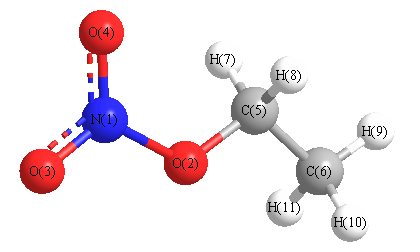
picture of Nitric acid, ethyl ester state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
N1 O2 C5 112.713 O2 N1 O3 112.380
O2 N1 O4 117.431 O2 C5 C6 105.080
O2 C5 H7 109.602 O2 C5 H8 109.602
O3 N1 O4 130.190 C5 C6 H9 109.325
C5 C6 H10 110.663 C5 C6 H11 110.663
C6 C5 H7 112.002 C6 C5 H8 112.002
H7 C5 H8 108.494 H9 C6 H10 108.725
H9 C6 H11 108.725 H10 C6 H11 108.692
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability