return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C2H5NO3 (Nitric acid, ethyl ester)

using model chemistry: CCSD(T)/cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A'
Energy calculated at CCSD(T)/cc-pVDZ
 hartrees
Energy at 0K-358.626928
Energy at 298.15K-358.634828
HF Energy-357.533151
Nuclear repulsion energy249.930592
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCSD(T)/cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3152 3085        
2 A' 3073 3008        
3 A' 3060 2996        
4 A' 1752 1715        
5 A' 1515 1483        
6 A' 1490 1458        
7 A' 1425 1395        
8 A' 1389 1359        
9 A' 1324 1296        
10 A' 1146 1122        
11 A' 1070 1048        
12 A' 933 913        
13 A' 864 846        
14 A' 704 689        
15 A' 564 552        
16 A' 379 371        
17 A' 230 225        
18 A" 3161 3094        
19 A" 3128 3062        
20 A" 1474 1443        
21 A" 1283 1255        
22 A" 1178 1153        
23 A" 823 806        
24 A" 761 744        
25 A" 259 253        
26 A" 122 119        
27 A" 101 99        

Unscaled Zero Point Vibrational Energy (zpe) 18178.4 cm-1
Scaled (by 0.9788) Zero Point Vibrational Energy (zpe) 17793.0 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at CCSD(T)/cc-pVDZ
ABC
0.32883 0.07318 0.06127

See section I.F.4 to change rotational constant units
Geometric Data calculated at CCSD(T)/cc-pVDZ

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
N1 1.109 -0.316 0.000
O2 0.000 0.576 0.000
O3 2.167 0.265 0.000
O4 0.870 -1.508 0.000
C5 -1.275 -0.114 0.000
C6 -2.323 0.992 0.000
H7 -1.349 -0.755 0.897
H8 -1.349 -0.755 -0.897
H9 -3.331 0.539 0.000
H10 -2.219 1.627 0.897
H11 -2.219 1.627 -0.897

Atom - Atom Distances (Å)
  N1 O2 O3 O4 C5 C6 H7 H8 H9 H10 H11
N11.42281.20671.21592.39233.67242.65342.65344.52193.95683.9568
O21.42282.18892.25811.44942.35982.09682.09683.33162.61452.6145
O31.20672.18892.19653.46244.54803.76943.76945.50504.67954.6795
O41.21592.25812.19652.55834.05502.50952.50954.67374.49204.4920
C52.39231.44943.46242.55831.52321.10521.10522.15772.17432.1743
C63.67242.35984.54804.05501.52322.19162.19161.10531.10451.1045
H72.65342.09683.76942.50951.10522.19161.79392.53142.53593.1066
H82.65342.09683.76942.50951.10522.19161.79392.53143.10662.5359
H94.52193.33165.50504.67372.15771.10532.53142.53141.79601.7960
H103.95682.61454.67954.49202.17431.10452.53593.10661.79601.7950
H113.95682.61454.67954.49202.17431.10453.10662.53591.79601.7950

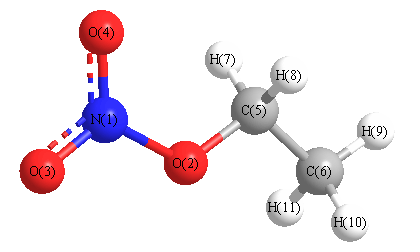
picture of Nitric acid, ethyl ester state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
N1 O2 C5 112.798 O2 N1 O3 112.437
O2 N1 O4 117.471 O2 C5 C6 105.069
O2 C5 H7 109.594 O2 C5 H8 109.594
O3 N1 O4 130.092 C5 C6 H9 109.314
C5 C6 H10 110.666 C5 C6 H11 110.666
C6 C5 H7 112.009 C6 C5 H8 112.009
H7 C5 H8 108.504 H9 C6 H10 108.726
H9 C6 H11 108.726 H10 C6 H11 108.696
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability