return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C2H5NO3 (Nitric acid, ethyl ester)

using model chemistry: QCISD/cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A'
Energy calculated at QCISD/cc-pVDZ
 hartrees
Energy at 0K-358.591970
Energy at 298.15K-358.599962
HF Energy-357.536707
Nuclear repulsion energy251.039958
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3165 3037 16.04      
2 A' 3095 2969 11.53      
3 A' 3075 2951 14.35      
4 A' 1762 1690 409.23      
5 A' 1526 1464 0.96      
6 A' 1500 1439 4.07      
7 A' 1443 1384 5.00      
8 A' 1409 1352 65.62      
9 A' 1356 1301 202.10      
10 A' 1158 1111 17.74      
11 A' 1085 1041 69.94      
12 A' 954 916 94.84      
13 A' 898 862 143.79      
14 A' 730 700 10.09      
15 A' 580 557 1.68      
16 A' 388 372 1.09      
17 A' 235 225 0.34      
18 A" 3175 3046 29.05      
19 A" 3149 3021 7.20      
20 A" 1483 1423 5.59      
21 A" 1296 1243 0.88      
22 A" 1189 1141 4.78      
23 A" 831 797 0.05      
24 A" 789 757 18.76      
25 A" 262 251 0.54      
26 A" 124 119 1.16      
27 A" 106 102 0.21      

Unscaled Zero Point Vibrational Energy (zpe) 18381.3 cm-1
Scaled (by 0.9594) Zero Point Vibrational Energy (zpe) 17635.0 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/cc-pVDZ
ABC
0.33175 0.07375 0.06177

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/cc-pVDZ

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
N1 1.105 -0.303 0.000
O2 0.000 0.564 0.000
O3 2.153 0.284 0.000
O4 0.883 -1.494 0.000
C5 -1.272 -0.127 0.000
C6 -2.322 0.974 0.000
H7 -1.343 -0.767 0.896
H8 -1.343 -0.767 -0.896
H9 -3.328 0.519 0.000
H10 -2.221 1.609 0.896
H11 -2.221 1.609 -0.896

Atom - Atom Distances (Å)
  N1 O2 O3 O4 C5 C6 H7 H8 H9 H10 H11
N11.40431.20141.21112.38323.65682.64742.64744.50833.93943.9394
O21.40432.17122.23911.44762.35782.09232.09233.32832.61312.6131
O31.20142.17122.18463.44964.52783.75873.75875.48604.65744.6574
O41.21112.23912.18462.55184.04462.50692.50694.66734.47934.4793
C52.38321.44763.44962.55181.52111.10331.10332.15512.17192.1719
C63.65682.35784.52784.04461.52112.18902.18901.10391.10321.1032
H72.64742.09233.75872.50691.10332.18901.79162.52942.53323.1030
H82.64742.09233.75872.50691.10332.18901.79162.52943.10302.5332
H94.50833.32835.48604.66732.15511.10392.52942.52941.79341.7934
H103.93942.61314.65744.47932.17191.10322.53323.10301.79341.7927
H113.93942.61314.65744.47932.17191.10323.10302.53321.79341.7927

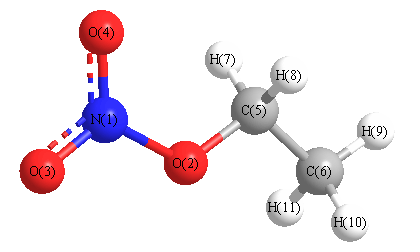
picture of Nitric acid, ethyl ester state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
N1 O2 C5 113.364 O2 N1 O3 112.635
O2 N1 O4 117.576 O2 C5 C6 105.141
O2 C5 H7 109.476 O2 C5 H8 109.476
O3 N1 O4 129.789 C5 C6 H9 109.341
C5 C6 H10 110.698 C5 C6 H11 110.698
C6 C5 H7 112.057 C6 C5 H8 112.057
H7 C5 H8 108.561 H9 C6 H10 108.690
H9 C6 H11 108.690 H10 C6 H11 108.672
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability