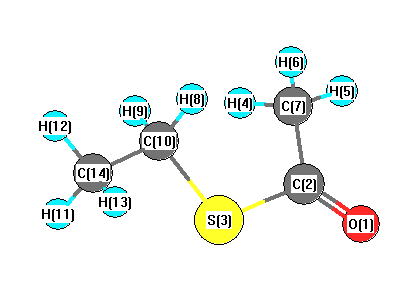
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3099 |
3081 |
5.03 |
|
|
|
| 2 |
A |
3070 |
3052 |
17.68 |
|
|
|
| 3 |
A |
3062 |
3044 |
3.91 |
|
|
|
| 4 |
A |
3059 |
3041 |
14.03 |
|
|
|
| 5 |
A |
3037 |
3019 |
3.65 |
|
|
|
| 6 |
A |
2982 |
2965 |
17.46 |
|
|
|
| 7 |
A |
2980 |
2963 |
1.62 |
|
|
|
| 8 |
A |
2977 |
2960 |
24.83 |
|
|
|
| 9 |
A |
1744 |
1733 |
280.41 |
|
|
|
| 10 |
A |
1426 |
1417 |
2.84 |
|
|
|
| 11 |
A |
1413 |
1405 |
9.48 |
|
|
|
| 12 |
A |
1405 |
1397 |
0.23 |
|
|
|
| 13 |
A |
1393 |
1385 |
11.50 |
|
|
|
| 14 |
A |
1378 |
1370 |
18.17 |
|
|
|
| 15 |
A |
1343 |
1335 |
2.54 |
|
|
|
| 16 |
A |
1303 |
1296 |
23.16 |
|
|
|
| 17 |
A |
1227 |
1220 |
24.07 |
|
|
|
| 18 |
A |
1215 |
1208 |
0.18 |
|
|
|
| 19 |
A |
1101 |
1094 |
158.74 |
|
|
|
| 20 |
A |
1046 |
1040 |
6.54 |
|
|
|
| 21 |
A |
992 |
986 |
0.02 |
|
|
|
| 22 |
A |
970 |
964 |
4.48 |
|
|
|
| 23 |
A |
968 |
963 |
2.23 |
|
|
|
| 24 |
A |
912 |
907 |
40.16 |
|
|
|
| 25 |
A |
763 |
758 |
5.38 |
|
|
|
| 26 |
A |
667 |
663 |
0.08 |
|
|
|
| 27 |
A |
615 |
612 |
66.02 |
|
|
|
| 28 |
A |
495 |
492 |
0.61 |
|
|
|
| 29 |
A |
421 |
418 |
2.67 |
|
|
|
| 30 |
A |
358 |
356 |
1.15 |
|
|
|
| 31 |
A |
291 |
289 |
1.08 |
|
|
|
| 32 |
A |
250 |
249 |
0.04 |
|
|
|
| 33 |
A |
151 |
151 |
0.72 |
|
|
|
| 34 |
A |
149 |
148 |
0.28 |
|
|
|
| 35 |
A |
92 |
91 |
0.33 |
|
|
|
| 36 |
A |
56 |
56 |
0.03 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 24203.6 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 24063.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.170 |
|
|
|
| 2 |
C |
-0.039 |
|
|
|
| 3 |
S |
0.008 |
|
|
|
| 4 |
H |
0.065 |
|
|
|
| 5 |
H |
0.055 |
|
|
|
| 6 |
H |
0.065 |
|
|
|
| 7 |
C |
-0.023 |
|
|
|
| 8 |
H |
0.071 |
|
|
|
| 9 |
H |
0.071 |
|
|
|
| 10 |
C |
-0.206 |
|
|
|
| 11 |
H |
0.054 |
|
|
|
| 12 |
H |
0.043 |
|
|
|
| 13 |
H |
0.054 |
|
|
|
| 14 |
C |
-0.047 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-2.378 |
3.113 |
-0.000 |
3.917 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-46.042 |
3.370 |
-0.003 |
| y |
3.370 |
-44.526 |
0.001 |
| z |
-0.003 |
0.001 |
-44.056 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.751 |
3.370 |
-0.003 |
| y |
3.370 |
0.523 |
0.001 |
| z |
-0.003 |
0.001 |
1.228 |
|
| Polar |
|---|
| 3z2-r2 | 2.456 |
|---|
| x2-y2 | -1.516 |
|---|
| xy | 3.370 |
|---|
| xz | -0.003 |
|---|
| yz | 0.001 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
14.001 |
0.111 |
0.000 |
| y |
0.111 |
9.449 |
0.000 |
| z |
0.000 |
0.000 |
6.579 |
<r2> (average value of r
2) Å
2
| <r2> |
255.351 |
| (<r2>)1/2 |
15.980 |