Jump to
S1C2
Vibrational Frequencies calculated at QCISD/cc-pVDZ
Geometric Data calculated at QCISD/cc-pVDZ
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at QCISD/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -553.176234 |
| Energy at 298.15K | -553.181673 |
| HF Energy | -552.448357 |
| Nuclear repulsion energy | 200.256895 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3277 |
3144 |
7.18 |
|
|
|
| 2 |
A |
3272 |
3139 |
5.31 |
|
|
|
| 3 |
A |
3204 |
3073 |
6.25 |
|
|
|
| 4 |
A |
3194 |
3064 |
6.96 |
|
|
|
| 5 |
A |
3178 |
3049 |
1.41 |
|
|
|
| 6 |
A |
3170 |
3042 |
2.12 |
|
|
|
| 7 |
A |
1675 |
1607 |
26.79 |
|
|
|
| 8 |
A |
1668 |
1600 |
17.31 |
|
|
|
| 9 |
A |
1426 |
1368 |
4.52 |
|
|
|
| 10 |
A |
1420 |
1362 |
4.69 |
|
|
|
| 11 |
A |
1302 |
1249 |
2.37 |
|
|
|
| 12 |
A |
1291 |
1238 |
12.77 |
|
|
|
| 13 |
A |
1061 |
1018 |
7.26 |
|
|
|
| 14 |
A |
1034 |
992 |
9.46 |
|
|
|
| 15 |
A |
997 |
956 |
20.92 |
|
|
|
| 16 |
A |
984 |
944 |
32.34 |
|
|
|
| 17 |
A |
928 |
890 |
23.15 |
|
|
|
| 18 |
A |
890 |
854 |
29.63 |
|
|
|
| 19 |
A |
750 |
720 |
9.44 |
|
|
|
| 20 |
A |
701 |
672 |
3.89 |
|
|
|
| 21 |
A |
625 |
599 |
13.13 |
|
|
|
| 22 |
A |
599 |
575 |
9.68 |
|
|
|
| 23 |
A |
456 |
437 |
0.25 |
|
|
|
| 24 |
A |
367 |
352 |
0.47 |
|
|
|
| 25 |
A |
225 |
216 |
0.91 |
|
|
|
| 26 |
A |
138 |
133 |
3.11 |
|
|
|
| 27 |
A |
85 |
81 |
1.20 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 18957.4 cm
-1
Scaled (by 0.9594) Zero Point Vibrational Energy (zpe) 18187.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD/cc-pVDZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| S1 |
0.015 |
-0.998 |
0.003 |
| C2 |
1.085 |
0.372 |
0.386 |
| C3 |
-1.572 |
-0.194 |
-0.028 |
| C4 |
2.268 |
0.564 |
-0.230 |
| C5 |
-1.823 |
1.127 |
-0.129 |
| H6 |
0.760 |
1.013 |
1.217 |
| H7 |
-2.396 |
-0.917 |
0.043 |
| H8 |
2.947 |
1.357 |
0.107 |
| H9 |
2.592 |
-0.066 |
-1.067 |
| H10 |
-1.025 |
1.871 |
-0.228 |
| H11 |
-2.857 |
1.487 |
-0.117 |
Atom - Atom Distances (Å)
| |
S1 |
C2 |
C3 |
C4 |
C5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| S1 | | 1.7802 | 1.7790 | 2.7518 | 2.8122 | 2.4640 | 2.4125 | 3.7619 | 2.9421 | 3.0604 | 3.7998 |
C2 | 1.7802 | | 2.7484 | 1.3478 | 3.0479 | 1.0986 | 3.7279 | 2.1246 | 2.1387 | 2.6602 | 4.1279 | C3 | 1.7790 | 2.7484 | | 3.9198 | 1.3488 | 2.9061 | 1.0979 | 4.7797 | 4.2936 | 2.1464 | 2.1183 | C4 | 2.7518 | 1.3478 | 3.9198 | | 4.1305 | 2.1382 | 4.9013 | 1.0965 | 1.0963 | 3.5428 | 5.2091 | C5 | 2.8122 | 3.0479 | 1.3488 | 4.1305 | | 2.9143 | 2.1297 | 4.7805 | 4.6679 | 1.0955 | 1.0957 | H6 | 2.4640 | 1.0986 | 2.9061 | 2.1382 | 2.9143 | | 3.8808 | 2.4769 | 3.1206 | 2.4517 | 3.8844 | H7 | 2.4125 | 3.7279 | 1.0979 | 4.9013 | 2.1297 | 3.8808 | | 5.8068 | 5.1803 | 3.1188 | 2.4530 | H8 | 3.7619 | 2.1246 | 4.7797 | 1.0965 | 4.7805 | 2.4769 | 5.8068 | | 1.8781 | 4.0185 | 5.8097 | H9 | 2.9421 | 2.1387 | 4.2936 | 1.0963 | 4.6679 | 3.1206 | 5.1803 | 1.8781 | | 4.1875 | 5.7451 | H10 | 3.0604 | 2.6602 | 2.1464 | 3.5428 | 1.0955 | 2.4517 | 3.1188 | 4.0185 | 4.1875 | | 1.8756 | H11 | 3.7998 | 4.1279 | 2.1183 | 5.2091 | 1.0957 | 3.8844 | 2.4530 | 5.8097 | 5.7451 | 1.8756 | |
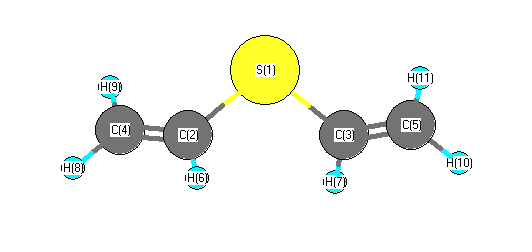 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| S1 |
C2 |
C4 |
122.623 |
|
S1 |
C2 |
H6 |
115.682 |
| S1 |
C3 |
C5 |
127.540 |
|
S1 |
C3 |
H7 |
111.784 |
| C2 |
S1 |
C3 |
101.106 |
|
C2 |
C4 |
H8 |
120.385 |
| C2 |
C4 |
H9 |
121.760 |
|
C3 |
C5 |
H10 |
122.495 |
| C3 |
C5 |
H11 |
119.765 |
|
C4 |
C2 |
H6 |
121.523 |
| C5 |
C3 |
H7 |
120.675 |
|
H8 |
C4 |
H9 |
117.850 |
| H10 |
C5 |
H11 |
117.739 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
