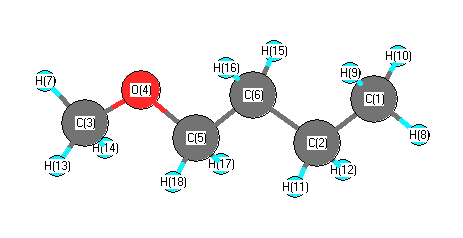
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3055 |
3037 |
27.41 |
|
|
|
| 2 |
A' |
3051 |
3033 |
31.62 |
|
|
|
| 3 |
A' |
2966 |
2948 |
49.54 |
|
|
|
| 4 |
A' |
2962 |
2945 |
21.66 |
|
|
|
| 5 |
A' |
2947 |
2930 |
20.64 |
|
|
|
| 6 |
A' |
2881 |
2864 |
90.40 |
|
|
|
| 7 |
A' |
2851 |
2835 |
43.30 |
|
|
|
| 8 |
A' |
1455 |
1446 |
1.01 |
|
|
|
| 9 |
A' |
1434 |
1426 |
2.33 |
|
|
|
| 10 |
A' |
1426 |
1417 |
12.12 |
|
|
|
| 11 |
A' |
1420 |
1412 |
0.25 |
|
|
|
| 12 |
A' |
1413 |
1405 |
0.09 |
|
|
|
| 13 |
A' |
1404 |
1395 |
0.04 |
|
|
|
| 14 |
A' |
1364 |
1356 |
33.75 |
|
|
|
| 15 |
A' |
1347 |
1339 |
0.43 |
|
|
|
| 16 |
A' |
1324 |
1316 |
1.69 |
|
|
|
| 17 |
A' |
1233 |
1226 |
2.33 |
|
|
|
| 18 |
A' |
1172 |
1165 |
39.70 |
|
|
|
| 19 |
A' |
1138 |
1132 |
152.76 |
|
|
|
| 20 |
A' |
1096 |
1089 |
8.33 |
|
|
|
| 21 |
A' |
1062 |
1056 |
0.73 |
|
|
|
| 22 |
A' |
1025 |
1020 |
2.23 |
|
|
|
| 23 |
A' |
960 |
955 |
21.49 |
|
|
|
| 24 |
A' |
887 |
882 |
5.24 |
|
|
|
| 25 |
A' |
475 |
472 |
1.03 |
|
|
|
| 26 |
A' |
374 |
371 |
2.52 |
|
|
|
| 27 |
A' |
303 |
301 |
0.59 |
|
|
|
| 28 |
A' |
131 |
130 |
0.68 |
|
|
|
| 29 |
A" |
3042 |
3025 |
45.52 |
|
|
|
| 30 |
A" |
3012 |
2994 |
37.35 |
|
|
|
| 31 |
A" |
2979 |
2962 |
4.66 |
|
|
|
| 32 |
A" |
2926 |
2909 |
65.39 |
|
|
|
| 33 |
A" |
2880 |
2863 |
77.16 |
|
|
|
| 34 |
A" |
1419 |
1411 |
6.51 |
|
|
|
| 35 |
A" |
1398 |
1390 |
5.88 |
|
|
|
| 36 |
A" |
1269 |
1261 |
0.09 |
|
|
|
| 37 |
A" |
1251 |
1243 |
0.04 |
|
|
|
| 38 |
A" |
1193 |
1186 |
1.73 |
|
|
|
| 39 |
A" |
1146 |
1140 |
6.03 |
|
|
|
| 40 |
A" |
1116 |
1109 |
0.13 |
|
|
|
| 41 |
A" |
915 |
909 |
0.00 |
|
|
|
| 42 |
A" |
789 |
785 |
0.44 |
|
|
|
| 43 |
A" |
730 |
725 |
3.91 |
|
|
|
| 44 |
A" |
250 |
249 |
0.70 |
|
|
|
| 45 |
A" |
232 |
231 |
1.67 |
|
|
|
| 46 |
A" |
135 |
134 |
1.04 |
|
|
|
| 47 |
A" |
96 |
96 |
1.07 |
|
|
|
| 48 |
A" |
71 |
71 |
0.64 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 35000.8 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 34797.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.045 |
|
|
|
| 2 |
C |
-0.089 |
|
|
|
| 3 |
C |
0.076 |
|
|
|
| 4 |
O |
-0.279 |
|
|
|
| 5 |
C |
0.114 |
|
|
|
| 6 |
C |
-0.046 |
|
|
|
| 7 |
H |
0.036 |
|
|
|
| 8 |
H |
0.027 |
|
|
|
| 9 |
H |
0.032 |
|
|
|
| 10 |
H |
0.032 |
|
|
|
| 11 |
H |
0.023 |
|
|
|
| 12 |
H |
0.023 |
|
|
|
| 13 |
H |
0.015 |
|
|
|
| 14 |
H |
0.015 |
|
|
|
| 15 |
H |
0.026 |
|
|
|
| 16 |
H |
0.026 |
|
|
|
| 17 |
H |
0.007 |
|
|
|
| 18 |
H |
0.007 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.786 |
-0.514 |
0.000 |
0.939 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-41.082 |
1.112 |
0.000 |
| y |
1.112 |
-37.152 |
0.000 |
| z |
0.000 |
0.000 |
-39.916 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.548 |
1.112 |
0.000 |
| y |
1.112 |
3.347 |
0.000 |
| z |
0.000 |
0.000 |
-0.799 |
|
| Polar |
|---|
| 3z2-r2 | -1.599 |
|---|
| x2-y2 | -3.930 |
|---|
| xy | 1.112 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.543 |
1.526 |
0.000 |
| y |
1.526 |
11.240 |
0.000 |
| z |
0.000 |
0.000 |
8.356 |
<r2> (average value of r
2) Å
2
| <r2> |
296.202 |
| (<r2>)1/2 |
17.211 |