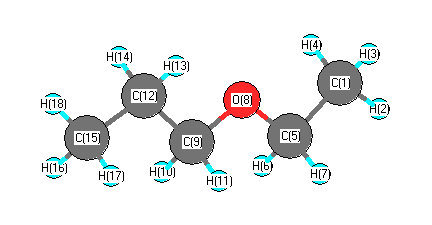
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3071 |
3053 |
20.30 |
|
|
|
| 2 |
A' |
3055 |
3038 |
26.26 |
|
|
|
| 3 |
A' |
2981 |
2964 |
19.39 |
|
|
|
| 4 |
A' |
2973 |
2956 |
26.95 |
|
|
|
| 5 |
A' |
2966 |
2949 |
27.36 |
|
|
|
| 6 |
A' |
2871 |
2854 |
96.58 |
|
|
|
| 7 |
A' |
2853 |
2836 |
27.71 |
|
|
|
| 8 |
A' |
1461 |
1453 |
1.23 |
|
|
|
| 9 |
A' |
1440 |
1432 |
6.62 |
|
|
|
| 10 |
A' |
1431 |
1423 |
3.24 |
|
|
|
| 11 |
A' |
1425 |
1417 |
1.38 |
|
|
|
| 12 |
A' |
1417 |
1409 |
1.23 |
|
|
|
| 13 |
A' |
1387 |
1379 |
5.07 |
|
|
|
| 14 |
A' |
1347 |
1340 |
1.25 |
|
|
|
| 15 |
A' |
1342 |
1334 |
32.18 |
|
|
|
| 16 |
A' |
1320 |
1312 |
18.35 |
|
|
|
| 17 |
A' |
1264 |
1257 |
15.36 |
|
|
|
| 18 |
A' |
1140 |
1134 |
51.86 |
|
|
|
| 19 |
A' |
1129 |
1122 |
189.50 |
|
|
|
| 20 |
A' |
1079 |
1072 |
5.16 |
|
|
|
| 21 |
A' |
1049 |
1043 |
4.08 |
|
|
|
| 22 |
A' |
1020 |
1014 |
13.26 |
|
|
|
| 23 |
A' |
888 |
883 |
2.65 |
|
|
|
| 24 |
A' |
872 |
867 |
9.17 |
|
|
|
| 25 |
A' |
478 |
475 |
3.16 |
|
|
|
| 26 |
A' |
397 |
395 |
0.76 |
|
|
|
| 27 |
A' |
295 |
293 |
0.50 |
|
|
|
| 28 |
A' |
133 |
133 |
0.27 |
|
|
|
| 29 |
A" |
3068 |
3051 |
22.00 |
|
|
|
| 30 |
A" |
3043 |
3025 |
47.59 |
|
|
|
| 31 |
A" |
3013 |
2995 |
7.39 |
|
|
|
| 32 |
A" |
2894 |
2877 |
37.51 |
|
|
|
| 33 |
A" |
2881 |
2864 |
102.24 |
|
|
|
| 34 |
A" |
1422 |
1414 |
6.68 |
|
|
|
| 35 |
A" |
1402 |
1394 |
6.15 |
|
|
|
| 36 |
A" |
1256 |
1248 |
0.26 |
|
|
|
| 37 |
A" |
1238 |
1231 |
1.72 |
|
|
|
| 38 |
A" |
1207 |
1200 |
1.86 |
|
|
|
| 39 |
A" |
1148 |
1141 |
7.17 |
|
|
|
| 40 |
A" |
1120 |
1114 |
0.79 |
|
|
|
| 41 |
A" |
868 |
863 |
1.70 |
|
|
|
| 42 |
A" |
795 |
791 |
1.00 |
|
|
|
| 43 |
A" |
748 |
744 |
2.45 |
|
|
|
| 44 |
A" |
247 |
246 |
0.70 |
|
|
|
| 45 |
A" |
234 |
232 |
0.05 |
|
|
|
| 46 |
A" |
150 |
149 |
2.45 |
|
|
|
| 47 |
A" |
94 |
93 |
0.90 |
|
|
|
| 48 |
A" |
66 |
65 |
0.21 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 34986.8 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 34783.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.060 |
|
|
|
| 2 |
H |
0.030 |
|
|
|
| 3 |
H |
0.042 |
|
|
|
| 4 |
H |
0.042 |
|
|
|
| 5 |
C |
0.069 |
|
|
|
| 6 |
H |
0.009 |
|
|
|
| 7 |
H |
0.009 |
|
|
|
| 8 |
O |
-0.283 |
|
|
|
| 9 |
C |
0.116 |
|
|
|
| 10 |
H |
0.006 |
|
|
|
| 11 |
H |
0.006 |
|
|
|
| 12 |
C |
-0.072 |
|
|
|
| 13 |
H |
0.029 |
|
|
|
| 14 |
H |
0.029 |
|
|
|
| 15 |
C |
-0.070 |
|
|
|
| 16 |
H |
0.033 |
|
|
|
| 17 |
H |
0.033 |
|
|
|
| 18 |
H |
0.032 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.691 |
-0.410 |
0.000 |
0.804 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-39.003 |
1.806 |
0.000 |
| y |
1.806 |
-38.890 |
0.000 |
| z |
0.000 |
0.000 |
-39.776 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.330 |
1.806 |
0.000 |
| y |
1.806 |
0.500 |
0.000 |
| z |
0.000 |
0.000 |
-0.829 |
|
| Polar |
|---|
| 3z2-r2 | -1.659 |
|---|
| x2-y2 | -0.113 |
|---|
| xy | 1.806 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.426 |
1.261 |
0.000 |
| y |
1.261 |
11.462 |
0.000 |
| z |
0.000 |
0.000 |
8.404 |
<r2> (average value of r
2) Å
2
| <r2> |
288.691 |
| (<r2>)1/2 |
16.991 |