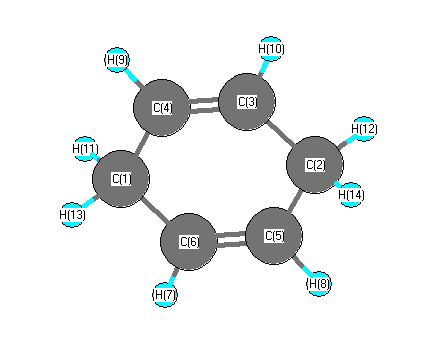
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3102 |
3084 |
0.00 |
|
|
|
| 2 |
Ag |
2913 |
2896 |
0.00 |
|
|
|
| 3 |
Ag |
1716 |
1706 |
0.00 |
|
|
|
| 4 |
Ag |
1381 |
1373 |
0.00 |
|
|
|
| 5 |
Ag |
1178 |
1171 |
0.00 |
|
|
|
| 6 |
Ag |
856 |
852 |
0.00 |
|
|
|
| 7 |
Ag |
524 |
521 |
0.00 |
|
|
|
| 8 |
Au |
1160 |
1153 |
0.00 |
|
|
|
| 9 |
Au |
957 |
951 |
0.00 |
|
|
|
| 10 |
Au |
371 |
369 |
0.00 |
|
|
|
| 11 |
B1g |
3077 |
3060 |
0.00 |
|
|
|
| 12 |
B1g |
1360 |
1352 |
0.00 |
|
|
|
| 13 |
B1g |
1298 |
1291 |
0.00 |
|
|
|
| 14 |
B1g |
1023 |
1017 |
0.00 |
|
|
|
| 15 |
B1g |
561 |
558 |
0.00 |
|
|
|
| 16 |
B1u |
2926 |
2909 |
43.15 |
|
|
|
| 17 |
B1u |
939 |
933 |
16.13 |
|
|
|
| 18 |
B1u |
605 |
602 |
53.64 |
|
|
|
| 19 |
B1u |
117 |
117 |
1.17 |
|
|
|
| 20 |
B2g |
2927 |
2910 |
0.00 |
|
|
|
| 21 |
B2g |
967 |
961 |
0.00 |
|
|
|
| 22 |
B2g |
924 |
918 |
0.00 |
|
|
|
| 23 |
B2g |
395 |
393 |
0.00 |
|
|
|
| 24 |
B2u |
3100 |
3082 |
71.69 |
|
|
|
| 25 |
B2u |
1670 |
1661 |
8.05 |
|
|
|
| 26 |
B2u |
1333 |
1325 |
0.01 |
|
|
|
| 27 |
B2u |
1128 |
1122 |
0.01 |
|
|
|
| 28 |
B2u |
938 |
932 |
0.04 |
|
|
|
| 29 |
B3g |
1169 |
1162 |
0.00 |
|
|
|
| 30 |
B3g |
697 |
693 |
0.00 |
|
|
|
| 31 |
B3u |
3078 |
3060 |
16.52 |
|
|
|
| 32 |
B3u |
2915 |
2898 |
94.48 |
|
|
|
| 33 |
B3u |
1387 |
1379 |
7.41 |
|
|
|
| 34 |
B3u |
1380 |
1372 |
0.98 |
|
|
|
| 35 |
B3u |
937 |
932 |
0.06 |
|
|
|
| 36 |
B3u |
887 |
882 |
14.36 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 25948.3 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 25797.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.036 |
|
|
|
| 2 |
C |
0.036 |
|
|
|
| 3 |
C |
-0.047 |
|
|
|
| 4 |
C |
-0.047 |
|
|
|
| 5 |
C |
-0.047 |
|
|
|
| 6 |
C |
-0.047 |
|
|
|
| 7 |
H |
-0.018 |
|
|
|
| 8 |
H |
-0.018 |
|
|
|
| 9 |
H |
-0.018 |
|
|
|
| 10 |
H |
-0.018 |
|
|
|
| 11 |
H |
0.047 |
|
|
|
| 12 |
H |
0.047 |
|
|
|
| 13 |
H |
0.047 |
|
|
|
| 14 |
H |
0.047 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-33.422 |
0.000 |
0.000 |
| y |
0.000 |
-35.889 |
0.000 |
| z |
0.000 |
0.000 |
-38.985 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
4.015 |
0.000 |
0.000 |
| y |
0.000 |
0.315 |
0.000 |
| z |
0.000 |
0.000 |
-4.329 |
|
| Polar |
|---|
| 3z2-r2 | -8.659 |
|---|
| x2-y2 | 2.467 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
12.726 |
0.000 |
0.000 |
| y |
0.000 |
9.514 |
0.000 |
| z |
0.000 |
0.000 |
5.519 |
<r2> (average value of r
2) Å
2
| <r2> |
146.337 |
| (<r2>)1/2 |
12.097 |