return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H7NO (Ethoxyacetonitrile)

using model chemistry: PBEPBE/cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at PBEPBE/cc-pVDZ
 hartrees
Energy at 0K-286.233641
Energy at 298.15K-286.241279
HF Energy-286.233641
Nuclear repulsion energy214.590088
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3076 3058 13.86      
2 A' 2986 2969 12.83      
3 A' 2883 2867 70.83      
4 A' 2873 2856 17.32      
5 A' 2286 2273 0.40      
6 A' 1453 1444 4.04      
7 A' 1426 1417 1.42      
8 A' 1414 1405 6.64      
9 A' 1378 1370 10.83      
10 A' 1335 1328 10.30      
11 A' 1307 1299 56.59      
12 A' 1123 1117 207.40      
13 A' 1107 1101 21.71      
14 A' 1028 1022 20.75      
15 A' 937 932 2.41      
16 A' 876 871 7.01      
17 A' 541 537 0.54      
18 A' 411 408 0.90      
19 A' 290 289 1.37      
20 A' 127 127 2.70      
21 A" 3076 3059 15.98      
22 A" 2914 2897 32.50      
23 A" 2897 2880 59.14      
24 A" 1403 1395 6.80      
25 A" 1239 1232 1.84      
26 A" 1189 1182 4.06      
27 A" 1130 1124 6.80      
28 A" 977 972 0.71      
29 A" 798 793 1.14      
30 A" 365 363 0.56      
31 A" 243 241 0.29      
32 A" 112 112 5.01      
33 A" 76 76 0.01      

Unscaled Zero Point Vibrational Energy (zpe) 22638.2 cm-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 22506.9 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at PBEPBE/cc-pVDZ
ABC
0.51941 0.04923 0.04617

See section I.F.4 to change rotational constant units
Geometric Data calculated at PBEPBE/cc-pVDZ

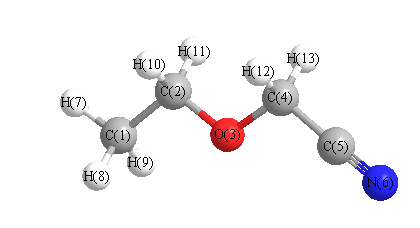
Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -1.032 2.522 0.000
C2 -1.272 1.023 0.000
O3 0.000 0.369 0.000
C4 -0.143 -1.036 0.000
C5 1.184 -1.665 0.000
N6 2.231 -2.194 0.000
H7 -1.997 3.065 0.000
H8 -0.458 2.825 0.898
H9 -0.458 2.825 -0.898
H10 -1.861 0.712 -0.899
H11 -1.861 0.712 0.899
H12 -0.705 -1.401 -0.898
H13 -0.705 -1.401 0.898

Atom - Atom Distances (Å)
  C1 C2 O3 C4 C5 N6 H7 H8 H9 H10 H11 H12 H13
C11.51772.38693.66674.73685.73481.10791.10781.10782.18442.18444.03754.0375
C21.51771.42992.34783.64094.75642.16712.17122.17121.11911.11912.64662.6466
O32.38691.42991.41242.35373.39873.35502.65392.65392.09512.09512.10642.1064
C43.66672.34781.41241.46912.64224.50053.97583.97582.61022.61021.12011.1201
C54.73683.64092.35371.46911.17335.70044.86374.86373.96623.96622.10792.1079
N65.73484.75643.39872.64221.17336.74855.76415.76415.09925.09923.17093.1709
H71.10792.16713.35504.50055.70046.74851.79841.79842.52292.52294.73544.7354
H81.10782.17122.65393.97584.86375.76411.79841.79533.10842.53664.59794.2329
H91.10782.17122.65393.97584.86375.76411.79841.79532.53663.10844.23294.5979
H102.18441.11912.09512.61023.96625.09922.52293.10842.53661.79782.40873.0050
H112.18441.11912.09512.61023.96625.09922.52292.53663.10841.79783.00502.4087
H124.03752.64662.10641.12012.10793.17094.73544.59794.23292.40873.00501.7957
H134.03752.64662.10641.12012.10793.17094.73544.23294.59793.00502.40871.7957

picture of Ethoxyacetonitrile state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 O3 108.111 C1 C2 H10 110.975
C1 C2 H11 110.975 C2 C1 H7 110.281
C2 C1 H8 110.607 C2 C1 H9 110.607
C2 O3 C4 111.383 O3 C2 H10 109.956
O3 C2 H11 109.956 O3 C4 C5 109.528
O3 C4 H12 112.043 O3 C4 H13 112.043
C4 C5 N6 178.523 C5 C4 H12 108.252
C5 C4 H13 108.252 H7 C1 H8 108.517
H7 C1 H9 108.517 H8 C1 H9 108.241
H10 C2 H11 106.875 H12 C4 H13 106.563
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability