Jump to
S1C2
Energy calculated at PBEPBE/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -189.572635 |
| Energy at 298.15K | -189.575331 |
| HF Energy | -189.572635 |
| Nuclear repulsion energy | 69.487369 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3546 |
3525 |
29.66 |
97.47 |
0.29 |
0.44 |
| 2 |
A' |
2949 |
2932 |
60.98 |
137.73 |
0.28 |
0.44 |
| 3 |
A' |
1792 |
1782 |
285.21 |
5.17 |
0.20 |
0.33 |
| 4 |
A' |
1346 |
1338 |
2.43 |
12.21 |
0.67 |
0.80 |
| 5 |
A' |
1270 |
1262 |
8.50 |
0.41 |
0.28 |
0.44 |
| 6 |
A' |
1096 |
1090 |
197.66 |
1.65 |
0.39 |
0.57 |
| 7 |
A' |
603 |
600 |
43.21 |
4.62 |
0.58 |
0.73 |
| 8 |
A" |
1000 |
994 |
3.51 |
2.24 |
0.75 |
0.86 |
| 9 |
A" |
704 |
700 |
119.99 |
2.92 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 7153.1 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 7111.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBEPBE/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.424 |
0.000 |
| O2 |
-1.043 |
-0.443 |
0.000 |
| O3 |
1.172 |
0.110 |
0.000 |
| H4 |
-0.403 |
1.468 |
0.000 |
| H5 |
-0.630 |
-1.341 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
O3 |
H4 |
H5 |
| C1 | | 1.3560 | 1.2129 | 1.1189 | 1.8735 |
O2 | 1.3560 | | 2.2822 | 2.0154 | 0.9875 | O3 | 1.2129 | 2.2822 | | 2.0788 | 2.3131 | H4 | 1.1189 | 2.0154 | 2.0788 | | 2.8176 | H5 | 1.8735 | 0.9875 | 2.3131 | 2.8176 | |
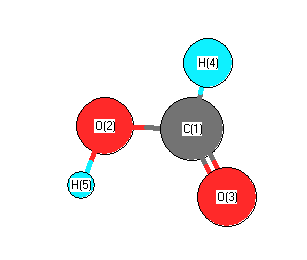 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
H5 |
105.069 |
|
O2 |
C1 |
O3 |
125.253 |
| O2 |
C1 |
H4 |
108.667 |
|
O3 |
C1 |
H4 |
126.080 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.170 |
|
|
|
| 2 |
O |
-0.139 |
|
|
|
| 3 |
O |
-0.210 |
|
|
|
| 4 |
H |
0.028 |
|
|
|
| 5 |
H |
0.151 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.311 |
-0.471 |
0.000 |
1.393 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-21.051 |
-0.070 |
0.000 |
| y |
-0.070 |
-12.902 |
0.000 |
| z |
0.000 |
0.000 |
-16.632 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-6.284 |
-0.070 |
0.000 |
| y |
-0.070 |
5.940 |
0.000 |
| z |
0.000 |
0.000 |
0.345 |
|
| Polar |
|---|
| 3z2-r2 | 0.689 |
|---|
| x2-y2 | -8.149 |
|---|
| xy | -0.070 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.208 |
0.075 |
0.000 |
| y |
0.075 |
3.143 |
0.000 |
| z |
0.000 |
0.000 |
1.399 |
<r2> (average value of r
2) Å
2
| <r2> |
37.465 |
| (<r2>)1/2 |
6.121 |
Jump to
S1C1
Energy calculated at PBEPBE/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -189.565030 |
| Energy at 298.15K | |
| HF Energy | -189.565030 |
| Nuclear repulsion energy | 69.284544 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3617 |
3596 |
25.40 |
129.33 |
0.29 |
0.45 |
| 2 |
A' |
2837 |
2821 |
112.74 |
118.62 |
0.28 |
0.44 |
| 3 |
A' |
1837 |
1826 |
235.71 |
9.27 |
0.24 |
0.39 |
| 4 |
A' |
1354 |
1346 |
0.27 |
9.07 |
0.65 |
0.79 |
| 5 |
A' |
1242 |
1235 |
279.51 |
4.16 |
0.74 |
0.85 |
| 6 |
A' |
1084 |
1077 |
25.43 |
8.96 |
0.53 |
0.70 |
| 7 |
A' |
634 |
630 |
8.56 |
0.54 |
0.75 |
0.86 |
| 8 |
A" |
987 |
981 |
0.10 |
2.93 |
0.75 |
0.86 |
| 9 |
A" |
544 |
541 |
75.27 |
3.36 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 7067.1 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 7026.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBEPBE/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.387 |
0.000 |
| O2 |
-0.905 |
-0.630 |
0.000 |
| O3 |
1.191 |
0.193 |
0.000 |
| H4 |
-0.480 |
1.408 |
0.000 |
| H5 |
-1.802 |
-0.232 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
O3 |
H4 |
H5 |
| C1 | | 1.3617 | 1.2063 | 1.1275 | 1.9053 |
O2 | 1.3617 | | 2.2515 | 2.0815 | 0.9811 | O3 | 1.2063 | 2.2515 | | 2.0655 | 3.0225 | H4 | 1.1275 | 2.0815 | 2.0655 | | 2.1061 | H5 | 1.9053 | 0.9811 | 3.0225 | 2.1061 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
H5 |
107.725 |
|
O2 |
C1 |
O3 |
122.385 |
| O2 |
C1 |
H4 |
113.145 |
|
O3 |
C1 |
H4 |
124.469 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.160 |
|
|
|
| 2 |
O |
-0.121 |
|
|
|
| 3 |
O |
-0.181 |
|
|
|
| 4 |
H |
-0.005 |
|
|
|
| 5 |
H |
0.148 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-3.315 |
1.283 |
0.000 |
3.555 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-15.175 |
-1.049 |
0.000 |
| y |
-1.049 |
-17.150 |
0.000 |
| z |
0.000 |
0.000 |
-16.638 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.719 |
-1.049 |
0.000 |
| y |
-1.049 |
-1.244 |
0.000 |
| z |
0.000 |
0.000 |
-0.475 |
|
| Polar |
|---|
| 3z2-r2 | -0.951 |
|---|
| x2-y2 | 1.976 |
|---|
| xy | -1.049 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.942 |
0.088 |
0.000 |
| y |
0.088 |
2.568 |
0.000 |
| z |
0.000 |
0.000 |
1.396 |
<r2> (average value of r
2) Å
2
| <r2> |
37.973 |
| (<r2>)1/2 |
6.162 |
