Jump to
S1C2
S1C3
Vibrational Frequencies calculated at QCISD/cc-pVDZ
Geometric Data calculated at QCISD/cc-pVDZ
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C3
Vibrational Frequencies calculated at QCISD/cc-pVDZ
Geometric Data calculated at QCISD/cc-pVDZ
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C2
Energy calculated at QCISD/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -267.632436 |
| Energy at 298.15K | -267.640783 |
| HF Energy | -266.808292 |
| Nuclear repulsion energy | 193.896884 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3158 |
3030 |
34.18 |
|
|
|
| 2 |
A |
3153 |
3025 |
41.29 |
|
|
|
| 3 |
A |
3116 |
2990 |
33.36 |
|
|
|
| 4 |
A |
3064 |
2939 |
65.34 |
|
|
|
| 5 |
A |
3032 |
2909 |
51.05 |
|
|
|
| 6 |
A |
2973 |
2852 |
116.11 |
|
|
|
| 7 |
A |
1549 |
1486 |
4.17 |
|
|
|
| 8 |
A |
1530 |
1468 |
3.12 |
|
|
|
| 9 |
A |
1515 |
1453 |
1.79 |
|
|
|
| 10 |
A |
1446 |
1387 |
18.63 |
|
|
|
| 11 |
A |
1389 |
1332 |
3.70 |
|
|
|
| 12 |
A |
1354 |
1299 |
0.63 |
|
|
|
| 13 |
A |
1286 |
1234 |
6.02 |
|
|
|
| 14 |
A |
1237 |
1187 |
2.40 |
|
|
|
| 15 |
A |
1232 |
1182 |
3.78 |
|
|
|
| 16 |
A |
1208 |
1159 |
102.36 |
|
|
|
| 17 |
A |
1168 |
1121 |
20.09 |
|
|
|
| 18 |
A |
1136 |
1089 |
96.69 |
|
|
|
| 19 |
A |
1108 |
1063 |
43.20 |
|
|
|
| 20 |
A |
1020 |
978 |
24.65 |
|
|
|
| 21 |
A |
989 |
949 |
54.02 |
|
|
|
| 22 |
A |
964 |
925 |
19.36 |
|
|
|
| 23 |
A |
866 |
831 |
8.04 |
|
|
|
| 24 |
A |
751 |
720 |
1.36 |
|
|
|
| 25 |
A |
706 |
678 |
5.85 |
|
|
|
| 26 |
A |
302 |
290 |
11.52 |
|
|
|
| 27 |
A |
47 |
45 |
3.05 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 20648.6 cm
-1
Scaled (by 0.9594) Zero Point Vibrational Energy (zpe) 19810.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD/cc-pVDZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
1.157 |
-0.089 |
0.175 |
| C2 |
-0.845 |
0.857 |
0.128 |
| C3 |
-1.027 |
-0.676 |
-0.020 |
| O4 |
0.502 |
1.069 |
-0.277 |
| O5 |
0.307 |
-1.179 |
-0.118 |
| H6 |
2.105 |
-0.208 |
-0.371 |
| H7 |
1.345 |
-0.026 |
1.272 |
| H8 |
-0.992 |
1.174 |
1.181 |
| H9 |
-1.572 |
-0.949 |
-0.941 |
| H10 |
-1.553 |
-1.113 |
0.850 |
| H11 |
-1.507 |
1.448 |
-0.524 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
O5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| C1 | | 2.2147 | 2.2700 | 1.4056 | 1.4128 | 1.1008 | 1.1151 | 2.6878 | 3.0712 | 2.9741 | 3.1539 |
C2 | 2.2147 | | 1.5500 | 1.4226 | 2.3517 | 3.1762 | 2.6246 | 1.1098 | 2.2204 | 2.2136 | 1.1010 | C3 | 2.2700 | 1.5500 | | 2.3342 | 1.4293 | 3.1869 | 2.7790 | 2.2058 | 1.1042 | 1.1067 | 2.2347 | O4 | 1.4056 | 1.4226 | 2.3342 | | 2.2622 | 2.0522 | 2.0769 | 2.0902 | 2.9689 | 3.2020 | 2.0589 | O5 | 1.4128 | 2.3517 | 1.4293 | 2.2622 | | 2.0597 | 2.0831 | 2.9849 | 2.0643 | 2.0973 | 3.2182 | H6 | 1.1008 | 3.1762 | 3.1869 | 2.0522 | 2.0597 | | 1.8193 | 3.7297 | 3.7947 | 3.9612 | 3.9769 | H7 | 1.1151 | 2.6246 | 2.7790 | 2.0769 | 2.0831 | 1.8193 | | 2.6291 | 3.7766 | 3.1238 | 3.6791 | H8 | 2.6878 | 1.1098 | 2.2058 | 2.0902 | 2.9849 | 3.7297 | 2.6291 | | 3.0572 | 2.3777 | 1.8017 | H9 | 3.0712 | 2.2204 | 1.1042 | 2.9689 | 2.0643 | 3.7947 | 3.7766 | 3.0572 | | 1.7987 | 2.4341 | H10 | 2.9741 | 2.2136 | 1.1067 | 3.2020 | 2.0973 | 3.9612 | 3.1238 | 2.3777 | 1.7987 | | 2.9065 | H11 | 3.1539 | 1.1010 | 2.2347 | 2.0589 | 3.2182 | 3.9769 | 3.6791 | 1.8017 | 2.4341 | 2.9065 | |
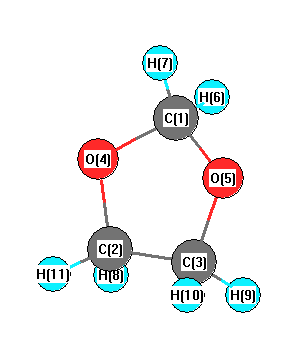 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O4 |
C2 |
103.088 |
|
C1 |
O5 |
C3 |
106.014 |
| C2 |
C3 |
O5 |
104.177 |
|
C2 |
C3 |
H9 |
112.477 |
| C2 |
C3 |
H10 |
111.784 |
|
C3 |
C2 |
O4 |
103.406 |
| C3 |
C2 |
H8 |
110.973 |
|
C3 |
C2 |
H11 |
113.837 |
| O4 |
C1 |
O5 |
106.772 |
|
O4 |
C1 |
H6 |
109.327 |
| O4 |
C1 |
H7 |
110.436 |
|
O4 |
C2 |
H8 |
110.648 |
| O4 |
C2 |
H11 |
108.676 |
|
O5 |
C1 |
H6 |
109.427 |
| O5 |
C1 |
H7 |
110.432 |
|
O5 |
C3 |
H9 |
108.458 |
| O5 |
C3 |
H10 |
110.948 |
|
H6 |
C1 |
H7 |
110.371 |
| H8 |
C2 |
H11 |
109.162 |
|
H9 |
C3 |
H10 |
108.898 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
