return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H5COOH (benzoic acid)

using model chemistry: QCISD/cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at QCISD/cc-pVDZ
 hartrees
Energy at 0K-419.671902
Energy at 298.15K-419.678982
HF Energy-418.358756
Nuclear repulsion energy402.458725
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3799 3645 87.29      
2 A' 3248 3116 2.49      
3 A' 3243 3111 4.45      
4 A' 3228 3096 15.00      
5 A' 3216 3086 11.34      
6 A' 3203 3073 0.26      
7 A' 1846 1772 280.74      
8 A' 1683 1615 12.97      
9 A' 1662 1594 4.05      
10 A' 1542 1479 6.41      
11 A' 1491 1431 19.51      
12 A' 1419 1361 118.01      
13 A' 1340 1286 6.70      
14 A' 1322 1268 3.94      
15 A' 1243 1193 207.39      
16 A' 1194 1146 26.43      
17 A' 1171 1123 0.48      
18 A' 1134 1088 55.97      
19 A' 1100 1055 22.88      
20 A' 1051 1008 7.64      
21 A' 1013 972 0.58      
22 A' 781 749 14.99      
23 A' 639 613 49.85      
24 A' 623 597 0.15      
25 A' 497 477 4.53      
26 A' 384 369 5.23      
27 A' 218 209 1.20      
28 A" 987 947 0.03      
29 A" 982 942 0.04      
30 A" 947 909 0.49      
31 A" 871 836 0.04      
32 A" 818 785 1.17      
33 A" 730 700 126.95      
34 A" 654 628 3.38      
35 A" 592 568 59.59      
36 A" 428 410 8.36      
37 A" 408 392 0.59      
38 A" 161 154 0.86      
39 A" 68 66 0.79      

Unscaled Zero Point Vibrational Energy (zpe) 25468.0 cm-1
Scaled (by 0.9594) Zero Point Vibrational Energy (zpe) 24434.0 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/cc-pVDZ
ABC
0.12775 0.04042 0.03070

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/cc-pVDZ

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.224 0.000
C2 1.283 -0.356 0.000
C3 1.420 -1.752 0.000
C4 0.276 -2.570 0.000
C5 -1.006 -1.991 0.000
C6 -1.147 -0.594 0.000
C7 -0.090 1.721 0.000
O8 0.864 2.470 0.000
O9 -1.370 2.168 0.000
H10 2.159 0.301 0.000
H11 2.419 -2.204 0.000
H12 0.382 -3.661 0.000
H13 -1.896 -2.629 0.000
H14 -2.140 -0.136 0.000
H15 -1.297 3.137 0.000

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 C7 O8 O9 H10 H11 H12 H13 H14 H15
C11.40842.43382.80822.43271.40851.49922.40642.37822.16023.42703.90423.42572.17003.1880
C21.40841.40282.43292.81292.44192.48992.85693.66231.09432.16863.42583.90853.43054.3422
C32.43381.40281.40722.43782.81673.78724.25864.81202.18151.09562.17313.43083.91035.5931
C42.80822.43291.40721.40622.43564.30665.07475.01623.43352.17411.09602.17293.42995.9197
C52.43272.81292.43781.40621.40463.82274.83704.17493.90703.43082.17171.09562.17475.1359
C61.40852.44192.81672.43561.40462.54383.66472.77053.42453.91233.42782.16881.09363.7331
C71.49922.48993.78724.30663.82272.54381.21381.35572.66004.65795.40264.70932.76521.8600
O82.40642.85694.25865.07474.83703.66471.21382.25482.52634.92546.15025.79813.97682.2614
O92.37823.66234.81205.01624.17492.77051.35572.25483.99275.78516.08704.82542.42880.9714
H102.16021.09432.18153.43353.90703.42452.66002.52633.99272.51774.34185.00264.32094.4702
H113.42702.16861.09562.17413.43083.91234.65794.92545.78512.51772.50424.33605.00586.5055
H123.90423.42582.17311.09602.17173.42785.40266.15026.08704.34182.50422.50194.33517.0022
H133.42573.90853.43082.17291.09562.16884.70935.79814.82545.00264.33602.50192.50495.7964
H142.17003.43053.91033.42992.17471.09362.76523.97682.42884.32095.00584.33512.50493.3792
H153.18804.34225.59315.91975.13593.73311.86002.26140.97144.47026.50557.00225.79643.3792

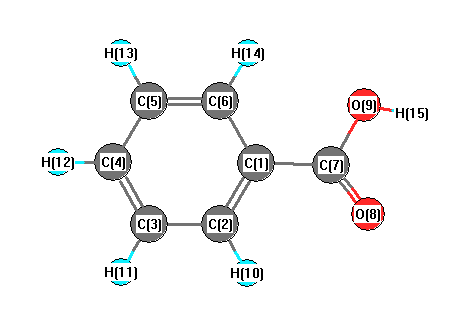
picture of benzoic acid state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 119.933 C1 C2 H10 118.805
C1 C6 C5 119.711 C1 C6 H14 119.763
C1 C7 O8 124.668 C1 C7 O9 112.729
C2 C1 C6 120.189 C2 C1 C7 117.785
C2 C3 C4 119.950 C2 C3 H11 119.947
C3 C2 H10 121.262 C3 C4 C5 120.107
C3 C4 H12 119.969 C4 C3 H11 120.103
C4 C5 C6 120.109 C4 C5 H13 120.072
C5 C4 H12 119.924 C5 C6 H14 120.525
C6 C1 C7 122.026 C6 C5 H13 119.819
C7 O9 H15 104.922 O8 C7 O9 122.603
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability