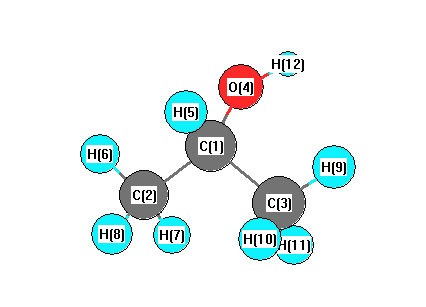
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3657 |
3635 |
7.37 |
|
|
|
| 2 |
A |
3073 |
3055 |
20.85 |
|
|
|
| 3 |
A |
3062 |
3044 |
33.24 |
|
|
|
| 4 |
A |
3054 |
3036 |
10.99 |
|
|
|
| 5 |
A |
3036 |
3019 |
30.46 |
|
|
|
| 6 |
A |
2977 |
2960 |
14.15 |
|
|
|
| 7 |
A |
2958 |
2941 |
23.03 |
|
|
|
| 8 |
A |
2865 |
2849 |
72.62 |
|
|
|
| 9 |
A |
1432 |
1424 |
4.61 |
|
|
|
| 10 |
A |
1422 |
1414 |
6.62 |
|
|
|
| 11 |
A |
1409 |
1401 |
5.09 |
|
|
|
| 12 |
A |
1402 |
1394 |
0.77 |
|
|
|
| 13 |
A |
1369 |
1361 |
21.16 |
|
|
|
| 14 |
A |
1341 |
1333 |
16.64 |
|
|
|
| 15 |
A |
1320 |
1312 |
2.44 |
|
|
|
| 16 |
A |
1308 |
1300 |
11.02 |
|
|
|
| 17 |
A |
1238 |
1230 |
47.42 |
|
|
|
| 18 |
A |
1142 |
1136 |
35.01 |
|
|
|
| 19 |
A |
1117 |
1110 |
23.91 |
|
|
|
| 20 |
A |
1063 |
1057 |
20.51 |
|
|
|
| 21 |
A |
939 |
934 |
32.79 |
|
|
|
| 22 |
A |
911 |
906 |
3.84 |
|
|
|
| 23 |
A |
888 |
883 |
0.23 |
|
|
|
| 24 |
A |
812 |
807 |
4.17 |
|
|
|
| 25 |
A |
466 |
463 |
5.27 |
|
|
|
| 26 |
A |
399 |
397 |
11.16 |
|
|
|
| 27 |
A |
348 |
346 |
3.87 |
|
|
|
| 28 |
A |
310 |
308 |
89.09 |
|
|
|
| 29 |
A |
262 |
260 |
3.29 |
|
|
|
| 30 |
A |
219 |
217 |
2.94 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 22898.9 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 22766.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.022 |
|
|
|
| 2 |
C |
-0.033 |
|
|
|
| 3 |
C |
-0.015 |
|
|
|
| 4 |
O |
-0.235 |
|
|
|
| 5 |
H |
-0.001 |
|
|
|
| 6 |
H |
0.018 |
|
|
|
| 7 |
H |
0.035 |
|
|
|
| 8 |
H |
0.027 |
|
|
|
| 9 |
H |
0.035 |
|
|
|
| 10 |
H |
0.026 |
|
|
|
| 11 |
H |
0.033 |
|
|
|
| 12 |
H |
0.133 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.082 |
-0.493 |
0.705 |
1.382 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-25.423 |
-2.518 |
-0.235 |
| y |
-2.518 |
-26.585 |
0.870 |
| z |
-0.235 |
0.870 |
-26.771 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.255 |
-2.518 |
-0.235 |
| y |
-2.518 |
-0.488 |
0.870 |
| z |
-0.235 |
0.870 |
-0.767 |
|
| Polar |
|---|
| 3z2-r2 | -1.534 |
|---|
| x2-y2 | 1.161 |
|---|
| xy | -2.518 |
|---|
| xz | -0.235 |
|---|
| yz | 0.870 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.564 |
-0.156 |
-0.060 |
| y |
-0.156 |
6.107 |
-0.005 |
| z |
-0.060 |
-0.005 |
5.515 |
<r2> (average value of r
2) Å
2
| <r2> |
89.904 |
| (<r2>)1/2 |
9.482 |