return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C2H6O2S (Dimethyl sulfone)

using model chemistry: MP4=FULL/cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at MP4=FULL/cc-pVDZ
 hartrees
Energy at 0K-627.295408
Energy at 298.15K 
HF Energy-626.398136
Nuclear repulsion energy269.469515
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP4=FULL/cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A1 3197 3085        
2 A1 3075 2968        
3 A1 1453 1402        
4 A1 1333 1287        
5 A1 1114 1075        
6 A1 989 955        
7 A1 667 644        
8 A1 457 441        
9 A1 262 253        
10 A2 3199 3087        
11 A2 1441 1390        
12 A2 907 876        
13 A2 275 265        
14 A2 174 168        
15 B1 3202 3090        
16 B1 1456 1406        
17 B1 1305 1260        
18 B1 969 935        
19 B1 339 327        
20 B1 210 203        
21 B2 3196 3084        
22 B2 3074 2966        
23 B2 1442 1392        
24 B2 1315 1269        
25 B2 928 896        
26 B2 731 705        
27 B2 426 411        

Unscaled Zero Point Vibrational Energy (zpe) 18567.7 cm-1
Scaled (by 0.9651) Zero Point Vibrational Energy (zpe) 17919.7 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP4=FULL/cc-pVDZ
ABC
0.14566 0.13670 0.13544

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP4=FULL/cc-pVDZ

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
S1 0.000 0.000 0.198
O2 -1.298 0.000 0.921
O3 1.298 0.000 0.921
C4 0.000 1.409 -0.929
C5 0.000 -1.409 -0.929
H6 0.000 2.301 -0.282
H7 0.000 -2.301 -0.282
H8 0.913 1.385 -1.546
H9 -0.913 1.385 -1.546
H10 -0.913 -1.385 -1.546
H11 0.913 -1.385 -1.546

Atom - Atom Distances (Å)
  S1 O2 O3 C4 C5 H6 H7 H8 H9 H10 H11
S11.48611.48611.80421.80422.35092.35092.40692.40692.40692.4069
O21.48612.59692.66342.66342.90342.90343.59082.85532.85533.5908
O31.48612.59692.66342.66342.90342.90342.85533.59083.59082.8553
C41.80422.66342.66342.81721.10253.76591.10191.10193.00293.0029
C51.80422.66342.66342.81723.76591.10253.00293.00291.10191.1019
H62.35092.90342.90341.10253.76594.60261.80821.80824.00234.0023
H72.35092.90342.90343.76591.10254.60264.00234.00231.80821.8082
H82.40693.59082.85531.10193.00291.80824.00231.82573.31752.7699
H92.40692.85533.59081.10193.00291.80824.00231.82572.76993.3175
H102.40692.85533.59083.00291.10194.00231.80823.31752.76991.8257
H112.40693.59082.85533.00291.10194.00231.80822.76993.31751.8257

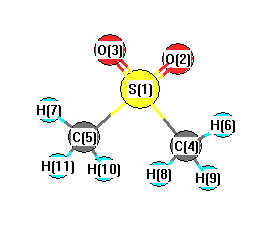
picture of Dimethyl sulfone state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
S1 C4 H6 105.394 S1 C4 H8 109.448
S1 C4 H9 109.448 S1 C5 H7 105.394
S1 C5 H10 109.448 S1 C5 H11 109.448
O2 S1 O3 121.785 O2 S1 C4 107.696
O2 S1 C5 107.696 O3 S1 C4 107.696
O3 S1 C5 107.696 C4 S1 C5 102.653
H6 C4 H8 110.228 H6 C4 H9 110.228
H7 C5 H10 110.228 H7 C5 H11 110.228
H8 C4 H9 111.886 H10 C5 H11 111.886
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability