Jump to
S1C2
Energy calculated at QCISD/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -498.771162 |
| Energy at 298.15K | |
| HF Energy | -498.486168 |
| Nuclear repulsion energy | 44.905211 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3190 |
3061 |
7.90 |
|
|
|
| 2 |
A' |
1423 |
1366 |
14.05 |
|
|
|
| 3 |
A' |
853 |
818 |
33.70 |
|
|
|
| 4 |
A' |
343 |
330 |
57.70 |
|
|
|
| 5 |
A" |
3339 |
3203 |
0.29 |
|
|
|
| 6 |
A" |
1001 |
960 |
0.01 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 5074.4 cm
-1
Scaled (by 0.9594) Zero Point Vibrational Energy (zpe) 4868.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.012 |
1.129 |
0.000 |
| Cl2 |
-0.012 |
-0.590 |
0.000 |
| H3 |
0.139 |
1.628 |
0.960 |
| H4 |
0.139 |
1.628 |
-0.960 |
Atom - Atom Distances (Å)
| |
C1 |
Cl2 |
H3 |
H4 |
| C1 | | 1.7196 | 1.0925 | 1.0925 |
Cl2 | 1.7196 | | 2.4222 | 2.4222 | H3 | 1.0925 | 2.4222 | | 1.9198 | H4 | 1.0925 | 2.4222 | 1.9198 | |
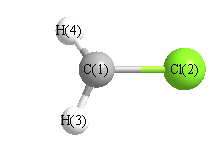 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| Br2 |
C1 |
H3 |
117.183 |
|
Br2 |
C1 |
H4 |
117.183 |
| H3 |
C1 |
H4 |
122.962 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at QCISD/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -498.771089 |
| Energy at 298.15K | |
| HF Energy | -498.485914 |
| Nuclear repulsion energy | 44.934917 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3200 |
3070 |
6.09 |
|
|
|
| 2 |
A1 |
1422 |
1365 |
14.70 |
|
|
|
| 3 |
A1 |
856 |
821 |
32.03 |
|
|
|
| 4 |
B1 |
247i |
237i |
69.01 |
|
|
|
| 5 |
B2 |
3353 |
3217 |
0.02 |
|
|
|
| 6 |
B2 |
993 |
953 |
0.01 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 4788.5 cm
-1
Scaled (by 0.9594) Zero Point Vibrational Energy (zpe) 4594.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD/cc-pVDZ
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
-1.127 |
| Cl2 |
0.000 |
0.000 |
0.590 |
| H3 |
0.000 |
0.964 |
-1.638 |
| H4 |
0.000 |
-0.964 |
-1.638 |
Atom - Atom Distances (Å)
| |
C1 |
Cl2 |
H3 |
H4 |
| C1 | | 1.7173 | 1.0914 | 1.0914 |
Cl2 | 1.7173 | | 2.4279 | 2.4279 | H3 | 1.0914 | 2.4279 | | 1.9289 | H4 | 1.0914 | 2.4279 | 1.9289 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| Br2 |
C1 |
H3 |
117.909 |
|
Br2 |
C1 |
H4 |
117.909 |
| H3 |
C1 |
H4 |
124.181 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
