Jump to
S1C2
Energy calculated at PBEPBEultrafine/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -499.199962 |
| Energy at 298.15K | |
| HF Energy | -499.199962 |
| Nuclear repulsion energy | 44.978168 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBEPBEultrafine/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3097 |
3079 |
4.80 |
120.79 |
0.12 |
0.22 |
| 2 |
A' |
1332 |
1325 |
8.53 |
3.45 |
0.62 |
0.76 |
| 3 |
A' |
838 |
833 |
34.22 |
8.27 |
0.29 |
0.45 |
| 4 |
A' |
119 |
118 |
72.99 |
0.49 |
0.42 |
0.59 |
| 5 |
A" |
3257 |
3238 |
0.00 |
56.53 |
0.75 |
0.86 |
| 6 |
A" |
950 |
944 |
0.23 |
4.94 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 4796.4 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 4768.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBEPBEultrafine/cc-pVDZ
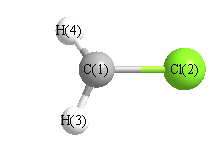
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.004 |
1.125 |
0.000 |
| Cl2 |
-0.004 |
-0.589 |
0.000 |
| H3 |
0.049 |
1.632 |
0.971 |
| H4 |
0.049 |
1.632 |
-0.971 |
Atom - Atom Distances (Å)
| |
C1 |
Cl2 |
H3 |
H4 |
| C1 | | 1.7139 | 1.0966 | 1.0966 |
Cl2 | 1.7139 | | 2.4243 | 2.4243 | H3 | 1.0966 | 2.4243 | | 1.9420 | H4 | 1.0966 | 2.4243 | 1.9420 | |
 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| Br2 |
C1 |
H3 |
117.530 |
|
Br2 |
C1 |
H4 |
117.530 |
| H3 |
C1 |
H4 |
124.614 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.144 |
|
|
|
| 2 |
Cl |
-0.034 |
|
|
|
| 3 |
H |
0.089 |
|
|
|
| 4 |
H |
0.089 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.093 |
0.941 |
0.000 |
0.945 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-20.211 |
0.159 |
0.000 |
| y |
0.159 |
-17.192 |
0.000 |
| z |
0.000 |
0.000 |
-18.276 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.477 |
0.159 |
0.000 |
| y |
0.159 |
2.052 |
0.000 |
| z |
0.000 |
0.000 |
0.426 |
|
| Polar |
|---|
| 3z2-r2 | 0.851 |
|---|
| x2-y2 | -3.019 |
|---|
| xy | 0.159 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
1.658 |
0.025 |
0.000 |
| y |
0.025 |
4.523 |
0.000 |
| z |
0.000 |
0.000 |
2.404 |
<r2> (average value of r
2) Å
2
| <r2> |
32.298 |
| (<r2>)1/2 |
5.683 |
Jump to
S1C1
Energy calculated at PBEPBEultrafine/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -499.199962 |
| Energy at 298.15K | |
| HF Energy | -499.199962 |
| Nuclear repulsion energy | 44.982791 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBEPBEultrafine/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3098 |
3081 |
4.61 |
120.49 |
0.12 |
0.22 |
| 2 |
A1 |
1332 |
1325 |
8.54 |
|
|
|
| 3 |
A1 |
838 |
833 |
34.04 |
|
|
|
| 4 |
B1 |
46i |
46i |
74.25 |
|
|
|
| 5 |
B2 |
3259 |
3240 |
0.00 |
|
|
|
| 6 |
B2 |
949 |
944 |
0.24 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 4715.4 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 4688.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBEPBEultrafine/cc-pVDZ
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
-1.125 |
| Cl2 |
0.000 |
0.000 |
0.589 |
| H3 |
0.000 |
0.972 |
-1.633 |
| H4 |
0.000 |
-0.972 |
-1.633 |
Atom - Atom Distances (Å)
| |
C1 |
Cl2 |
H3 |
H4 |
| C1 | | 1.7135 | 1.0965 | 1.0965 |
Cl2 | 1.7135 | | 2.4250 | 2.4250 | H3 | 1.0965 | 2.4250 | | 1.9430 | H4 | 1.0965 | 2.4250 | 1.9430 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| Br2 |
C1 |
H3 |
117.621 |
|
Br2 |
C1 |
H4 |
117.621 |
| H3 |
C1 |
H4 |
124.758 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.144 |
|
|
|
| 2 |
Cl |
-0.034 |
|
|
|
| 3 |
H |
0.089 |
|
|
|
| 4 |
H |
0.089 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-0.942 |
0.942 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-20.218 |
0.000 |
0.000 |
| y |
0.000 |
-18.271 |
0.000 |
| z |
0.000 |
0.000 |
-17.183 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.491 |
0.000 |
0.000 |
| y |
0.000 |
0.429 |
0.000 |
| z |
0.000 |
0.000 |
2.061 |
|
| Polar |
|---|
| 3z2-r2 | 4.122 |
|---|
| x2-y2 | -1.947 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
1.655 |
0.000 |
0.000 |
| y |
0.000 |
2.403 |
0.000 |
| z |
0.000 |
0.000 |
4.522 |
<r2> (average value of r
2) Å
2
| <r2> |
32.296 |
| (<r2>)1/2 |
5.683 |
