Jump to
S1C2
Energy calculated at ROHF/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -498.482138 |
| Energy at 298.15K | |
| HF Energy | -498.482138 |
| Nuclear repulsion energy | 44.896376 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at ROHF/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3311 |
2849 |
13.60 |
|
|
|
| 2 |
A' |
1503 |
1294 |
18.29 |
|
|
|
| 3 |
A' |
857 |
737 |
48.72 |
|
|
|
| 4 |
A' |
689 |
593 |
36.43 |
|
|
|
| 5 |
A" |
3464 |
2981 |
0.59 |
|
|
|
| 6 |
A" |
1071 |
922 |
0.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 5446.6 cm
-1
Scaled (by 0.8607) Zero Point Vibrational Energy (zpe) 4687.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at ROHF/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.021 |
1.138 |
0.000 |
| Cl2 |
-0.021 |
-0.590 |
0.000 |
| H3 |
0.241 |
1.600 |
0.940 |
| H4 |
0.241 |
1.600 |
-0.940 |
Atom - Atom Distances (Å)
| |
C1 |
Cl2 |
H3 |
H4 |
| C1 | | 1.7284 | 1.0796 | 1.0796 |
Cl2 | 1.7284 | | 2.3973 | 2.3973 | H3 | 1.0796 | 2.3973 | | 1.8806 | H4 | 1.0796 | 2.3973 | 1.8806 | |
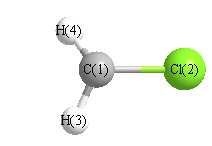 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| Br2 |
C1 |
H3 |
115.285 |
|
Br2 |
C1 |
H4 |
115.285 |
| H3 |
C1 |
H4 |
121.137 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at ROHF/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.068 |
|
|
|
| 2 |
Cl |
-0.128 |
|
|
|
| 3 |
H |
0.098 |
|
|
|
| 4 |
H |
0.098 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.416 |
1.565 |
0.000 |
1.619 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-20.259 |
0.692 |
-0.000 |
| y |
0.692 |
-17.306 |
0.000 |
| z |
-0.000 |
0.000 |
-18.595 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.309 |
0.692 |
-0.000 |
| y |
0.692 |
2.121 |
0.000 |
| z |
-0.000 |
0.000 |
0.188 |
|
| Polar |
|---|
| 3z2-r2 | 0.376 |
|---|
| x2-y2 | -2.953 |
|---|
| xy | 0.692 |
|---|
| xz | -0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
32.399 |
| (<r2>)1/2 |
5.692 |
Jump to
S1C1
Energy calculated at ROHF/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -498.481087 |
| Energy at 298.15K | |
| HF Energy | -498.481087 |
| Nuclear repulsion energy | 45.028929 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at ROHF/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3345 |
2879 |
6.84 |
|
|
|
| 2 |
A1 |
1499 |
1290 |
20.88 |
|
|
|
| 3 |
A1 |
867 |
746 |
42.98 |
|
|
|
| 4 |
B1 |
550i |
474i |
64.57 |
|
|
|
| 5 |
B2 |
3513 |
3023 |
0.39 |
|
|
|
| 6 |
B2 |
1047 |
901 |
0.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 4860.1 cm
-1
Scaled (by 0.8607) Zero Point Vibrational Energy (zpe) 4183.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at ROHF/cc-pVDZ
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
-1.129 |
| Cl2 |
0.000 |
0.000 |
0.590 |
| H3 |
0.000 |
0.954 |
-1.627 |
| H4 |
0.000 |
-0.954 |
-1.627 |
Atom - Atom Distances (Å)
| |
C1 |
Cl2 |
H3 |
H4 |
| C1 | | 1.7193 | 1.0760 | 1.0760 |
Cl2 | 1.7193 | | 2.4133 | 2.4133 | H3 | 1.0760 | 2.4133 | | 1.9082 | H4 | 1.0760 | 2.4133 | 1.9082 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| Br2 |
C1 |
H3 |
117.536 |
|
Br2 |
C1 |
H4 |
117.536 |
| H3 |
C1 |
H4 |
124.927 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at ROHF/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.087 |
|
|
|
| 2 |
Cl |
-0.118 |
|
|
|
| 3 |
H |
0.102 |
|
|
|
| 4 |
H |
0.102 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-1.556 |
1.556 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-20.429 |
0.000 |
0.000 |
| y |
0.000 |
-18.443 |
-0.001 |
| z |
0.000 |
-0.001 |
-17.111 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.652 |
0.000 |
0.000 |
| y |
0.000 |
0.327 |
-0.001 |
| z |
0.000 |
-0.001 |
2.325 |
|
| Polar |
|---|
| 3z2-r2 | 4.650 |
|---|
| x2-y2 | -1.986 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | -0.001 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
32.338 |
| (<r2>)1/2 |
5.687 |
