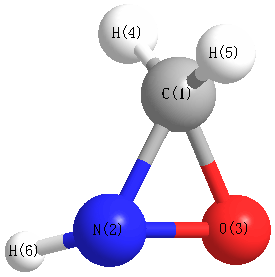
Vibrational Frequencies calculated at BLYP/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3182 |
3187 |
12.48 |
|
|
|
| 2 |
A |
3045 |
3049 |
41.53 |
|
|
|
| 3 |
A |
2955 |
2960 |
38.28 |
|
|
|
| 4 |
A |
1469 |
1472 |
3.24 |
|
|
|
| 5 |
A |
1321 |
1323 |
18.43 |
|
|
|
| 6 |
A |
1207 |
1209 |
31.33 |
|
|
|
| 7 |
A |
1194 |
1196 |
4.87 |
|
|
|
| 8 |
A |
1145 |
1147 |
1.65 |
|
|
|
| 9 |
A |
1030 |
1032 |
9.69 |
|
|
|
| 10 |
A |
915 |
916 |
13.08 |
|
|
|
| 11 |
A |
846 |
847 |
22.06 |
|
|
|
| 12 |
A |
703 |
704 |
5.33 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 9505.7 cm
-1
Scaled (by 1.0016) Zero Point Vibrational Energy (zpe) 9520.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at BLYP/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.184 |
|
|
|
| 2 |
N |
-0.094 |
|
|
|
| 3 |
O |
-0.237 |
|
|
|
| 4 |
H |
0.016 |
|
|
|
| 5 |
H |
0.026 |
|
|
|
| 6 |
H |
0.104 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.947 |
-1.544 |
1.346 |
2.257 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-16.504 |
-0.287 |
-1.840 |
| y |
-0.287 |
-18.654 |
-0.882 |
| z |
-1.840 |
-0.882 |
-17.108 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.376 |
-0.287 |
-1.840 |
| y |
-0.287 |
-1.848 |
-0.882 |
| z |
-1.840 |
-0.882 |
0.472 |
|
| Polar |
|---|
| 3z2-r2 | 0.943 |
|---|
| x2-y2 | 2.150 |
|---|
| xy | -0.287 |
|---|
| xz | -1.840 |
|---|
| yz | -0.882 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.706 |
-0.088 |
-0.183 |
| y |
-0.088 |
2.850 |
-0.119 |
| z |
-0.183 |
-0.119 |
2.824 |
<r2> (average value of r
2) Å
2
| <r2> |
34.048 |
| (<r2>)1/2 |
5.835 |