Jump to
S1C2
Energy calculated at CCD/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -749.654064 |
| Energy at 298.15K | |
| HF Energy | -748.030397 |
| Nuclear repulsion energy | 560.460358 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
1913 |
1831 |
0.00 |
|
|
|
| 2 |
Ag |
1490 |
1425 |
0.00 |
|
|
|
| 3 |
Ag |
1385 |
1325 |
0.00 |
|
|
|
| 4 |
Ag |
1216 |
1164 |
0.00 |
|
|
|
| 5 |
Ag |
727 |
696 |
0.00 |
|
|
|
| 6 |
Ag |
595 |
569 |
0.00 |
|
|
|
| 7 |
Ag |
383 |
366 |
0.00 |
|
|
|
| 8 |
Ag |
355 |
340 |
0.00 |
|
|
|
| 9 |
Ag |
223 |
214 |
0.00 |
|
|
|
| 10 |
Au |
585 |
560 |
5.37 |
|
|
|
| 11 |
Au |
355 |
340 |
5.91 |
|
|
|
| 12 |
Au |
113 |
108 |
0.10 |
|
|
|
| 13 |
Au |
17i |
16i |
0.10 |
|
|
|
| 14 |
Bg |
655 |
627 |
0.00 |
|
|
|
| 15 |
Bg |
492 |
470 |
0.00 |
|
|
|
| 16 |
Bg |
185 |
177 |
0.00 |
|
|
|
| 17 |
Bu |
1850 |
1770 |
307.76 |
|
|
|
| 18 |
Bu |
1409 |
1348 |
401.65 |
|
|
|
| 19 |
Bu |
1256 |
1202 |
309.52 |
|
|
|
| 20 |
Bu |
977 |
935 |
269.60 |
|
|
|
| 21 |
Bu |
632 |
605 |
6.79 |
|
|
|
| 22 |
Bu |
504 |
482 |
4.85 |
|
|
|
| 23 |
Bu |
288 |
276 |
6.66 |
|
|
|
| 24 |
Bu |
139 |
133 |
0.95 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 8855.5 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 8472.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCD/cc-pVDZ
Point Group is C2h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.158 |
1.864 |
0.000 |
| C2 |
0.474 |
0.559 |
0.000 |
| C3 |
-0.474 |
-0.559 |
0.000 |
| C4 |
-0.158 |
-1.864 |
0.000 |
| F5 |
1.066 |
2.817 |
0.000 |
| F6 |
-1.066 |
2.345 |
0.000 |
| F7 |
1.777 |
0.226 |
0.000 |
| F8 |
-1.777 |
-0.226 |
0.000 |
| F9 |
1.066 |
-2.345 |
0.000 |
| F10 |
-1.066 |
-2.817 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
F5 |
F6 |
F7 |
F8 |
F9 |
F10 |
| C1 | | 1.3421 | 2.5042 | 3.7408 | 1.3168 | 1.3151 | 2.3030 | 2.8480 | 4.3051 | 4.8382 |
C2 | 1.3421 | | 1.4667 | 2.5042 | 2.3340 | 2.3578 | 1.3449 | 2.3844 | 2.9637 | 3.7113 | C3 | 2.5042 | 1.4667 | | 1.3421 | 3.7113 | 2.9637 | 2.3844 | 1.3449 | 2.3578 | 2.3340 | C4 | 3.7408 | 2.5042 | 1.3421 | | 4.8382 | 4.3051 | 2.8480 | 2.3030 | 1.3151 | 1.3168 | F5 | 1.3168 | 2.3340 | 3.7113 | 4.8382 | | 2.1840 | 2.6869 | 4.1647 | 5.1616 | 6.0242 | F6 | 1.3151 | 2.3578 | 2.9637 | 4.3051 | 2.1840 | | 3.5457 | 2.6670 | 5.1511 | 5.1616 | F7 | 2.3030 | 1.3449 | 2.3844 | 2.8480 | 2.6869 | 3.5457 | | 3.5828 | 2.6670 | 4.1647 | F8 | 2.8480 | 2.3844 | 1.3449 | 2.3030 | 4.1647 | 2.6670 | 3.5828 | | 3.5457 | 2.6869 | F9 | 4.3051 | 2.9637 | 2.3578 | 1.3151 | 5.1616 | 5.1511 | 2.6670 | 3.5457 | | 2.1840 | F10 | 4.8382 | 3.7113 | 2.3340 | 1.3168 | 6.0242 | 5.1616 | 4.1647 | 2.6869 | 2.1840 | |
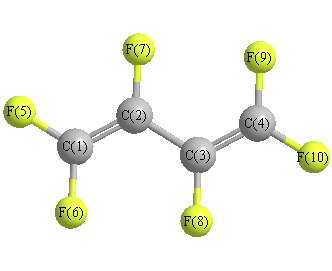 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
126.081 |
|
C1 |
C2 |
F7 |
117.987 |
| C2 |
C1 |
F5 |
122.757 |
|
C2 |
C1 |
F6 |
125.077 |
| C2 |
C3 |
C4 |
126.081 |
|
C2 |
C3 |
F8 |
115.933 |
| C3 |
C2 |
F7 |
115.933 |
|
C3 |
C4 |
F9 |
125.077 |
| C3 |
C4 |
F10 |
122.757 |
|
C4 |
C3 |
F8 |
117.987 |
| F5 |
C1 |
F6 |
112.165 |
|
F9 |
C4 |
F10 |
112.165 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CCD/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -749.655091 |
| Energy at 298.15K | -749.656347 |
| HF Energy | -748.031382 |
| Nuclear repulsion energy | 564.727824 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
1908 |
1825 |
60.77 |
|
|
|
| 2 |
A |
1464 |
1401 |
10.65 |
|
|
|
| 3 |
A |
1402 |
1341 |
179.63 |
|
|
|
| 4 |
A |
1177 |
1126 |
281.04 |
|
|
|
| 5 |
A |
727 |
695 |
0.13 |
|
|
|
| 6 |
A |
673 |
644 |
1.59 |
|
|
|
| 7 |
A |
546 |
523 |
0.03 |
|
|
|
| 8 |
A |
482 |
461 |
0.63 |
|
|
|
| 9 |
A |
385 |
368 |
1.11 |
|
|
|
| 10 |
A |
259 |
248 |
0.19 |
|
|
|
| 11 |
A |
190 |
182 |
0.35 |
|
|
|
| 12 |
A |
99 |
95 |
0.32 |
|
|
|
| 13 |
A |
48 |
46 |
0.00 |
|
|
|
| 14 |
B |
1883 |
1801 |
195.39 |
|
|
|
| 15 |
B |
1405 |
1345 |
279.76 |
|
|
|
| 16 |
B |
1253 |
1199 |
113.59 |
|
|
|
| 17 |
B |
998 |
955 |
185.39 |
|
|
|
| 18 |
B |
638 |
610 |
6.13 |
|
|
|
| 19 |
B |
632 |
605 |
4.38 |
|
|
|
| 20 |
B |
572 |
547 |
4.13 |
|
|
|
| 21 |
B |
425 |
406 |
4.82 |
|
|
|
| 22 |
B |
299 |
286 |
4.10 |
|
|
|
| 23 |
B |
211 |
202 |
4.48 |
|
|
|
| 24 |
B |
111 |
106 |
0.36 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 8892.4 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 8507.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCD/cc-pVDZ
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.093 |
1.569 |
-0.375 |
| C2 |
0.274 |
0.679 |
0.608 |
| C3 |
-0.274 |
-0.679 |
0.608 |
| C4 |
-0.093 |
-1.569 |
-0.375 |
| F5 |
0.619 |
2.773 |
-0.388 |
| F6 |
-0.619 |
1.325 |
-1.456 |
| F7 |
0.995 |
1.034 |
1.689 |
| F8 |
-0.995 |
-1.034 |
1.689 |
| F9 |
0.619 |
-1.325 |
-1.456 |
| F10 |
-0.619 |
-2.773 |
-0.388 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
F5 |
F6 |
F7 |
F8 |
F9 |
F10 |
| C1 | | 1.3384 | 2.4810 | 3.1438 | 1.3134 | 1.3173 | 2.3155 | 3.4958 | 3.1340 | 4.3997 |
C2 | 1.3384 | | 1.4645 | 2.4810 | 2.3437 | 2.3400 | 1.3473 | 2.3903 | 2.8977 | 3.7016 | C3 | 2.4810 | 1.4645 | | 1.3384 | 3.7016 | 2.8977 | 2.3903 | 1.3473 | 2.3400 | 2.3437 | C4 | 3.1438 | 2.4810 | 1.3384 | | 4.3997 | 3.1340 | 3.4958 | 2.3155 | 1.3173 | 1.3134 | F5 | 1.3134 | 2.3437 | 3.7016 | 4.3997 | | 2.1840 | 2.7345 | 4.6268 | 4.2346 | 5.6816 | F6 | 1.3173 | 2.3400 | 2.8977 | 3.1340 | 2.1840 | | 3.5474 | 3.9498 | 2.9249 | 4.2346 | F7 | 2.3155 | 1.3473 | 2.3903 | 3.4958 | 2.7345 | 3.5474 | | 2.8698 | 3.9498 | 4.6268 | F8 | 3.4958 | 2.3903 | 1.3473 | 2.3155 | 4.6268 | 3.9498 | 2.8698 | | 3.5474 | 2.7345 | F9 | 3.1340 | 2.8977 | 2.3400 | 1.3173 | 4.2346 | 2.9249 | 3.9498 | 3.5474 | | 2.1840 | F10 | 4.3997 | 3.7016 | 2.3437 | 1.3134 | 5.6816 | 4.2346 | 4.6268 | 2.7345 | 2.1840 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
124.486 |
|
C1 |
C2 |
F7 |
119.124 |
| C2 |
C1 |
F5 |
124.208 |
|
C2 |
C1 |
F6 |
123.551 |
| C2 |
C3 |
C4 |
124.486 |
|
C2 |
C3 |
F8 |
116.388 |
| C3 |
C2 |
F7 |
116.388 |
|
C3 |
C4 |
F9 |
123.551 |
| C3 |
C4 |
F10 |
124.208 |
|
C4 |
C3 |
F8 |
119.124 |
| F5 |
C1 |
F6 |
112.231 |
|
F9 |
C4 |
F10 |
112.231 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
