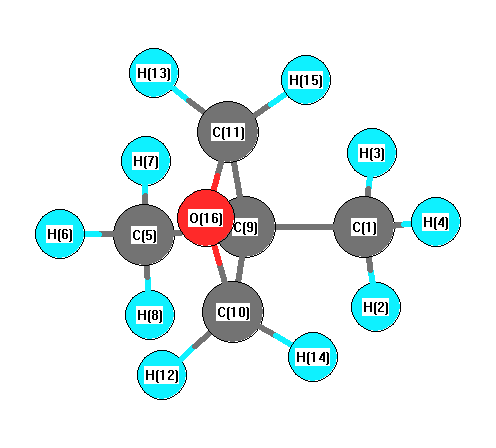
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3050 |
3033 |
27.41 |
|
|
|
| 2 |
A' |
3049 |
3031 |
20.36 |
|
|
|
| 3 |
A' |
2970 |
2952 |
88.11 |
|
|
|
| 4 |
A' |
2963 |
2946 |
30.34 |
|
|
|
| 5 |
A' |
2958 |
2941 |
15.38 |
|
|
|
| 6 |
A' |
2918 |
2901 |
32.87 |
|
|
|
| 7 |
A' |
1457 |
1449 |
1.49 |
|
|
|
| 8 |
A' |
1427 |
1419 |
3.81 |
|
|
|
| 9 |
A' |
1408 |
1400 |
2.56 |
|
|
|
| 10 |
A' |
1353 |
1345 |
2.23 |
|
|
|
| 11 |
A' |
1337 |
1330 |
6.51 |
|
|
|
| 12 |
A' |
1323 |
1315 |
0.68 |
|
|
|
| 13 |
A' |
1253 |
1245 |
1.02 |
|
|
|
| 14 |
A' |
1223 |
1216 |
10.81 |
|
|
|
| 15 |
A' |
1107 |
1100 |
0.41 |
|
|
|
| 16 |
A' |
994 |
988 |
4.20 |
|
|
|
| 17 |
A' |
950 |
945 |
8.24 |
|
|
|
| 18 |
A' |
920 |
915 |
1.63 |
|
|
|
| 19 |
A' |
905 |
900 |
7.12 |
|
|
|
| 20 |
A' |
828 |
823 |
8.03 |
|
|
|
| 21 |
A' |
628 |
624 |
0.80 |
|
|
|
| 22 |
A' |
400 |
398 |
0.20 |
|
|
|
| 23 |
A' |
331 |
329 |
0.12 |
|
|
|
| 24 |
A' |
64 |
64 |
3.51 |
|
|
|
| 25 |
A" |
3041 |
3023 |
44.75 |
|
|
|
| 26 |
A" |
3037 |
3020 |
0.00 |
|
|
|
| 27 |
A" |
2969 |
2951 |
17.83 |
|
|
|
| 28 |
A" |
2904 |
2888 |
132.15 |
|
|
|
| 29 |
A" |
1437 |
1428 |
8.66 |
|
|
|
| 30 |
A" |
1421 |
1413 |
2.20 |
|
|
|
| 31 |
A" |
1404 |
1396 |
0.07 |
|
|
|
| 32 |
A" |
1246 |
1239 |
2.03 |
|
|
|
| 33 |
A" |
1172 |
1165 |
0.43 |
|
|
|
| 34 |
A" |
1110 |
1104 |
0.06 |
|
|
|
| 35 |
A" |
1012 |
1006 |
9.55 |
|
|
|
| 36 |
A" |
1010 |
1004 |
69.93 |
|
|
|
| 37 |
A" |
898 |
893 |
0.05 |
|
|
|
| 38 |
A" |
864 |
859 |
4.32 |
|
|
|
| 39 |
A" |
380 |
378 |
0.56 |
|
|
|
| 40 |
A" |
304 |
303 |
0.15 |
|
|
|
| 41 |
A" |
262 |
261 |
0.02 |
|
|
|
| 42 |
A" |
212 |
211 |
0.01 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 30248.9 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 30073.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.025 |
|
|
|
| 2 |
H |
0.031 |
|
|
|
| 3 |
H |
0.031 |
|
|
|
| 4 |
H |
0.028 |
|
|
|
| 5 |
C |
0.025 |
|
|
|
| 6 |
H |
0.033 |
|
|
|
| 7 |
H |
0.030 |
|
|
|
| 8 |
H |
0.030 |
|
|
|
| 9 |
C |
-0.435 |
|
|
|
| 10 |
C |
0.238 |
|
|
|
| 11 |
C |
0.238 |
|
|
|
| 12 |
H |
0.012 |
|
|
|
| 13 |
H |
0.012 |
|
|
|
| 14 |
H |
0.006 |
|
|
|
| 15 |
H |
0.006 |
|
|
|
| 16 |
O |
-0.308 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.259 |
1.753 |
0.000 |
1.772 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-38.070 |
0.597 |
0.000 |
| y |
0.597 |
-42.861 |
0.000 |
| z |
0.000 |
0.000 |
-36.552 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.636 |
0.597 |
0.000 |
| y |
0.597 |
-5.549 |
0.000 |
| z |
0.000 |
0.000 |
3.913 |
|
| Polar |
|---|
| 3z2-r2 | 7.826 |
|---|
| x2-y2 | 4.790 |
|---|
| xy | 0.597 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.786 |
0.027 |
0.000 |
| y |
0.027 |
8.653 |
0.000 |
| z |
0.000 |
0.000 |
9.035 |
<r2> (average value of r
2) Å
2
| <r2> |
160.403 |
| (<r2>)1/2 |
12.665 |