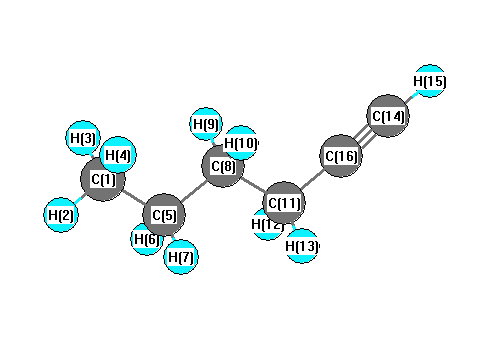
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3421 |
3401 |
69.59 |
|
|
|
| 2 |
A' |
3052 |
3034 |
29.70 |
|
|
|
| 3 |
A' |
2970 |
2953 |
49.42 |
|
|
|
| 4 |
A' |
2965 |
2948 |
18.70 |
|
|
|
| 5 |
A' |
2948 |
2931 |
29.89 |
|
|
|
| 6 |
A' |
2946 |
2929 |
5.29 |
|
|
|
| 7 |
A' |
2169 |
2156 |
9.66 |
|
|
|
| 8 |
A' |
1434 |
1426 |
4.77 |
|
|
|
| 9 |
A' |
1421 |
1413 |
0.53 |
|
|
|
| 10 |
A' |
1412 |
1403 |
0.62 |
|
|
|
| 11 |
A' |
1392 |
1384 |
1.49 |
|
|
|
| 12 |
A' |
1349 |
1342 |
1.32 |
|
|
|
| 13 |
A' |
1338 |
1330 |
1.37 |
|
|
|
| 14 |
A' |
1296 |
1288 |
2.09 |
|
|
|
| 15 |
A' |
1214 |
1207 |
8.34 |
|
|
|
| 16 |
A' |
1090 |
1083 |
1.97 |
|
|
|
| 17 |
A' |
1055 |
1049 |
0.23 |
|
|
|
| 18 |
A' |
1019 |
1013 |
1.48 |
|
|
|
| 19 |
A' |
940 |
935 |
0.12 |
|
|
|
| 20 |
A' |
886 |
881 |
2.55 |
|
|
|
| 21 |
A' |
613 |
610 |
39.77 |
|
|
|
| 22 |
A' |
498 |
495 |
4.06 |
|
|
|
| 23 |
A' |
376 |
374 |
0.01 |
|
|
|
| 24 |
A' |
265 |
263 |
0.76 |
|
|
|
| 25 |
A' |
116 |
115 |
0.04 |
|
|
|
| 26 |
A" |
3043 |
3026 |
44.72 |
|
|
|
| 27 |
A" |
3019 |
3001 |
32.50 |
|
|
|
| 28 |
A" |
2984 |
2967 |
0.37 |
|
|
|
| 29 |
A" |
2978 |
2961 |
9.23 |
|
|
|
| 30 |
A" |
1422 |
1414 |
6.76 |
|
|
|
| 31 |
A" |
1274 |
1267 |
0.10 |
|
|
|
| 32 |
A" |
1257 |
1250 |
0.42 |
|
|
|
| 33 |
A" |
1185 |
1178 |
0.16 |
|
|
|
| 34 |
A" |
1080 |
1074 |
0.10 |
|
|
|
| 35 |
A" |
897 |
891 |
0.38 |
|
|
|
| 36 |
A" |
766 |
761 |
0.66 |
|
|
|
| 37 |
A" |
722 |
718 |
3.71 |
|
|
|
| 38 |
A" |
602 |
599 |
38.97 |
|
|
|
| 39 |
A" |
337 |
335 |
4.26 |
|
|
|
| 40 |
A" |
251 |
250 |
0.04 |
|
|
|
| 41 |
A" |
118 |
117 |
0.00 |
|
|
|
| 42 |
A" |
92 |
91 |
0.01 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 30105.0 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 29930.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.042 |
|
|
|
| 2 |
H |
0.029 |
|
|
|
| 3 |
H |
0.034 |
|
|
|
| 4 |
H |
0.034 |
|
|
|
| 5 |
C |
-0.072 |
|
|
|
| 6 |
H |
0.022 |
|
|
|
| 7 |
H |
0.022 |
|
|
|
| 8 |
C |
-0.047 |
|
|
|
| 9 |
H |
0.031 |
|
|
|
| 10 |
H |
0.031 |
|
|
|
| 11 |
C |
-0.031 |
|
|
|
| 12 |
H |
0.055 |
|
|
|
| 13 |
H |
0.055 |
|
|
|
| 14 |
C |
-0.464 |
|
|
|
| 15 |
H |
-0.030 |
|
|
|
| 16 |
C |
0.375 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.741 |
0.569 |
0.000 |
0.935 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-36.352 |
0.758 |
0.000 |
| y |
0.758 |
-39.711 |
0.000 |
| z |
0.000 |
0.000 |
-38.954 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.981 |
0.758 |
0.000 |
| y |
0.758 |
-2.059 |
0.000 |
| z |
0.000 |
0.000 |
-0.923 |
|
| Polar |
|---|
| 3z2-r2 | -1.845 |
|---|
| x2-y2 | 3.360 |
|---|
| xy | 0.758 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
11.646 |
2.869 |
0.000 |
| y |
2.869 |
10.629 |
0.000 |
| z |
0.000 |
0.000 |
7.376 |
<r2> (average value of r
2) Å
2
| <r2> |
269.281 |
| (<r2>)1/2 |
16.410 |