return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H8 (Ethenylcyclopropane)

using model chemistry: CCD/cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at CCD/cc-pVDZ
 hartrees
Energy at 0K-194.692016
Energy at 298.15K-194.700194
HF Energy-193.954032
Nuclear repulsion energy161.754758
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3272 3130 16.99      
2 A' 3269 3127 14.07      
3 A' 3191 3053 8.40      
4 A' 3180 3042 2.27      
5 A' 3172 3035 11.79      
6 A' 3163 3026 7.85      
7 A' 1742 1666 14.43      
8 A' 1528 1462 1.07      
9 A' 1476 1412 0.11      
10 A' 1370 1310 0.72      
11 A' 1316 1259 3.78      
12 A' 1244 1190 2.03      
13 A' 1234 1180 2.30      
14 A' 1077 1030 1.14      
15 A' 1009 965 2.38      
16 A' 983 940 30.20      
17 A' 833 797 1.13      
18 A' 773 739 0.22      
19 A' 445 426 0.86      
20 A' 267 256 1.02      
21 A" 3255 3114 0.12      
22 A" 3165 3028 15.59      
23 A" 1468 1405 1.05      
24 A" 1203 1151 0.65      
25 A" 1138 1089 1.36      
26 A" 1084 1037 2.91      
27 A" 1018 974 10.66      
28 A" 946 905 9.04      
29 A" 923 883 34.04      
30 A" 838 802 4.96      
31 A" 673 644 4.43      
32 A" 315 301 0.89      
33 A" 116 111 0.04      

Unscaled Zero Point Vibrational Energy (zpe) 25341.8 cm-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 24244.5 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at CCD/cc-pVDZ
ABC
0.49776 0.10042 0.09661

See section I.F.4 to change rotational constant units
Geometric Data calculated at CCD/cc-pVDZ

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -0.393 0.292 0.000
C2 0.259 -1.047 0.000
C3 -0.402 -2.220 0.000
C4 0.259 1.442 0.756
C5 0.259 1.442 -0.756
H6 -1.489 0.272 0.000
H7 1.359 -1.048 0.000
H8 0.133 -3.176 0.000
H9 -1.499 -2.258 0.000
H10 -0.398 2.147 1.275
H11 1.204 1.234 1.270
H12 -0.398 2.147 -1.275
H13 1.204 1.234 -1.270

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 H6 H7 H8 H9 H10 H11 H12 H13
C11.48922.51151.52271.52271.09662.20573.50692.77932.25122.24722.25122.2472
C21.48921.34582.60142.60142.19041.10072.13192.13413.50162.77673.50162.7767
C32.51151.34583.79683.79682.71912.11571.09531.09724.54904.01504.54904.0150
C41.52272.60143.79681.51192.23482.82524.68054.16531.09481.09562.24842.2449
C51.52272.60143.79681.51192.23482.82524.68054.16532.24842.24491.09481.0956
H61.09662.19042.71912.23482.23483.13993.81062.53082.51593.12892.51593.1289
H72.20571.10072.11572.82522.82523.13992.45583.10383.86302.61613.86302.6161
H83.50692.13191.09534.68054.68053.81062.45581.87175.49884.71215.49884.7121
H92.77932.13411.09724.16534.16532.53083.10381.87174.71614.59484.71614.5948
H102.25123.50164.54901.09482.24842.51593.86305.49884.71611.84412.55073.1428
H112.24722.77674.01501.09562.24493.12892.61614.71214.59481.84413.14282.5392
H122.25123.50164.54902.24841.09482.51593.86305.49884.71612.55073.14281.8441
H132.24722.77674.01502.24491.09563.12892.61614.71214.59483.14282.53921.8441

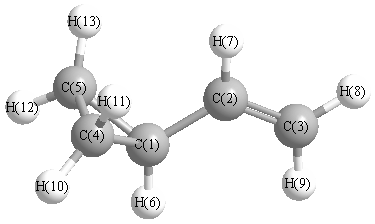
picture of Ethenylcyclopropane state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 124.646 C1 C2 H7 115.978
C1 C4 C5 60.234 C1 C4 H10 117.715
C1 C4 H11 117.303 C1 C5 C4 60.234
C1 C5 H12 117.715 C1 C5 H13 117.303
C2 C1 C4 119.466 C2 C1 C5 119.466
C2 C1 H6 114.946 C2 C3 H8 121.356
C2 C3 H9 121.414 C3 C2 H7 119.376
C4 C1 H6 116.170 C4 C5 H12 118.321
C4 C5 H13 117.957 C5 C1 H6 116.170
C5 C4 H10 118.321 C5 C4 H11 117.957
H8 C3 H9 117.231 H10 C4 H11 114.675
H12 C5 H13 114.675
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability