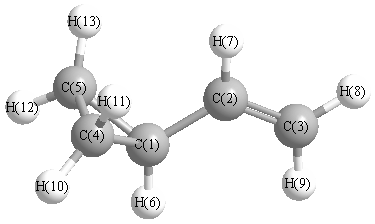
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3172 |
3154 |
13.69 |
|
|
|
| 2 |
A' |
3164 |
3146 |
12.61 |
|
|
|
| 3 |
A' |
3084 |
3066 |
10.94 |
|
|
|
| 4 |
A' |
3075 |
3057 |
3.01 |
|
|
|
| 5 |
A' |
3066 |
3048 |
10.83 |
|
|
|
| 6 |
A' |
3052 |
3034 |
10.25 |
|
|
|
| 7 |
A' |
1664 |
1654 |
33.29 |
|
|
|
| 8 |
A' |
1434 |
1426 |
0.96 |
|
|
|
| 9 |
A' |
1396 |
1388 |
0.06 |
|
|
|
| 10 |
A' |
1305 |
1298 |
1.13 |
|
|
|
| 11 |
A' |
1260 |
1253 |
2.89 |
|
|
|
| 12 |
A' |
1196 |
1189 |
4.71 |
|
|
|
| 13 |
A' |
1189 |
1182 |
0.81 |
|
|
|
| 14 |
A' |
1001 |
996 |
1.78 |
|
|
|
| 15 |
A' |
964 |
959 |
3.24 |
|
|
|
| 16 |
A' |
951 |
945 |
38.79 |
|
|
|
| 17 |
A' |
806 |
801 |
0.91 |
|
|
|
| 18 |
A' |
741 |
737 |
0.17 |
|
|
|
| 19 |
A' |
434 |
432 |
1.12 |
|
|
|
| 20 |
A' |
261 |
260 |
1.20 |
|
|
|
| 21 |
A" |
3150 |
3131 |
0.10 |
|
|
|
| 22 |
A" |
3062 |
3044 |
17.36 |
|
|
|
| 23 |
A" |
1383 |
1375 |
1.77 |
|
|
|
| 24 |
A" |
1143 |
1137 |
0.68 |
|
|
|
| 25 |
A" |
1074 |
1068 |
2.63 |
|
|
|
| 26 |
A" |
1009 |
1004 |
5.64 |
|
|
|
| 27 |
A" |
979 |
973 |
5.53 |
|
|
|
| 28 |
A" |
880 |
875 |
1.84 |
|
|
|
| 29 |
A" |
866 |
861 |
37.24 |
|
|
|
| 30 |
A" |
801 |
797 |
4.35 |
|
|
|
| 31 |
A" |
656 |
652 |
4.72 |
|
|
|
| 32 |
A" |
301 |
299 |
0.81 |
|
|
|
| 33 |
A" |
123 |
122 |
0.05 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 24321.5 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 24180.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.082 |
|
|
|
| 2 |
C |
-0.006 |
|
|
|
| 3 |
C |
-0.013 |
|
|
|
| 4 |
C |
-0.021 |
|
|
|
| 5 |
C |
-0.021 |
|
|
|
| 6 |
H |
-0.014 |
|
|
|
| 7 |
H |
-0.021 |
|
|
|
| 8 |
H |
0.022 |
|
|
|
| 9 |
H |
0.014 |
|
|
|
| 10 |
H |
0.037 |
|
|
|
| 11 |
H |
0.034 |
|
|
|
| 12 |
H |
0.037 |
|
|
|
| 13 |
H |
0.034 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.023 |
0.613 |
0.000 |
0.613 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-29.760 |
-1.011 |
0.000 |
| y |
-1.011 |
-31.589 |
0.000 |
| z |
0.000 |
0.000 |
-33.571 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.820 |
-1.011 |
0.000 |
| y |
-1.011 |
0.077 |
0.000 |
| z |
0.000 |
0.000 |
-2.897 |
|
| Polar |
|---|
| 3z2-r2 | -5.794 |
|---|
| x2-y2 | 1.828 |
|---|
| xy | -1.011 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.603 |
0.865 |
0.000 |
| y |
0.865 |
11.450 |
0.000 |
| z |
0.000 |
0.000 |
5.976 |
<r2> (average value of r
2) Å
2
| <r2> |
136.144 |
| (<r2>)1/2 |
11.668 |