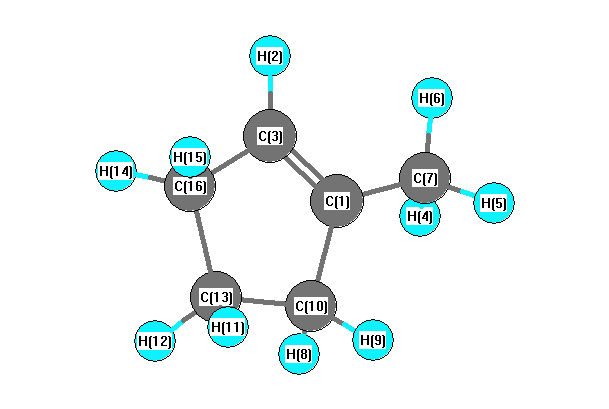
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3115 |
3097 |
18.36 |
|
|
|
| 2 |
A |
3059 |
3041 |
18.42 |
|
|
|
| 3 |
A |
3040 |
3023 |
35.45 |
|
|
|
| 4 |
A |
3004 |
2987 |
17.79 |
|
|
|
| 5 |
A |
2997 |
2979 |
54.53 |
|
|
|
| 6 |
A |
2986 |
2969 |
35.10 |
|
|
|
| 7 |
A |
2976 |
2959 |
7.42 |
|
|
|
| 8 |
A |
2949 |
2932 |
42.64 |
|
|
|
| 9 |
A |
2927 |
2910 |
55.62 |
|
|
|
| 10 |
A |
2926 |
2909 |
36.02 |
|
|
|
| 11 |
A |
1677 |
1667 |
2.89 |
|
|
|
| 12 |
A |
1425 |
1417 |
1.55 |
|
|
|
| 13 |
A |
1409 |
1401 |
7.14 |
|
|
|
| 14 |
A |
1404 |
1396 |
0.69 |
|
|
|
| 15 |
A |
1399 |
1391 |
0.27 |
|
|
|
| 16 |
A |
1397 |
1389 |
8.65 |
|
|
|
| 17 |
A |
1347 |
1339 |
1.62 |
|
|
|
| 18 |
A |
1310 |
1302 |
1.22 |
|
|
|
| 19 |
A |
1277 |
1269 |
0.57 |
|
|
|
| 20 |
A |
1269 |
1262 |
1.95 |
|
|
|
| 21 |
A |
1221 |
1214 |
1.60 |
|
|
|
| 22 |
A |
1208 |
1201 |
1.61 |
|
|
|
| 23 |
A |
1178 |
1172 |
1.26 |
|
|
|
| 24 |
A |
1137 |
1130 |
0.35 |
|
|
|
| 25 |
A |
1101 |
1094 |
1.51 |
|
|
|
| 26 |
A |
1039 |
1033 |
0.29 |
|
|
|
| 27 |
A |
1020 |
1014 |
2.68 |
|
|
|
| 28 |
A |
987 |
981 |
7.96 |
|
|
|
| 29 |
A |
973 |
967 |
5.65 |
|
|
|
| 30 |
A |
921 |
916 |
1.00 |
|
|
|
| 31 |
A |
903 |
898 |
0.69 |
|
|
|
| 32 |
A |
874 |
869 |
0.15 |
|
|
|
| 33 |
A |
853 |
848 |
0.55 |
|
|
|
| 34 |
A |
805 |
801 |
1.81 |
|
|
|
| 35 |
A |
775 |
771 |
8.96 |
|
|
|
| 36 |
A |
630 |
626 |
1.74 |
|
|
|
| 37 |
A |
563 |
560 |
0.25 |
|
|
|
| 38 |
A |
427 |
424 |
3.25 |
|
|
|
| 39 |
A |
312 |
310 |
0.60 |
|
|
|
| 40 |
A |
237 |
236 |
1.95 |
|
|
|
| 41 |
A |
178 |
177 |
0.90 |
|
|
|
| 42 |
A |
144 |
143 |
0.09 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 30686.8 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 30508.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.140 |
|
|
|
| 2 |
H |
-0.040 |
|
|
|
| 3 |
C |
-0.044 |
|
|
|
| 4 |
H |
0.041 |
|
|
|
| 5 |
H |
0.042 |
|
|
|
| 6 |
H |
0.029 |
|
|
|
| 7 |
C |
0.006 |
|
|
|
| 8 |
H |
0.031 |
|
|
|
| 9 |
H |
0.018 |
|
|
|
| 10 |
C |
0.003 |
|
|
|
| 11 |
H |
0.030 |
|
|
|
| 12 |
H |
0.011 |
|
|
|
| 13 |
C |
-0.040 |
|
|
|
| 14 |
H |
0.033 |
|
|
|
| 15 |
H |
0.023 |
|
|
|
| 16 |
C |
-0.000 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.186 |
-0.206 |
0.060 |
0.284 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-37.394 |
0.251 |
0.073 |
| y |
0.251 |
-37.795 |
-0.062 |
| z |
0.073 |
-0.062 |
-38.968 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.987 |
0.251 |
0.073 |
| y |
0.251 |
0.386 |
-0.062 |
| z |
0.073 |
-0.062 |
-1.373 |
|
| Polar |
|---|
| 3z2-r2 | -2.746 |
|---|
| x2-y2 | 0.401 |
|---|
| xy | 0.251 |
|---|
| xz | 0.073 |
|---|
| yz | -0.062 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
11.733 |
0.858 |
0.091 |
| y |
0.858 |
10.509 |
-0.113 |
| z |
0.091 |
-0.113 |
7.363 |
<r2> (average value of r
2) Å
2
| <r2> |
171.889 |
| (<r2>)1/2 |
13.111 |