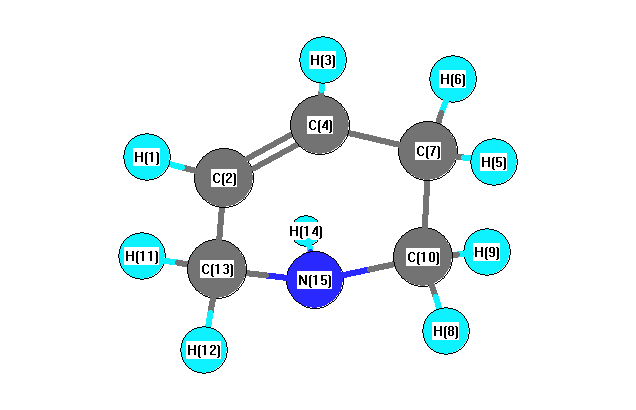
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3375 |
3355 |
1.13 |
|
|
|
| 2 |
A |
3097 |
3079 |
36.35 |
|
|
|
| 3 |
A |
3072 |
3054 |
10.24 |
|
|
|
| 4 |
A |
3012 |
2995 |
37.28 |
|
|
|
| 5 |
A |
2982 |
2965 |
25.41 |
|
|
|
| 6 |
A |
2973 |
2955 |
53.86 |
|
|
|
| 7 |
A |
2954 |
2937 |
24.58 |
|
|
|
| 8 |
A |
2926 |
2909 |
36.21 |
|
|
|
| 9 |
A |
2906 |
2889 |
47.27 |
|
|
|
| 10 |
A |
1675 |
1666 |
2.73 |
|
|
|
| 11 |
A |
1436 |
1428 |
4.48 |
|
|
|
| 12 |
A |
1408 |
1399 |
2.82 |
|
|
|
| 13 |
A |
1391 |
1383 |
9.78 |
|
|
|
| 14 |
A |
1387 |
1379 |
1.02 |
|
|
|
| 15 |
A |
1360 |
1352 |
0.66 |
|
|
|
| 16 |
A |
1332 |
1325 |
5.36 |
|
|
|
| 17 |
A |
1312 |
1305 |
4.83 |
|
|
|
| 18 |
A |
1288 |
1281 |
0.30 |
|
|
|
| 19 |
A |
1273 |
1266 |
0.52 |
|
|
|
| 20 |
A |
1209 |
1202 |
4.87 |
|
|
|
| 21 |
A |
1170 |
1164 |
12.16 |
|
|
|
| 22 |
A |
1146 |
1139 |
3.52 |
|
|
|
| 23 |
A |
1096 |
1090 |
15.11 |
|
|
|
| 24 |
A |
1065 |
1059 |
3.61 |
|
|
|
| 25 |
A |
1023 |
1017 |
4.87 |
|
|
|
| 26 |
A |
979 |
973 |
5.50 |
|
|
|
| 27 |
A |
951 |
946 |
0.11 |
|
|
|
| 28 |
A |
938 |
933 |
1.66 |
|
|
|
| 29 |
A |
901 |
896 |
5.48 |
|
|
|
| 30 |
A |
876 |
871 |
11.24 |
|
|
|
| 31 |
A |
838 |
833 |
7.75 |
|
|
|
| 32 |
A |
790 |
785 |
88.65 |
|
|
|
| 33 |
A |
733 |
729 |
18.02 |
|
|
|
| 34 |
A |
624 |
621 |
21.13 |
|
|
|
| 35 |
A |
511 |
508 |
1.00 |
|
|
|
| 36 |
A |
470 |
468 |
0.37 |
|
|
|
| 37 |
A |
385 |
383 |
1.01 |
|
|
|
| 38 |
A |
286 |
285 |
6.41 |
|
|
|
| 39 |
A |
159 |
158 |
1.86 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 28654.8 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 28488.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
H |
-0.020 |
|
|
|
| 2 |
C |
-0.060 |
|
|
|
| 3 |
H |
-0.017 |
|
|
|
| 4 |
C |
-0.035 |
|
|
|
| 5 |
H |
0.025 |
|
|
|
| 6 |
H |
0.031 |
|
|
|
| 7 |
C |
-0.020 |
|
|
|
| 8 |
H |
0.032 |
|
|
|
| 9 |
H |
0.014 |
|
|
|
| 10 |
C |
0.026 |
|
|
|
| 11 |
H |
0.028 |
|
|
|
| 12 |
H |
0.043 |
|
|
|
| 13 |
C |
0.059 |
|
|
|
| 14 |
H |
0.079 |
|
|
|
| 15 |
N |
-0.184 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.423 |
0.507 |
-0.500 |
0.828 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-35.906 |
-1.701 |
0.213 |
| y |
-1.701 |
-39.870 |
0.873 |
| z |
0.213 |
0.873 |
-36.908 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.483 |
-1.701 |
0.213 |
| y |
-1.701 |
-3.464 |
0.873 |
| z |
0.213 |
0.873 |
0.981 |
|
| Polar |
|---|
| 3z2-r2 | 1.961 |
|---|
| x2-y2 | 3.964 |
|---|
| xy | -1.701 |
|---|
| xz | 0.213 |
|---|
| yz | 0.873 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
11.190 |
-0.181 |
-0.252 |
| y |
-0.181 |
9.008 |
0.060 |
| z |
-0.252 |
0.060 |
6.929 |
<r2> (average value of r
2) Å
2
| <r2> |
148.659 |
| (<r2>)1/2 |
12.193 |