Jump to
S1C2
Energy calculated at PBEPBE/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -194.099528 |
| Energy at 298.15K | -194.108339 |
| Nuclear repulsion energy | 129.557877 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3677 |
3655 |
13.65 |
|
|
|
| 2 |
A' |
3055 |
3037 |
26.14 |
|
|
|
| 3 |
A' |
2972 |
2954 |
30.54 |
|
|
|
| 4 |
A' |
2965 |
2948 |
25.41 |
|
|
|
| 5 |
A' |
2866 |
2849 |
72.20 |
|
|
|
| 6 |
A' |
1451 |
1442 |
4.10 |
|
|
|
| 7 |
A' |
1433 |
1424 |
4.71 |
|
|
|
| 8 |
A' |
1418 |
1409 |
0.97 |
|
|
|
| 9 |
A' |
1402 |
1394 |
1.94 |
|
|
|
| 10 |
A' |
1345 |
1337 |
0.20 |
|
|
|
| 11 |
A' |
1290 |
1282 |
10.43 |
|
|
|
| 12 |
A' |
1220 |
1213 |
45.60 |
|
|
|
| 13 |
A' |
1073 |
1067 |
19.10 |
|
|
|
| 14 |
A' |
1045 |
1039 |
29.49 |
|
|
|
| 15 |
A' |
1030 |
1024 |
53.63 |
|
|
|
| 16 |
A' |
869 |
864 |
4.84 |
|
|
|
| 17 |
A' |
447 |
444 |
9.49 |
|
|
|
| 18 |
A' |
262 |
260 |
3.25 |
|
|
|
| 19 |
A" |
3042 |
3025 |
51.69 |
|
|
|
| 20 |
A" |
3012 |
2994 |
9.04 |
|
|
|
| 21 |
A" |
2893 |
2876 |
70.62 |
|
|
|
| 22 |
A" |
1421 |
1413 |
6.93 |
|
|
|
| 23 |
A" |
1261 |
1254 |
0.14 |
|
|
|
| 24 |
A" |
1207 |
1200 |
0.00 |
|
|
|
| 25 |
A" |
1132 |
1126 |
0.74 |
|
|
|
| 26 |
A" |
865 |
860 |
2.83 |
|
|
|
| 27 |
A" |
749 |
744 |
1.55 |
|
|
|
| 28 |
A" |
272 |
271 |
100.16 |
|
|
|
| 29 |
A" |
231 |
230 |
2.73 |
|
|
|
| 30 |
A" |
125 |
124 |
3.94 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 23013.3 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 22879.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBEPBE/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.445 |
1.243 |
0.000 |
| C2 |
0.000 |
0.739 |
0.000 |
| C3 |
0.096 |
-0.782 |
0.000 |
| O4 |
1.479 |
-1.135 |
0.000 |
| H5 |
-1.486 |
2.350 |
0.000 |
| H6 |
-1.999 |
0.889 |
0.895 |
| H7 |
-1.999 |
0.889 |
-0.895 |
| H8 |
0.546 |
1.121 |
0.889 |
| H9 |
0.546 |
1.121 |
-0.889 |
| H10 |
-0.435 |
-1.188 |
0.899 |
| H11 |
-0.435 |
-1.188 |
-0.899 |
| H12 |
1.524 |
-2.110 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
| C1 | | 1.5305 | 2.5451 | 3.7688 | 1.1082 | 1.1105 | 1.1105 | 2.1838 | 2.1838 | 2.7815 | 2.7815 | 4.4786 |
C2 | 1.5305 | | 1.5242 | 2.3871 | 2.1920 | 2.1958 | 2.1958 | 1.1113 | 1.1113 | 2.1700 | 2.1700 | 3.2308 | C3 | 2.5451 | 1.5242 | | 1.4269 | 3.5095 | 2.8263 | 2.8263 | 2.1486 | 2.1486 | 1.1199 | 1.1199 | 1.9497 | O4 | 3.7688 | 2.3871 | 1.4269 | | 4.5757 | 4.1229 | 4.1229 | 2.5984 | 2.5984 | 2.1149 | 2.1149 | 0.9759 | H5 | 1.1082 | 2.1920 | 3.5095 | 4.5757 | | 1.7885 | 1.7885 | 2.5356 | 2.5356 | 3.7988 | 3.7988 | 5.3809 | H6 | 1.1105 | 2.1958 | 2.8263 | 4.1229 | 1.7885 | | 1.7899 | 2.5558 | 3.1170 | 2.6006 | 3.1592 | 4.7131 | H7 | 1.1105 | 2.1958 | 2.8263 | 4.1229 | 1.7885 | 1.7899 | | 3.1170 | 2.5558 | 3.1592 | 2.6006 | 4.7131 | H8 | 2.1838 | 1.1113 | 2.1486 | 2.5984 | 2.5356 | 2.5558 | 3.1170 | | 1.7786 | 2.5087 | 3.0806 | 3.4911 | H9 | 2.1838 | 1.1113 | 2.1486 | 2.5984 | 2.5356 | 3.1170 | 2.5558 | 1.7786 | | 3.0806 | 2.5087 | 3.4911 | H10 | 2.7815 | 2.1700 | 1.1199 | 2.1149 | 3.7988 | 2.6006 | 3.1592 | 2.5087 | 3.0806 | | 1.7974 | 2.3444 | H11 | 2.7815 | 2.1700 | 1.1199 | 2.1149 | 3.7988 | 3.1592 | 2.6006 | 3.0806 | 2.5087 | 1.7974 | | 2.3444 | H12 | 4.4786 | 3.2308 | 1.9497 | 0.9759 | 5.3809 | 4.7131 | 4.7131 | 3.4911 | 3.4911 | 2.3444 | 2.3444 | |
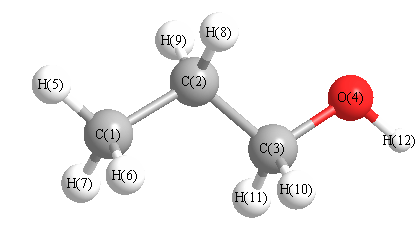 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
112.850 |
|
C1 |
C2 |
H8 |
110.511 |
| C1 |
C2 |
H9 |
110.511 |
|
C2 |
C1 |
H5 |
111.342 |
| C2 |
C1 |
H6 |
111.504 |
|
C2 |
C1 |
H7 |
111.504 |
| C2 |
C3 |
O4 |
107.927 |
|
C2 |
C3 |
H10 |
109.354 |
| C2 |
C3 |
H11 |
109.354 |
|
C3 |
C2 |
H8 |
108.202 |
| C3 |
C2 |
H9 |
108.202 |
|
C3 |
O4 |
H12 |
106.973 |
| O4 |
C3 |
H10 |
111.721 |
|
O4 |
C3 |
H11 |
111.721 |
| H5 |
C1 |
H6 |
107.436 |
|
H5 |
C1 |
H7 |
107.436 |
| H6 |
C1 |
H7 |
107.393 |
|
H8 |
C2 |
H9 |
106.313 |
| H10 |
C3 |
H11 |
106.735 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.072 |
|
|
|
| 2 |
C |
-0.073 |
|
|
|
| 3 |
C |
0.081 |
|
|
|
| 4 |
O |
-0.242 |
|
|
|
| 5 |
H |
0.033 |
|
|
|
| 6 |
H |
0.032 |
|
|
|
| 7 |
H |
0.032 |
|
|
|
| 8 |
H |
0.029 |
|
|
|
| 9 |
H |
0.029 |
|
|
|
| 10 |
H |
0.008 |
|
|
|
| 11 |
H |
0.008 |
|
|
|
| 12 |
H |
0.136 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.878 |
-0.974 |
0.000 |
1.312 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-28.882 |
-1.172 |
0.000 |
| y |
-1.172 |
-22.020 |
0.000 |
| z |
0.000 |
0.000 |
-26.653 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-4.545 |
-1.172 |
0.000 |
| y |
-1.172 |
5.747 |
0.000 |
| z |
0.000 |
0.000 |
-1.202 |
|
| Polar |
|---|
| 3z2-r2 | -2.405 |
|---|
| x2-y2 | -6.862 |
|---|
| xy | -1.172 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.247 |
-0.589 |
0.000 |
| y |
-0.589 |
6.491 |
0.000 |
| z |
0.000 |
0.000 |
5.312 |
<r2> (average value of r
2) Å
2
| <r2> |
0.000 |
| (<r2>)1/2 |
0.000 |
Jump to
S1C1
Energy calculated at PBEPBE/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -194.100136 |
| Energy at 298.15K | |
| HF Energy | -194.100136 |
| Nuclear repulsion energy | 131.886405 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3679 |
3658 |
16.80 |
126.74 |
0.30 |
0.46 |
| 2 |
A |
3070 |
3052 |
18.70 |
49.78 |
0.75 |
0.86 |
| 3 |
A |
3048 |
3030 |
38.04 |
62.73 |
0.60 |
0.75 |
| 4 |
A |
3009 |
2992 |
20.56 |
104.09 |
0.70 |
0.83 |
| 5 |
A |
2970 |
2953 |
19.99 |
169.52 |
0.02 |
0.03 |
| 6 |
A |
2968 |
2951 |
46.06 |
90.91 |
0.10 |
0.19 |
| 7 |
A |
2904 |
2888 |
70.89 |
109.46 |
0.73 |
0.85 |
| 8 |
A |
2874 |
2857 |
74.45 |
131.59 |
0.10 |
0.18 |
| 9 |
A |
1443 |
1435 |
3.09 |
6.86 |
0.63 |
0.78 |
| 10 |
A |
1434 |
1426 |
6.93 |
8.37 |
0.75 |
0.86 |
| 11 |
A |
1414 |
1406 |
7.33 |
20.18 |
0.75 |
0.86 |
| 12 |
A |
1399 |
1391 |
4.80 |
14.19 |
0.74 |
0.85 |
| 13 |
A |
1397 |
1388 |
1.07 |
2.56 |
0.58 |
0.73 |
| 14 |
A |
1343 |
1336 |
4.57 |
3.30 |
0.75 |
0.86 |
| 15 |
A |
1313 |
1305 |
1.23 |
1.55 |
0.74 |
0.85 |
| 16 |
A |
1276 |
1268 |
23.54 |
11.71 |
0.75 |
0.86 |
| 17 |
A |
1214 |
1207 |
5.06 |
10.93 |
0.71 |
0.83 |
| 18 |
A |
1206 |
1199 |
25.61 |
3.95 |
0.53 |
0.69 |
| 19 |
A |
1112 |
1106 |
4.93 |
1.08 |
0.34 |
0.51 |
| 20 |
A |
1091 |
1084 |
20.68 |
2.87 |
0.61 |
0.76 |
| 21 |
A |
1049 |
1043 |
25.89 |
2.99 |
0.74 |
0.85 |
| 22 |
A |
960 |
954 |
51.17 |
3.49 |
0.75 |
0.85 |
| 23 |
A |
892 |
886 |
2.15 |
0.32 |
0.49 |
0.66 |
| 24 |
A |
860 |
855 |
2.14 |
8.89 |
0.18 |
0.31 |
| 25 |
A |
749 |
745 |
0.88 |
0.50 |
0.63 |
0.78 |
| 26 |
A |
463 |
461 |
6.91 |
0.29 |
0.41 |
0.58 |
| 27 |
A |
321 |
319 |
5.89 |
0.27 |
0.56 |
0.72 |
| 28 |
A |
247 |
245 |
90.59 |
4.75 |
0.74 |
0.85 |
| 29 |
A |
221 |
220 |
3.01 |
0.22 |
0.73 |
0.84 |
| 30 |
A |
148 |
147 |
7.88 |
0.17 |
0.74 |
0.85 |
Unscaled Zero Point Vibrational Energy (zpe) 23035.9 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 22902.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBEPBE/cc-pVDZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.539 |
-0.516 |
0.131 |
| C2 |
-0.635 |
0.645 |
-0.289 |
| C3 |
0.773 |
0.546 |
0.292 |
| O4 |
1.379 |
-0.640 |
-0.226 |
| H5 |
-2.532 |
-0.458 |
-0.358 |
| H6 |
-1.704 |
-0.520 |
1.229 |
| H7 |
-1.073 |
-1.483 |
-0.142 |
| H8 |
-0.546 |
0.676 |
-1.397 |
| H9 |
-1.074 |
1.618 |
0.021 |
| H10 |
0.712 |
0.521 |
1.410 |
| H11 |
1.358 |
1.458 |
0.016 |
| H12 |
2.236 |
-0.745 |
0.228 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
| C1 | | 1.5308 | 2.5497 | 2.9426 | 1.1084 | 1.1108 | 1.1077 | 2.1778 | 2.1867 | 2.7892 | 3.5082 | 3.7836 |
C2 | 1.5308 | | 1.5258 | 2.3892 | 2.1962 | 2.1925 | 2.1774 | 1.1120 | 1.1116 | 2.1715 | 2.1738 | 3.2312 | C3 | 2.5497 | 1.5258 | | 1.4292 | 3.5150 | 2.8547 | 2.7768 | 2.1469 | 2.1524 | 1.1198 | 1.1184 | 1.9528 | O4 | 2.9426 | 2.3892 | 1.4292 | | 3.9177 | 3.4109 | 2.5936 | 2.6093 | 3.3429 | 2.1135 | 2.1123 | 0.9757 | H5 | 1.1084 | 2.1962 | 3.5150 | 3.9177 | | 1.7919 | 1.7968 | 2.5123 | 2.5648 | 3.8223 | 4.3530 | 4.8129 | H6 | 1.1108 | 2.1925 | 2.8547 | 3.4109 | 1.7919 | | 1.7905 | 3.1098 | 2.5353 | 2.6366 | 3.8422 | 4.0711 | H7 | 1.1077 | 2.1774 | 2.7768 | 2.5936 | 1.7968 | 1.7905 | | 2.5520 | 3.1048 | 3.0995 | 3.8188 | 3.4098 | H8 | 2.1778 | 1.1120 | 2.1469 | 2.6093 | 2.5123 | 3.1098 | 2.5520 | | 1.7824 | 3.0800 | 2.4969 | 3.5215 | H9 | 2.1867 | 1.1116 | 2.1524 | 3.3429 | 2.5648 | 2.5353 | 3.1048 | 1.7824 | | 2.5147 | 2.4375 | 4.0724 | H10 | 2.7892 | 2.1715 | 1.1198 | 2.1135 | 3.8223 | 2.6366 | 3.0995 | 3.0800 | 2.5147 | | 1.8000 | 2.3068 | H11 | 3.5082 | 2.1738 | 1.1184 | 2.1123 | 4.3530 | 3.8422 | 3.8188 | 2.4969 | 2.4375 | 1.8000 | | 2.3817 | H12 | 3.7836 | 3.2312 | 1.9528 | 0.9757 | 4.8129 | 4.0711 | 3.4098 | 3.5215 | 4.0724 | 2.3068 | 2.3817 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
113.057 |
|
C1 |
C2 |
H8 |
109.976 |
| C1 |
C2 |
H9 |
110.695 |
|
C2 |
C1 |
H5 |
111.637 |
| C2 |
C1 |
H6 |
111.199 |
|
C2 |
C1 |
H7 |
110.192 |
| C2 |
C3 |
O4 |
107.865 |
|
C2 |
C3 |
H10 |
109.375 |
| C2 |
C3 |
H11 |
109.626 |
|
C3 |
C2 |
H8 |
107.932 |
| C3 |
C2 |
H9 |
108.371 |
|
C3 |
O4 |
H12 |
107.074 |
| O4 |
C3 |
H10 |
111.448 |
|
O4 |
C3 |
H11 |
111.440 |
| H5 |
C1 |
H6 |
107.693 |
|
H5 |
C1 |
H7 |
108.352 |
| H6 |
C1 |
H7 |
107.623 |
|
H8 |
C2 |
H9 |
106.563 |
| H10 |
C3 |
H11 |
107.072 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.059 |
|
|
|
| 2 |
C |
-0.074 |
|
|
|
| 3 |
C |
0.080 |
|
|
|
| 4 |
O |
-0.241 |
|
|
|
| 5 |
H |
0.026 |
|
|
|
| 6 |
H |
0.025 |
|
|
|
| 7 |
H |
0.042 |
|
|
|
| 8 |
H |
0.030 |
|
|
|
| 9 |
H |
0.016 |
|
|
|
| 10 |
H |
0.008 |
|
|
|
| 11 |
H |
0.008 |
|
|
|
| 12 |
H |
0.138 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.775 |
0.660 |
0.918 |
1.371 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-22.988 |
-0.495 |
1.921 |
| y |
-0.495 |
-26.986 |
-0.570 |
| z |
1.921 |
-0.570 |
-26.774 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.892 |
-0.495 |
1.921 |
| y |
-0.495 |
-2.105 |
-0.570 |
| z |
1.921 |
-0.570 |
-1.787 |
|
| Polar |
|---|
| 3z2-r2 | -3.574 |
|---|
| x2-y2 | 3.998 |
|---|
| xy | -0.495 |
|---|
| xz | 1.921 |
|---|
| yz | -0.570 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.639 |
0.078 |
0.184 |
| y |
0.078 |
5.864 |
-0.009 |
| z |
0.184 |
-0.009 |
5.517 |
<r2> (average value of r
2) Å
2
| <r2> |
95.719 |
| (<r2>)1/2 |
9.784 |
