return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H10O (1-Butanol)

using model chemistry: CCD/cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at CCD/cc-pVDZ
 hartrees
Energy at 0K-232.994227
Energy at 298.15K-233.005442
HF Energy-232.165714
Nuclear repulsion energy184.247047
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3886 3718 24.65      
2 A' 3150 3013 34.86      
3 A' 3068 2936 37.07      
4 A' 3062 2929 17.94      
5 A' 3053 2920 36.47      
6 A' 3008 2878 53.42      
7 A' 1535 1469 3.15      
8 A' 1512 1446 4.43      
9 A' 1502 1437 2.86      
10 A' 1494 1429 1.26      
11 A' 1490 1426 3.57      
12 A' 1430 1368 1.24      
13 A' 1410 1349 4.92      
14 A' 1329 1271 34.67      
15 A' 1264 1209 35.98      
16 A' 1143 1093 16.30      
17 A' 1115 1067 53.99      
18 A' 1096 1049 9.18      
19 A' 1037 992 0.33      
20 A' 922 882 8.35      
21 A' 444 425 11.76      
22 A' 399 382 0.09      
23 A' 185 177 2.33      
24 A" 3146 3010 55.33      
25 A" 3123 2987 41.17      
26 A" 3091 2957 6.18      
27 A" 3045 2913 54.61      
28 A" 1500 1435 5.68      
29 A" 1329 1271 0.24      
30 A" 1324 1266 0.35      
31 A" 1258 1204 0.53      
32 A" 1201 1149 2.69      
33 A" 956 915 0.23      
34 A" 820 784 1.38      
35 A" 750 718 1.02      
36 A" 288 275 109.27      
37 A" 258 247 1.56      
38 A" 119 114 1.16      
39 A" 111 106 4.84      

Unscaled Zero Point Vibrational Energy (zpe) 30424.5 cm-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 29107.1 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at CCD/cc-pVDZ
ABC
0.61729 0.06569 0.06217

See section I.F.4 to change rotational constant units
Geometric Data calculated at CCD/cc-pVDZ

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 1.364 -0.340 0.000
C2 0.000 0.341 0.000
C3 -1.167 -0.655 0.000
C4 -2.535 0.039 0.000
O5 2.358 0.676 0.000
H6 1.454 -0.993 0.895
H7 1.454 -0.993 -0.895
H8 -0.062 0.999 0.888
H9 -0.062 0.999 -0.888
H10 -1.088 -1.315 0.887
H11 -1.088 -1.315 -0.887
H12 -3.360 -0.695 0.000
H13 -2.651 0.681 0.892
H14 -2.651 0.681 -0.892
H15 3.214 0.235 0.000

Atom - Atom Distances (Å)
  C1 C2 C3 C4 O5 H6 H7 H8 H9 H10 H11 H12 H13 H14 H15
C11.52472.55043.91761.42031.11141.11142.14842.14842.78382.78384.73754.23824.23821.9372
C21.52471.53372.55282.38122.16642.16641.10701.10702.17052.17053.51602.81792.81793.2162
C32.55041.53371.53423.76692.78952.78952.17812.17811.10821.10822.19362.18752.18754.4704
C43.91762.55281.53424.93374.21614.21612.79742.79742.17112.17111.10441.10561.10565.7526
O51.42032.38123.76694.93372.09782.09782.59762.59764.07654.07655.87945.08765.08760.9637
H61.11142.16642.78954.21612.09781.78972.50313.07312.56193.12044.90514.43314.77982.3253
H71.11142.16642.78954.21612.09781.78973.07312.50313.12042.56194.90514.77984.43312.3253
H82.14841.10702.17812.79742.59762.50313.07311.77582.53123.09133.81252.60863.15823.4797
H92.14841.10702.17812.79742.59763.07312.50311.77583.09132.53123.81253.15822.60863.4797
H102.78382.17051.10822.17114.07652.56193.12042.53123.09131.77332.51652.53543.09744.6578
H112.78382.17051.10822.17114.07653.12042.56193.09132.53121.77332.51653.09742.53544.6578
H124.73753.51602.19361.10445.87944.90514.90513.81253.81252.51652.51651.78701.78706.6397
H134.23822.81792.18751.10565.08764.43314.77982.60863.15822.53543.09741.78701.78505.9499
H144.23822.81792.18751.10565.08764.77984.43313.15822.60863.09742.53541.78701.78505.9499
H151.93723.21624.47045.75260.96372.32532.32533.47973.47974.65784.65786.63975.94995.9499

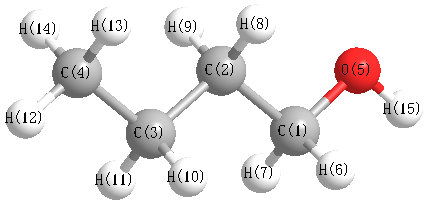
picture of 1-Butanol state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 113.001 C1 C2 H8 108.394
C1 C2 H9 108.394 C1 O5 H15 107.137
C2 C1 O5 107.856 C2 C1 H6 109.533
C2 C1 H7 109.533 C2 C3 C4 112.628
C2 C3 H10 109.425 C2 C3 H11 109.425
C3 C2 H8 110.087 C3 C2 H9 110.087
C3 C4 H12 111.438 C3 C4 H13 110.881
C3 C4 H14 110.881 C4 C3 H10 109.442
C4 C3 H11 109.442 O5 C1 H6 111.327
O5 C1 H7 111.327 H6 C1 H7 107.253
H8 C2 H9 106.655 H10 C3 H11 106.279
H12 C4 H13 107.916 H12 C4 H14 107.916
H13 C4 H14 107.657
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability