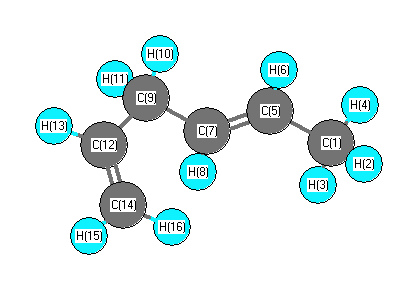
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3172 |
3153 |
13.90 |
|
|
|
| 2 |
A |
3079 |
3061 |
7.83 |
|
|
|
| 3 |
A |
3065 |
3047 |
22.62 |
|
|
|
| 4 |
A |
3062 |
3044 |
50.45 |
|
|
|
| 5 |
A |
3055 |
3037 |
6.27 |
|
|
|
| 6 |
A |
3044 |
3027 |
1.82 |
|
|
|
| 7 |
A |
3013 |
2995 |
15.54 |
|
|
|
| 8 |
A |
2962 |
2945 |
14.10 |
|
|
|
| 9 |
A |
2954 |
2937 |
33.13 |
|
|
|
| 10 |
A |
2925 |
2908 |
28.22 |
|
|
|
| 11 |
A |
1703 |
1693 |
1.66 |
|
|
|
| 12 |
A |
1667 |
1657 |
16.17 |
|
|
|
| 13 |
A |
1412 |
1404 |
10.57 |
|
|
|
| 14 |
A |
1399 |
1391 |
5.81 |
|
|
|
| 15 |
A |
1383 |
1375 |
13.55 |
|
|
|
| 16 |
A |
1372 |
1364 |
0.96 |
|
|
|
| 17 |
A |
1343 |
1335 |
1.39 |
|
|
|
| 18 |
A |
1308 |
1301 |
0.12 |
|
|
|
| 19 |
A |
1275 |
1268 |
2.29 |
|
|
|
| 20 |
A |
1272 |
1265 |
0.75 |
|
|
|
| 21 |
A |
1250 |
1243 |
4.45 |
|
|
|
| 22 |
A |
1172 |
1165 |
1.24 |
|
|
|
| 23 |
A |
1100 |
1093 |
6.51 |
|
|
|
| 24 |
A |
1087 |
1080 |
6.56 |
|
|
|
| 25 |
A |
1015 |
1009 |
5.06 |
|
|
|
| 26 |
A |
1012 |
1006 |
0.03 |
|
|
|
| 27 |
A |
995 |
989 |
8.57 |
|
|
|
| 28 |
A |
967 |
961 |
24.46 |
|
|
|
| 29 |
A |
920 |
915 |
13.68 |
|
|
|
| 30 |
A |
908 |
903 |
2.10 |
|
|
|
| 31 |
A |
893 |
888 |
22.65 |
|
|
|
| 32 |
A |
863 |
858 |
2.83 |
|
|
|
| 33 |
A |
753 |
748 |
0.25 |
|
|
|
| 34 |
A |
578 |
575 |
3.94 |
|
|
|
| 35 |
A |
528 |
525 |
6.37 |
|
|
|
| 36 |
A |
455 |
452 |
0.23 |
|
|
|
| 37 |
A |
306 |
304 |
0.30 |
|
|
|
| 38 |
A |
274 |
272 |
1.90 |
|
|
|
| 39 |
A |
212 |
211 |
1.33 |
|
|
|
| 40 |
A |
156 |
155 |
0.59 |
|
|
|
| 41 |
A |
132 |
131 |
0.05 |
|
|
|
| 42 |
A |
74 |
74 |
0.29 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 30055.4 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 29881.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.016 |
|
|
|
| 2 |
H |
0.047 |
|
|
|
| 3 |
H |
0.029 |
|
|
|
| 4 |
H |
0.046 |
|
|
|
| 5 |
C |
-0.078 |
|
|
|
| 6 |
H |
-0.017 |
|
|
|
| 7 |
C |
-0.049 |
|
|
|
| 8 |
H |
-0.015 |
|
|
|
| 9 |
C |
0.048 |
|
|
|
| 10 |
H |
0.031 |
|
|
|
| 11 |
H |
0.042 |
|
|
|
| 12 |
C |
-0.105 |
|
|
|
| 13 |
H |
-0.008 |
|
|
|
| 14 |
C |
0.003 |
|
|
|
| 15 |
H |
0.021 |
|
|
|
| 16 |
H |
0.021 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.394 |
-0.318 |
-0.038 |
0.507 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-36.523 |
1.378 |
-0.434 |
| y |
1.378 |
-37.822 |
0.627 |
| z |
-0.434 |
0.627 |
-39.188 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.982 |
1.378 |
-0.434 |
| y |
1.378 |
0.034 |
0.627 |
| z |
-0.434 |
0.627 |
-2.015 |
|
| Polar |
|---|
| 3z2-r2 | -4.031 |
|---|
| x2-y2 | 1.299 |
|---|
| xy | 1.378 |
|---|
| xz | -0.434 |
|---|
| yz | 0.627 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
13.379 |
0.371 |
-0.748 |
| y |
0.371 |
10.087 |
0.339 |
| z |
-0.748 |
0.339 |
7.496 |
<r2> (average value of r
2) Å
2
| <r2> |
237.654 |
| (<r2>)1/2 |
15.416 |