Jump to
S1C2
Energy calculated at QCISD/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -79.575006 |
| Energy at 298.15K | -79.580937 |
| HF Energy | -79.234269 |
| Nuclear repulsion energy | 41.879596 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1g |
3061 |
2936 |
0.00 |
|
|
|
| 2 |
A1g |
1437 |
1379 |
0.00 |
|
|
|
| 3 |
A1g |
1032 |
990 |
0.00 |
|
|
|
| 4 |
A1u |
327 |
313 |
0.00 |
|
|
|
| 5 |
A2u |
3055 |
2931 |
57.38 |
|
|
|
| 6 |
A2u |
1405 |
1348 |
0.00 |
|
|
|
| 7 |
Eg |
3126 |
2999 |
0.00 |
|
|
|
| 7 |
Eg |
3126 |
2999 |
0.00 |
|
|
|
| 8 |
Eg |
1501 |
1440 |
0.00 |
|
|
|
| 8 |
Eg |
1501 |
1440 |
0.00 |
|
|
|
| 9 |
Eg |
1225 |
1176 |
0.00 |
|
|
|
| 9 |
Eg |
1225 |
1176 |
0.00 |
|
|
|
| 10 |
Eu |
3148 |
3020 |
65.59 |
|
|
|
| 10 |
Eu |
3148 |
3020 |
65.59 |
|
|
|
| 11 |
Eu |
1504 |
1442 |
6.13 |
|
|
|
| 11 |
Eu |
1504 |
1442 |
6.13 |
|
|
|
| 12 |
Eu |
822 |
789 |
2.57 |
|
|
|
| 12 |
Eu |
822 |
789 |
2.57 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16483.9 cm
-1
Scaled (by 0.9594) Zero Point Vibrational Energy (zpe) 15814.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD/cc-pVDZ
Point Group is D3d
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.767 |
| C2 |
0.000 |
0.000 |
-0.767 |
| H3 |
0.000 |
1.030 |
1.167 |
| H4 |
-0.892 |
-0.515 |
1.167 |
| H5 |
0.892 |
-0.515 |
1.167 |
| H6 |
0.000 |
-1.030 |
-1.167 |
| H7 |
-0.892 |
0.515 |
-1.167 |
| H8 |
0.892 |
0.515 |
-1.167 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
H3 |
H4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5338 | 1.1054 | 1.1054 | 1.1054 | 2.1915 | 2.1915 | 2.1915 |
C2 | 1.5338 | | 2.1915 | 2.1915 | 2.1915 | 1.1054 | 1.1054 | 1.1054 | H3 | 1.1054 | 2.1915 | | 1.7847 | 1.7847 | 3.1140 | 2.5518 | 2.5518 | H4 | 1.1054 | 2.1915 | 1.7847 | | 1.7847 | 2.5518 | 2.5518 | 3.1140 | H5 | 1.1054 | 2.1915 | 1.7847 | 1.7847 | | 2.5518 | 3.1140 | 2.5518 | H6 | 2.1915 | 1.1054 | 3.1140 | 2.5518 | 2.5518 | | 1.7847 | 1.7847 | H7 | 2.1915 | 1.1054 | 2.5518 | 2.5518 | 3.1140 | 1.7847 | | 1.7847 | H8 | 2.1915 | 1.1054 | 2.5518 | 3.1140 | 2.5518 | 1.7847 | 1.7847 | |
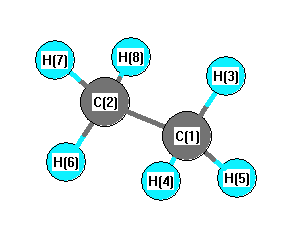 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H6 |
111.234 |
|
C1 |
C2 |
H7 |
111.234 |
| C1 |
C2 |
H8 |
111.234 |
|
C2 |
C1 |
H3 |
111.234 |
| C2 |
C1 |
H4 |
111.234 |
|
C2 |
C1 |
H5 |
111.234 |
| H3 |
C1 |
H4 |
107.652 |
|
H3 |
C1 |
H5 |
107.652 |
| H4 |
C1 |
H5 |
107.652 |
|
H6 |
C2 |
H7 |
107.652 |
| H6 |
C2 |
H8 |
107.652 |
|
H7 |
C2 |
H8 |
107.652 |
Electronic energy levels
Electronic state
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at QCISD/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -79.570033 |
| Energy at 298.15K | |
| HF Energy | -79.229016 |
| Nuclear repulsion energy | 41.708968 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1' |
3072 |
2947 |
0.00 |
|
|
|
| 2 |
A1' |
1449 |
1390 |
0.00 |
|
|
|
| 3 |
A1' |
1028 |
987 |
0.00 |
|
|
|
| 4 |
A1" |
318i |
305i |
0.00 |
|
|
|
| 5 |
A2" |
3060 |
2936 |
59.77 |
|
|
|
| 6 |
A2" |
1401 |
1344 |
0.04 |
|
|
|
| 7 |
E' |
3151 |
3023 |
54.81 |
|
|
|
| 7 |
E' |
3151 |
3023 |
54.81 |
|
|
|
| 8 |
E' |
1506 |
1445 |
7.83 |
|
|
|
| 8 |
E' |
1506 |
1445 |
7.83 |
|
|
|
| 9 |
E' |
895 |
858 |
2.09 |
|
|
|
| 9 |
E' |
895 |
858 |
2.08 |
|
|
|
| 10 |
E" |
3131 |
3004 |
0.00 |
|
|
|
| 10 |
E" |
3131 |
3004 |
0.00 |
|
|
|
| 11 |
E" |
1497 |
1436 |
0.00 |
|
|
|
| 11 |
E" |
1497 |
1436 |
0.00 |
|
|
|
| 12 |
E" |
1174 |
1126 |
0.00 |
|
|
|
| 12 |
E" |
1174 |
1126 |
0.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16199.9 cm
-1
Scaled (by 0.9594) Zero Point Vibrational Energy (zpe) 15542.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD/cc-pVDZ
Point Group is D3h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.774 |
| C2 |
0.000 |
0.000 |
-0.774 |
| H3 |
0.000 |
1.026 |
1.183 |
| H4 |
-0.889 |
-0.513 |
1.183 |
| H5 |
0.889 |
-0.513 |
1.183 |
| H6 |
0.000 |
1.026 |
-1.183 |
| H7 |
0.889 |
-0.513 |
-1.183 |
| H8 |
-0.889 |
-0.513 |
-1.183 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
H3 |
H4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5477 | 1.1046 | 1.1046 | 1.1046 | 2.2094 | 2.2094 | 2.2094 |
C2 | 1.5477 | | 2.2094 | 2.2094 | 2.2094 | 1.1046 | 1.1046 | 1.1046 | H3 | 1.1046 | 2.2094 | | 1.7771 | 1.7771 | 2.3658 | 2.9589 | 2.9589 | H4 | 1.1046 | 2.2094 | 1.7771 | | 1.7771 | 2.9589 | 2.9589 | 2.3658 | H5 | 1.1046 | 2.2094 | 1.7771 | 1.7771 | | 2.9589 | 2.3658 | 2.9589 | H6 | 2.2094 | 1.1046 | 2.3658 | 2.9589 | 2.9589 | | 1.7771 | 1.7771 | H7 | 2.2094 | 1.1046 | 2.9589 | 2.9589 | 2.3658 | 1.7771 | | 1.7771 | H8 | 2.2094 | 1.1046 | 2.9589 | 2.3658 | 2.9589 | 1.7771 | 1.7771 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H6 |
111.737 |
|
C1 |
C2 |
H7 |
111.737 |
| C1 |
C2 |
H8 |
111.737 |
|
C2 |
C1 |
H3 |
111.737 |
| C2 |
C1 |
H4 |
111.737 |
|
C2 |
C1 |
H5 |
111.737 |
| H3 |
C1 |
H4 |
107.114 |
|
H3 |
C1 |
H5 |
107.114 |
| H4 |
C1 |
H5 |
107.114 |
|
H6 |
C2 |
H7 |
107.114 |
| H6 |
C2 |
H8 |
107.114 |
|
H7 |
C2 |
H8 |
107.114 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
