Jump to
S1C2
Energy calculated at PBEPBEultrafine/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -79.699365 |
| Energy at 298.15K | -79.705228 |
| HF Energy | -79.699365 |
| Nuclear repulsion energy | 41.825329 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBEPBEultrafine/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1g |
2971 |
2954 |
0.00 |
271.70 |
0.00 |
0.01 |
| 2 |
A1g |
1361 |
1353 |
0.00 |
5.81 |
0.71 |
0.83 |
| 3 |
A1g |
1005 |
1000 |
0.00 |
8.43 |
0.28 |
0.44 |
| 4 |
A1u |
312 |
310 |
0.00 |
0.00 |
0.00 |
0.00 |
| 5 |
A2u |
2970 |
2953 |
58.55 |
0.00 |
0.00 |
0.00 |
| 6 |
A2u |
1336 |
1328 |
0.00 |
0.00 |
0.00 |
0.00 |
| 7 |
Eg |
3035 |
3017 |
0.00 |
155.89 |
0.75 |
0.86 |
| 7 |
Eg |
3035 |
3017 |
0.00 |
155.89 |
0.75 |
0.86 |
| 8 |
Eg |
1428 |
1419 |
0.00 |
31.38 |
0.75 |
0.86 |
| 8 |
Eg |
1428 |
1419 |
0.00 |
31.38 |
0.75 |
0.86 |
| 9 |
Eg |
1162 |
1155 |
0.00 |
0.78 |
0.75 |
0.86 |
| 9 |
Eg |
1162 |
1155 |
0.00 |
0.78 |
0.75 |
0.86 |
| 10 |
Eu |
3059 |
3041 |
53.68 |
0.00 |
0.00 |
0.00 |
| 10 |
Eu |
3059 |
3041 |
53.68 |
0.00 |
0.00 |
0.00 |
| 11 |
Eu |
1428 |
1420 |
8.02 |
0.00 |
0.00 |
0.00 |
| 11 |
Eu |
1428 |
1420 |
8.02 |
0.00 |
0.00 |
0.00 |
| 12 |
Eu |
797 |
792 |
5.62 |
0.00 |
0.00 |
0.00 |
| 12 |
Eu |
797 |
792 |
5.62 |
0.00 |
0.00 |
0.00 |
Unscaled Zero Point Vibrational Energy (zpe) 15885.8 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 15793.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBEPBEultrafine/cc-pVDZ
Point Group is D3d
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.765 |
| C2 |
0.000 |
0.000 |
-0.765 |
| H3 |
0.000 |
1.032 |
1.173 |
| H4 |
-0.894 |
-0.516 |
1.173 |
| H5 |
0.894 |
-0.516 |
1.173 |
| H6 |
0.000 |
-1.032 |
-1.173 |
| H7 |
-0.894 |
0.516 |
-1.173 |
| H8 |
0.894 |
0.516 |
-1.173 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
H3 |
H4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5294 | 1.1096 | 1.1096 | 1.1096 | 2.1954 | 2.1954 | 2.1954 |
C2 | 1.5294 | | 2.1954 | 2.1954 | 2.1954 | 1.1096 | 1.1096 | 1.1096 | H3 | 1.1096 | 2.1954 | | 1.7870 | 1.7870 | 3.1246 | 2.5631 | 2.5631 | H4 | 1.1096 | 2.1954 | 1.7870 | | 1.7870 | 2.5631 | 2.5631 | 3.1246 | H5 | 1.1096 | 2.1954 | 1.7870 | 1.7870 | | 2.5631 | 3.1246 | 2.5631 | H6 | 2.1954 | 1.1096 | 3.1246 | 2.5631 | 2.5631 | | 1.7870 | 1.7870 | H7 | 2.1954 | 1.1096 | 2.5631 | 2.5631 | 3.1246 | 1.7870 | | 1.7870 | H8 | 2.1954 | 1.1096 | 2.5631 | 3.1246 | 2.5631 | 1.7870 | 1.7870 | |
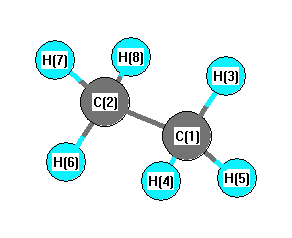 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H6 |
111.597 |
|
C1 |
C2 |
H7 |
111.597 |
| C1 |
C2 |
H8 |
111.597 |
|
C2 |
C1 |
H3 |
111.597 |
| C2 |
C1 |
H4 |
111.597 |
|
C2 |
C1 |
H5 |
111.597 |
| H3 |
C1 |
H4 |
107.264 |
|
H3 |
C1 |
H5 |
107.264 |
| H4 |
C1 |
H5 |
107.264 |
|
H6 |
C2 |
H7 |
107.264 |
| H6 |
C2 |
H8 |
107.264 |
|
H7 |
C2 |
H8 |
107.264 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.104 |
|
|
|
| 2 |
C |
-0.104 |
|
|
|
| 3 |
H |
0.035 |
|
|
|
| 4 |
H |
0.035 |
|
|
|
| 5 |
H |
0.035 |
|
|
|
| 6 |
H |
0.035 |
|
|
|
| 7 |
H |
0.035 |
|
|
|
| 8 |
H |
0.035 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-14.855 |
0.000 |
0.000 |
| y |
0.000 |
-14.855 |
0.000 |
| z |
0.000 |
0.000 |
-15.375 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.260 |
0.000 |
0.000 |
| y |
0.000 |
0.260 |
0.000 |
| z |
0.000 |
0.000 |
-0.520 |
|
| Polar |
|---|
| 3z2-r2 | -1.041 |
|---|
| x2-y2 | 0.000 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.609 |
0.000 |
0.000 |
| y |
0.000 |
3.609 |
0.000 |
| z |
0.000 |
0.000 |
4.029 |
<r2> (average value of r
2) Å
2
| <r2> |
31.048 |
| (<r2>)1/2 |
5.572 |
Jump to
S1C1
Energy calculated at PBEPBEultrafine/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -79.694811 |
| Energy at 298.15K | |
| HF Energy | -79.694811 |
| Nuclear repulsion energy | 41.664154 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBEPBEultrafine/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1' |
2986 |
2968 |
0.00 |
|
|
|
| 2 |
A1' |
1370 |
1362 |
0.00 |
|
|
|
| 3 |
A1' |
1001 |
995 |
0.00 |
|
|
|
| 4 |
A1" |
304i |
302i |
0.00 |
|
|
|
| 5 |
A2" |
2978 |
2960 |
60.04 |
|
|
|
| 6 |
A2" |
1332 |
1324 |
0.00 |
|
|
|
| 7 |
E' |
3064 |
3047 |
41.85 |
|
|
|
| 7 |
E' |
3064 |
3047 |
41.85 |
|
|
|
| 8 |
E' |
1430 |
1421 |
10.06 |
|
|
|
| 8 |
E' |
1430 |
1421 |
10.06 |
|
|
|
| 9 |
E' |
858 |
853 |
4.68 |
|
|
|
| 9 |
E' |
858 |
853 |
4.68 |
|
|
|
| 10 |
E" |
3044 |
3027 |
0.00 |
|
|
|
| 10 |
E" |
3044 |
3027 |
0.00 |
|
|
|
| 11 |
E" |
1426 |
1418 |
0.00 |
|
|
|
| 11 |
E" |
1426 |
1418 |
0.00 |
|
|
|
| 12 |
E" |
1113 |
1106 |
0.00 |
|
|
|
| 12 |
E" |
1113 |
1106 |
0.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 15615.7 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 15525.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBEPBEultrafine/cc-pVDZ
Point Group is D3h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.772 |
| C2 |
0.000 |
0.000 |
-0.772 |
| H3 |
0.000 |
1.027 |
1.188 |
| H4 |
-0.890 |
-0.514 |
1.188 |
| H5 |
0.890 |
-0.514 |
1.188 |
| H6 |
0.000 |
1.027 |
-1.188 |
| H7 |
0.890 |
-0.514 |
-1.188 |
| H8 |
-0.890 |
-0.514 |
-1.188 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
H3 |
H4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5430 | 1.1085 | 1.1085 | 1.1085 | 2.2126 | 2.2126 | 2.2126 |
C2 | 1.5430 | | 2.2126 | 2.2126 | 2.2126 | 1.1085 | 1.1085 | 1.1085 | H3 | 1.1085 | 2.2126 | | 1.7792 | 1.7792 | 2.3764 | 2.9686 | 2.9686 | H4 | 1.1085 | 2.2126 | 1.7792 | | 1.7792 | 2.9686 | 2.9686 | 2.3764 | H5 | 1.1085 | 2.2126 | 1.7792 | 1.7792 | | 2.9686 | 2.3764 | 2.9686 | H6 | 2.2126 | 1.1085 | 2.3764 | 2.9686 | 2.9686 | | 1.7792 | 1.7792 | H7 | 2.2126 | 1.1085 | 2.9686 | 2.9686 | 2.3764 | 1.7792 | | 1.7792 | H8 | 2.2126 | 1.1085 | 2.9686 | 2.3764 | 2.9686 | 1.7792 | 1.7792 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H6 |
112.080 |
|
C1 |
C2 |
H7 |
112.080 |
| C1 |
C2 |
H8 |
112.080 |
|
C2 |
C1 |
H3 |
112.080 |
| C2 |
C1 |
H4 |
112.080 |
|
C2 |
C1 |
H5 |
112.080 |
| H3 |
C1 |
H4 |
106.741 |
|
H3 |
C1 |
H5 |
106.741 |
| H4 |
C1 |
H5 |
106.741 |
|
H6 |
C2 |
H7 |
106.741 |
| H6 |
C2 |
H8 |
106.741 |
|
H7 |
C2 |
H8 |
106.741 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.105 |
|
|
|
| 2 |
C |
-0.105 |
|
|
|
| 3 |
H |
0.035 |
|
|
|
| 4 |
H |
0.035 |
|
|
|
| 5 |
H |
0.035 |
|
|
|
| 6 |
H |
0.035 |
|
|
|
| 7 |
H |
0.035 |
|
|
|
| 8 |
H |
0.035 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-14.844 |
0.000 |
0.000 |
| y |
0.000 |
-14.844 |
0.000 |
| z |
0.000 |
0.000 |
-15.367 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.261 |
0.000 |
0.000 |
| y |
0.000 |
0.261 |
0.000 |
| z |
0.000 |
0.000 |
-0.523 |
|
| Polar |
|---|
| 3z2-r2 | -1.046 |
|---|
| x2-y2 | 0.000 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.530 |
0.000 |
0.000 |
| y |
0.000 |
3.530 |
0.000 |
| z |
0.000 |
0.000 |
3.969 |
<r2> (average value of r
2) Å
2
| <r2> |
31.325 |
| (<r2>)1/2 |
5.597 |
