return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3NH2 (methyl amine)

using model chemistry: CISD/cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at CISD/cc-pVDZ
 hartrees
Energy at 0K-95.548057
Energy at 298.15K 
HF Energy-95.221249
Nuclear repulsion energy41.909143
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CISD/cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3575 3296 0.97      
2 A' 3154 2908 46.73      
3 A' 3062 2822 80.00      
4 A' 1702 1569 17.30      
5 A' 1528 1408 7.08      
6 A' 1493 1376 3.67      
7 A' 1221 1126 12.69      
8 A' 1112 1025 12.46      
9 A' 924 851 125.80      
10 A" 3652 3367 0.01      
11 A" 3195 2945 35.34      
12 A" 1546 1425 1.85      
13 A" 1395 1286 0.15      
14 A" 1002 924 0.15      
15 A" 343 316 37.01      

Unscaled Zero Point Vibrational Energy (zpe) 14451.9 cm-1
Scaled (by 0.9218) Zero Point Vibrational Energy (zpe) 13321.7 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at CISD/cc-pVDZ
ABC
3.40295 0.75961 0.73251

See section I.F.4 to change rotational constant units
Geometric Data calculated at CISD/cc-pVDZ

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.054 0.701 0.000
N2 0.054 -0.762 0.000
H3 -0.946 1.176 0.000
H4 0.597 1.065 0.885
H5 0.597 1.065 -0.885
H6 -0.475 -1.089 -0.806
H7 -0.475 -1.089 0.806

Atom - Atom Distances (Å)
  C1 N2 H3 H4 H5 H6 H7
C11.46291.10691.09991.09992.03362.0336
N21.46292.18072.10122.10121.01831.0183
H31.10692.18071.78171.78172.45032.4503
H41.09992.10121.78171.76942.94102.4077
H51.09992.10121.78171.76942.40772.9410
H62.03361.01832.45032.94102.40771.6121
H72.03361.01832.45032.40772.94101.6121

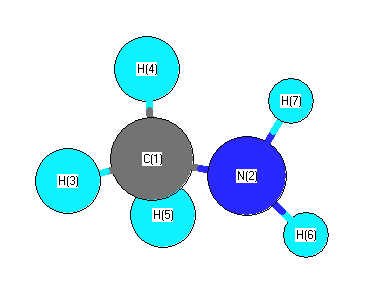
picture of methyl amine state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 N2 H6 108.760 C1 N2 H7 108.760
N2 C1 H3 115.420 N2 C1 H4 109.327
N2 C1 H5 109.327 H3 C1 H4 107.673
H3 C1 H5 107.673 H4 C1 H5 107.096
H6 N2 H7 104.659
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability