return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C3H8 (Propane)

using model chemistry: CCD/cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at CCD/cc-pVDZ
 hartrees
Energy at 0K-118.766408
Energy at 298.15K-118.774816
HF Energy-118.271629
Nuclear repulsion energy81.971536
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A1 3149 3013 45.49      
2 A1 3065 2933 10.36      
3 A1 3059 2927 37.84      
4 A1 1513 1448 2.96      
5 A1 1492 1427 0.12      
6 A1 1429 1367 1.30      
7 A1 1188 1136 0.71      
8 A1 901 862 0.66      
9 A1 368 352 0.04      
10 A2 3138 3002 0.00      
11 A2 1490 1426 0.00      
12 A2 1322 1264 0.00      
13 A2 910 871 0.00      
14 A2 229 219 0.00      
15 B1 3147 3010 89.54      
16 B1 3102 2968 6.20      
17 B1 1507 1442 10.37      
18 B1 1222 1169 0.09      
19 B1 754 722 1.97      
20 B1 286 274 0.00      
21 B2 3147 3010 26.10      
22 B2 3058 2925 34.01      
23 B2 1499 1434 1.06      
24 B2 1424 1362 3.91      
25 B2 1375 1315 3.26      
26 B2 1095 1048 0.21      
27 B2 934 894 0.99      

Unscaled Zero Point Vibrational Energy (zpe) 22900.3 cm-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 21908.7 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at CCD/cc-pVDZ
ABC
0.96724 0.27890 0.24688

See section I.F.4 to change rotational constant units
Geometric Data calculated at CCD/cc-pVDZ

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.000 0.591
C2 0.000 1.276 -0.261
C3 0.000 -1.276 -0.261
H4 0.886 0.000 1.255
H5 -0.886 0.000 1.255
H6 0.000 2.184 0.368
H7 0.000 -2.184 0.368
H8 0.893 1.315 -0.913
H9 -0.893 1.315 -0.913
H10 -0.893 -1.315 -0.913
H11 0.893 -1.315 -0.913

Atom - Atom Distances (Å)
  C1 C2 C3 H4 H5 H6 H7 H8 H9 H10 H11
C11.53431.53431.10721.10722.19512.19512.18812.18812.18812.1881
C21.53432.55172.17072.17071.10473.51641.10601.10602.81672.8167
C31.53432.55172.17072.17073.51641.10472.81672.81671.10601.1060
H41.10722.17072.17071.77162.51802.51802.53583.09733.09732.5358
H51.10722.17072.17071.77162.51802.51803.09732.53582.53583.0973
H62.19511.10473.51642.51802.51804.36751.78701.78703.83133.8313
H72.19513.51641.10472.51802.51804.36753.83133.83131.78701.7870
H82.18811.10602.81672.53583.09731.78703.83131.78543.17852.6298
H92.18811.10602.81673.09732.53581.78703.83131.78542.62983.1785
H102.18812.81671.10603.09732.53583.83131.78703.17852.62981.7854
H112.18812.81671.10602.53583.09733.83131.78702.62983.17851.7854

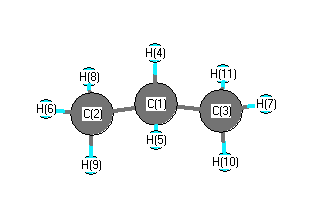
picture of Propane state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 H6 111.531 C1 C2 H8 110.898
C1 C2 H9 110.898 C1 C3 H7 111.531
C1 C3 H10 110.898 C1 C3 H11 110.898
C2 C1 C3 112.518 C2 C1 H4 109.464
C2 C1 H5 109.464 C3 C1 H4 109.464
C3 C1 H5 109.464 H4 C1 H5 106.273
H6 C2 H8 107.863 H6 C2 H9 107.863
H7 C3 H10 107.863 H7 C3 H11 107.863
H8 C2 H9 107.626 H10 C3 H11 107.626
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability