Jump to
S1C2
Energy calculated at B3PW91/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -135.131502 |
| Energy at 298.15K | -135.139692 |
| Nuclear repulsion energy | 82.597382 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3PW91/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3468 |
3347 |
1.75 |
|
|
|
| 2 |
A' |
3112 |
3003 |
40.03 |
|
|
|
| 3 |
A' |
3036 |
2930 |
37.12 |
|
|
|
| 4 |
A' |
3026 |
2920 |
28.46 |
|
|
|
| 5 |
A' |
1650 |
1592 |
15.80 |
|
|
|
| 6 |
A' |
1477 |
1425 |
1.97 |
|
|
|
| 7 |
A' |
1458 |
1407 |
0.04 |
|
|
|
| 8 |
A' |
1391 |
1343 |
13.20 |
|
|
|
| 9 |
A' |
1363 |
1315 |
13.21 |
|
|
|
| 10 |
A' |
1160 |
1120 |
7.96 |
|
|
|
| 11 |
A' |
1082 |
1044 |
31.98 |
|
|
|
| 12 |
A' |
904 |
872 |
68.45 |
|
|
|
| 13 |
A' |
882 |
851 |
87.61 |
|
|
|
| 14 |
A' |
400 |
386 |
5.39 |
|
|
|
| 15 |
A" |
3546 |
3422 |
0.12 |
|
|
|
| 16 |
A" |
3112 |
3004 |
50.90 |
|
|
|
| 17 |
A" |
3073 |
2966 |
11.05 |
|
|
|
| 18 |
A" |
1464 |
1413 |
8.13 |
|
|
|
| 19 |
A" |
1386 |
1337 |
0.00 |
|
|
|
| 20 |
A" |
1260 |
1216 |
0.07 |
|
|
|
| 21 |
A" |
1005 |
970 |
1.24 |
|
|
|
| 22 |
A" |
777 |
750 |
1.74 |
|
|
|
| 23 |
A" |
313 |
302 |
35.71 |
|
|
|
| 24 |
A" |
258 |
249 |
8.98 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 20301.8 cm
-1
Scaled (by 0.965) Zero Point Vibrational Energy (zpe) 19591.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3PW91/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| N1 |
-1.310 |
-0.073 |
0.000 |
| C2 |
0.000 |
0.571 |
0.000 |
| C3 |
1.209 |
-0.363 |
0.000 |
| H4 |
2.159 |
0.199 |
0.000 |
| H5 |
1.208 |
-1.015 |
0.890 |
| H6 |
1.208 |
-1.015 |
-0.890 |
| H7 |
0.047 |
1.237 |
-0.879 |
| H8 |
0.047 |
1.237 |
0.879 |
| H9 |
-1.377 |
-0.694 |
0.810 |
| H10 |
-1.377 |
-0.694 |
-0.810 |
Atom - Atom Distances (Å)
| |
N1 |
C2 |
C3 |
H4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| N1 | | 1.4596 | 2.5359 | 3.4792 | 2.8319 | 2.8319 | 2.0812 | 2.0812 | 1.0226 | 1.0226 |
C2 | 1.4596 | | 1.5277 | 2.1903 | 2.1832 | 2.1832 | 1.1042 | 1.1042 | 2.0376 | 2.0376 | C3 | 2.5359 | 1.5277 | | 1.1031 | 1.1036 | 1.1036 | 2.1639 | 2.1639 | 2.7304 | 2.7304 | H4 | 3.4792 | 2.1903 | 1.1031 | | 1.7806 | 1.7806 | 2.5114 | 2.5114 | 3.7355 | 3.7355 | H5 | 2.8319 | 2.1832 | 1.1036 | 1.7806 | | 1.7802 | 3.0902 | 2.5335 | 2.6061 | 3.1104 | H6 | 2.8319 | 2.1832 | 1.1036 | 1.7806 | 1.7802 | | 2.5335 | 3.0902 | 3.1104 | 2.6061 | H7 | 2.0812 | 1.1042 | 2.1639 | 2.5114 | 3.0902 | 2.5335 | | 1.7585 | 2.9344 | 2.4006 | H8 | 2.0812 | 1.1042 | 2.1639 | 2.5114 | 2.5335 | 3.0902 | 1.7585 | | 2.4006 | 2.9344 | H9 | 1.0226 | 2.0376 | 2.7304 | 3.7355 | 2.6061 | 3.1104 | 2.9344 | 2.4006 | | 1.6193 | H10 | 1.0226 | 2.0376 | 2.7304 | 3.7355 | 3.1104 | 2.6061 | 2.4006 | 2.9344 | 1.6193 | |
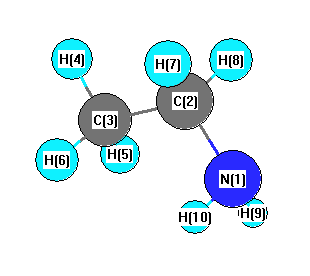 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| N1 |
C2 |
C3 |
116.165 |
|
N1 |
C2 |
H7 |
107.736 |
| N1 |
C2 |
H8 |
107.736 |
|
C2 |
N1 |
H9 |
109.075 |
| C2 |
N1 |
H10 |
109.075 |
|
C2 |
C3 |
H4 |
111.709 |
| C2 |
C3 |
H5 |
111.111 |
|
C2 |
C3 |
H6 |
111.111 |
| C3 |
C2 |
H7 |
109.555 |
|
C3 |
C2 |
H8 |
109.555 |
| H4 |
C3 |
H5 |
107.592 |
|
H4 |
C3 |
H6 |
107.592 |
| H5 |
C3 |
H6 |
107.528 |
|
H7 |
C2 |
H8 |
105.557 |
| H9 |
N1 |
H10 |
104.695 |
|
Electronic energy levels
Electronic state
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.217 |
|
|
|
| 2 |
C |
-0.035 |
|
|
|
| 3 |
C |
-0.105 |
|
|
|
| 4 |
H |
0.036 |
|
|
|
| 5 |
H |
0.036 |
|
|
|
| 6 |
H |
0.036 |
|
|
|
| 7 |
H |
0.042 |
|
|
|
| 8 |
H |
0.042 |
|
|
|
| 9 |
H |
0.083 |
|
|
|
| 10 |
H |
0.083 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.802 |
-0.956 |
0.000 |
1.248 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
58.533 |
| (<r2>)1/2 |
7.651 |
Jump to
S1C1
Energy calculated at B3PW91/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -135.130707 |
| Energy at 298.15K | |
| Nuclear repulsion energy | |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3PW91/cc-pVDZ
Geometric Data calculated at B3PW91/cc-pVDZ
Point Group is Cs
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
