Jump to
S1C2
Energy calculated at QCISD/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -134.775809 |
| Energy at 298.15K | -134.784057 |
| HF Energy | -134.261018 |
| Nuclear repulsion energy | 82.322181 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3475 |
3334 |
2.63 |
|
|
|
| 2 |
A' |
3129 |
3002 |
42.50 |
|
|
|
| 3 |
A' |
3060 |
2936 |
35.25 |
|
|
|
| 4 |
A' |
3042 |
2919 |
27.71 |
|
|
|
| 5 |
A' |
1671 |
1603 |
15.23 |
|
|
|
| 6 |
A' |
1506 |
1445 |
0.94 |
|
|
|
| 7 |
A' |
1487 |
1427 |
0.15 |
|
|
|
| 8 |
A' |
1420 |
1362 |
11.57 |
|
|
|
| 9 |
A' |
1386 |
1330 |
15.40 |
|
|
|
| 10 |
A' |
1180 |
1132 |
9.80 |
|
|
|
| 11 |
A' |
1101 |
1057 |
32.05 |
|
|
|
| 12 |
A' |
946 |
907 |
115.83 |
|
|
|
| 13 |
A' |
899 |
863 |
14.13 |
|
|
|
| 14 |
A' |
404 |
388 |
5.88 |
|
|
|
| 15 |
A" |
3556 |
3411 |
1.00 |
|
|
|
| 16 |
A" |
3129 |
3002 |
61.92 |
|
|
|
| 17 |
A" |
3099 |
2973 |
7.64 |
|
|
|
| 18 |
A" |
1493 |
1432 |
6.51 |
|
|
|
| 19 |
A" |
1406 |
1349 |
0.02 |
|
|
|
| 20 |
A" |
1281 |
1229 |
0.26 |
|
|
|
| 21 |
A" |
1018 |
976 |
1.56 |
|
|
|
| 22 |
A" |
782 |
750 |
0.32 |
|
|
|
| 23 |
A" |
325 |
312 |
37.71 |
|
|
|
| 24 |
A" |
266 |
255 |
8.44 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 20529.6 cm
-1
Scaled (by 0.9594) Zero Point Vibrational Energy (zpe) 19696.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| N1 |
-1.315 |
-0.076 |
0.000 |
| C2 |
0.000 |
0.580 |
0.000 |
| C3 |
1.210 |
-0.364 |
0.000 |
| H4 |
2.164 |
0.197 |
0.000 |
| H5 |
1.199 |
-1.017 |
0.894 |
| H6 |
1.199 |
-1.017 |
-0.894 |
| H7 |
0.046 |
1.243 |
-0.885 |
| H8 |
0.046 |
1.243 |
0.885 |
| H9 |
-1.354 |
-0.706 |
0.809 |
| H10 |
-1.354 |
-0.706 |
-0.809 |
Atom - Atom Distances (Å)
| |
N1 |
C2 |
C3 |
H4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| N1 | | 1.4700 | 2.5416 | 3.4895 | 2.8291 | 2.8291 | 2.0923 | 2.0923 | 1.0257 | 1.0257 |
C2 | 1.4700 | | 1.5353 | 2.1974 | 2.1881 | 2.1881 | 1.1068 | 1.1068 | 2.0354 | 2.0354 | C3 | 2.5416 | 1.5353 | | 1.1064 | 1.1070 | 1.1070 | 2.1733 | 2.1733 | 2.7105 | 2.7105 | H4 | 3.4895 | 2.1974 | 1.1064 | | 1.7897 | 1.7897 | 2.5225 | 2.5225 | 3.7209 | 3.7209 | H5 | 2.8291 | 2.1881 | 1.1070 | 1.7897 | | 1.7889 | 3.0989 | 2.5370 | 2.5733 | 3.0847 | H6 | 2.8291 | 2.1881 | 1.1070 | 1.7897 | 1.7889 | | 2.5370 | 3.0989 | 3.0847 | 2.5733 | H7 | 2.0923 | 1.1068 | 2.1733 | 2.5225 | 3.0989 | 2.5370 | | 1.7703 | 2.9377 | 2.4014 | H8 | 2.0923 | 1.1068 | 2.1733 | 2.5225 | 2.5370 | 3.0989 | 1.7703 | | 2.4014 | 2.9377 | H9 | 1.0257 | 2.0354 | 2.7105 | 3.7209 | 2.5733 | 3.0847 | 2.9377 | 2.4014 | | 1.6174 | H10 | 1.0257 | 2.0354 | 2.7105 | 3.7209 | 3.0847 | 2.5733 | 2.4014 | 2.9377 | 1.6174 | |
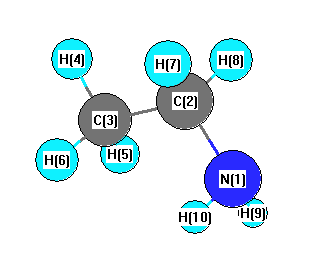 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| N1 |
C2 |
C3 |
115.480 |
|
N1 |
C2 |
H7 |
107.749 |
| N1 |
C2 |
H8 |
107.749 |
|
C2 |
N1 |
H9 |
107.967 |
| C2 |
N1 |
H10 |
107.967 |
|
C2 |
C3 |
H4 |
111.545 |
| C2 |
C3 |
H5 |
110.763 |
|
C2 |
C3 |
H6 |
110.763 |
| C3 |
C2 |
H7 |
109.619 |
|
C3 |
C2 |
H8 |
109.619 |
| H4 |
C3 |
H5 |
107.915 |
|
H4 |
C3 |
H6 |
107.915 |
| H5 |
C3 |
H6 |
107.793 |
|
H7 |
C2 |
H8 |
106.207 |
| H9 |
N1 |
H10 |
104.086 |
|
Electronic energy levels
Electronic state
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at QCISD/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -134.775281 |
| Energy at 298.15K | |
| Nuclear repulsion energy | |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/cc-pVDZ
Geometric Data calculated at QCISD/cc-pVDZ
Point Group is Cs
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
