Jump to
S1C2
Energy calculated at PBEPBE/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -134.985860 |
| Energy at 298.15K | -134.993984 |
| Nuclear repulsion energy | 82.109350 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3347 |
3328 |
4.76 |
|
|
|
| 2 |
A' |
3034 |
3017 |
41.17 |
|
|
|
| 3 |
A' |
2957 |
2940 |
36.07 |
|
|
|
| 4 |
A' |
2951 |
2934 |
29.90 |
|
|
|
| 5 |
A' |
1601 |
1592 |
12.66 |
|
|
|
| 6 |
A' |
1428 |
1420 |
1.47 |
|
|
|
| 7 |
A' |
1408 |
1399 |
0.01 |
|
|
|
| 8 |
A' |
1341 |
1333 |
10.11 |
|
|
|
| 9 |
A' |
1317 |
1309 |
15.47 |
|
|
|
| 10 |
A' |
1130 |
1123 |
6.36 |
|
|
|
| 11 |
A' |
1048 |
1042 |
28.01 |
|
|
|
| 12 |
A' |
884 |
879 |
93.98 |
|
|
|
| 13 |
A' |
857 |
852 |
56.53 |
|
|
|
| 14 |
A' |
389 |
386 |
5.16 |
|
|
|
| 15 |
A" |
3424 |
3404 |
1.52 |
|
|
|
| 16 |
A" |
3038 |
3020 |
46.73 |
|
|
|
| 17 |
A" |
2993 |
2976 |
12.75 |
|
|
|
| 18 |
A" |
1416 |
1408 |
7.98 |
|
|
|
| 19 |
A" |
1346 |
1338 |
0.01 |
|
|
|
| 20 |
A" |
1222 |
1215 |
0.12 |
|
|
|
| 21 |
A" |
977 |
972 |
1.13 |
|
|
|
| 22 |
A" |
757 |
753 |
1.79 |
|
|
|
| 23 |
A" |
311 |
310 |
34.76 |
|
|
|
| 24 |
A" |
257 |
256 |
6.71 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 19716.1 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 19601.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBEPBE/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| N1 |
-1.320 |
-0.071 |
0.000 |
| C2 |
0.000 |
0.574 |
0.000 |
| C3 |
1.215 |
-0.364 |
0.000 |
| H4 |
2.173 |
0.199 |
0.000 |
| H5 |
1.212 |
-1.021 |
0.896 |
| H6 |
1.212 |
-1.021 |
-0.896 |
| H7 |
0.048 |
1.245 |
-0.886 |
| H8 |
0.048 |
1.245 |
0.886 |
| H9 |
-1.371 |
-0.705 |
0.813 |
| H10 |
-1.371 |
-0.705 |
-0.813 |
Atom - Atom Distances (Å)
| |
N1 |
C2 |
C3 |
H4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| N1 | | 1.4684 | 2.5511 | 3.5025 | 2.8485 | 2.8485 | 2.0937 | 2.0937 | 1.0326 | 1.0326 |
C2 | 1.4684 | | 1.5344 | 2.2046 | 2.1942 | 2.1942 | 1.1124 | 1.1124 | 2.0437 | 2.0437 | C3 | 2.5511 | 1.5344 | | 1.1111 | 1.1113 | 1.1113 | 2.1758 | 2.1758 | 2.7322 | 2.7322 | H4 | 3.5025 | 2.2046 | 1.1111 | | 1.7932 | 1.7932 | 2.5286 | 2.5286 | 3.7467 | 3.7467 | H5 | 2.8485 | 2.1942 | 1.1113 | 1.7932 | | 1.7921 | 3.1088 | 2.5474 | 2.6036 | 3.1133 | H6 | 2.8485 | 2.1942 | 1.1113 | 1.7932 | 1.7921 | | 2.5474 | 3.1088 | 3.1133 | 2.6036 | H7 | 2.0937 | 1.1124 | 2.1758 | 2.5286 | 3.1088 | 2.5474 | | 1.7721 | 2.9498 | 2.4124 | H8 | 2.0937 | 1.1124 | 2.1758 | 2.5286 | 2.5474 | 3.1088 | 1.7721 | | 2.4124 | 2.9498 | H9 | 1.0326 | 2.0437 | 2.7322 | 3.7467 | 2.6036 | 3.1133 | 2.9498 | 2.4124 | | 1.6261 | H10 | 1.0326 | 2.0437 | 2.7322 | 3.7467 | 3.1133 | 2.6036 | 2.4124 | 2.9498 | 1.6261 | |
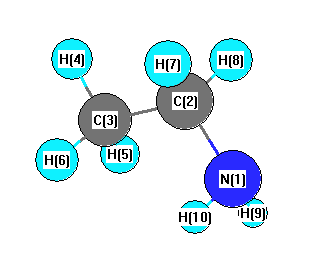 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| N1 |
C2 |
C3 |
116.320 |
|
N1 |
C2 |
H7 |
107.640 |
| N1 |
C2 |
H8 |
107.640 |
|
C2 |
N1 |
H9 |
108.339 |
| C2 |
N1 |
H10 |
108.339 |
|
C2 |
C3 |
H4 |
111.896 |
| C2 |
C3 |
H5 |
111.056 |
|
C2 |
C3 |
H6 |
111.056 |
| C3 |
C2 |
H7 |
109.552 |
|
C3 |
C2 |
H8 |
109.552 |
| H4 |
C3 |
H5 |
107.581 |
|
H4 |
C3 |
H6 |
107.581 |
| H5 |
C3 |
H6 |
107.468 |
|
H7 |
C2 |
H8 |
105.589 |
| H9 |
N1 |
H10 |
103.876 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.174 |
|
|
|
| 2 |
C |
-0.032 |
|
|
|
| 3 |
C |
-0.077 |
|
|
|
| 4 |
H |
0.025 |
|
|
|
| 5 |
H |
0.026 |
|
|
|
| 6 |
H |
0.026 |
|
|
|
| 7 |
H |
0.032 |
|
|
|
| 8 |
H |
0.032 |
|
|
|
| 9 |
H |
0.072 |
|
|
|
| 10 |
H |
0.072 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.748 |
-0.934 |
0.000 |
1.197 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-23.609 |
2.384 |
0.000 |
| y |
2.384 |
-20.270 |
0.000 |
| z |
0.000 |
0.000 |
-19.340 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-3.804 |
2.384 |
0.000 |
| y |
2.384 |
1.204 |
0.000 |
| z |
0.000 |
0.000 |
2.600 |
|
| Polar |
|---|
| 3z2-r2 | 5.200 |
|---|
| x2-y2 | -3.339 |
|---|
| xy | 2.384 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.331 |
0.189 |
0.000 |
| y |
0.189 |
4.764 |
0.000 |
| z |
0.000 |
0.000 |
4.655 |
<r2> (average value of r
2) Å
2
| <r2> |
59.144 |
| (<r2>)1/2 |
7.691 |
Jump to
S1C1
Energy calculated at PBEPBE/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -134.984738 |
| Energy at 298.15K | |
| Nuclear repulsion energy | |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
Geometric Data calculated at PBEPBE/cc-pVDZ
Point Group is Cs
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
