return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3CHO (Acetaldehyde)

using model chemistry: CCD/cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at CCD/cc-pVDZ
 hartrees
Energy at 0K-153.409880
Energy at 298.15K 
HF Energy-152.927692
Nuclear repulsion energy69.321049
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3195 3057 8.77      
2 A' 3070 2937 3.78      
3 A' 2951 2823 123.55      
4 A' 1875 1794 131.85      
5 A' 1466 1403 14.73      
6 A' 1444 1382 15.27      
7 A' 1389 1329 11.59      
8 A' 1140 1090 22.43      
9 A' 911 872 4.47      
10 A' 509 487 14.16      
11 A" 3151 3015 9.58      
12 A" 1477 1413 8.63      
13 A" 1142 1092 0.20      
14 A" 784 750 0.44      
15 A" 169 161 0.64      

Unscaled Zero Point Vibrational Energy (zpe) 12337.3 cm-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 11803.1 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at CCD/cc-pVDZ
ABC
1.87343 0.33541 0.30068

See section I.F.4 to change rotational constant units
Geometric Data calculated at CCD/cc-pVDZ

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
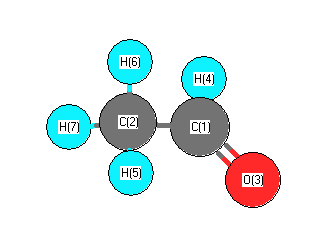
C1 0.000 0.466 0.000
C2 -0.935 -0.725 0.000
O3 1.207 0.390 0.000
H4 -0.509 1.465 0.000
H5 -0.356 -1.662 0.000
H6 -1.589 -0.684 0.890
H7 -1.589 -0.684 -0.890

Atom - Atom Distances (Å)
  C1 C2 O3 H4 H5 H6 H7
C11.51421.20901.12142.15772.15432.1543
C21.51422.41402.23131.10131.10541.1054
O31.20902.41402.02512.57873.12423.1242
H41.12142.23132.02513.13102.56492.5649
H52.15771.10132.57873.13101.80811.8081
H62.15431.10543.12422.56491.80811.7800
H72.15431.10543.12422.56491.80811.7800

picture of Acetaldehyde state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 H5 110.166 C1 C2 H6 109.657
C1 C2 H7 109.657 C2 C1 O3 124.489
C2 C1 H4 114.870 O3 C1 H4 120.641
H5 C2 H6 110.033 H5 C2 H7 110.033
H6 C2 H7 107.245
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability